Announcing AI-Native Market Access Product for Regulated Products

Learn More

Blogs

MedTech

Medical device audits - preparation and responses

By

Wendy Levine

May 2, 2023

4 min read

The word “audit” can strike panic in poorly prepared medtech companies. However, audits serve an important purpose in ensuring a compliant and effective quality system and production of safe and effective medical devices. And organizations can limit the stress and risk around audits through proper preparation. 

The key to a positive audit is to ensure that your organization’s focus is on building and implementing quality processes and procedures that cover the entire product life cycle and are continuously evaluated and improved upon. Not only is it the right thing to do, but focusing too closely on simply passing an inspection or audit may leave gaps in your processes and present a false sense of compliance. This article covers audit basics, how to prepare for them, and what to do when you receive an audit finding.

What is an audit?

Per ISO 19011 an audit is a systematic documented and independent process for obtaining objective evidence and evaluating it objectively to determine the extent to which the audit criteria are fulfilled. Audits can be internally conducted, externally conducted by interested parties (i.e., customers/ suppliers), and externally conducted by government agencies and notified bodies to ensure that product design, manufacturing, safety, and documentation requirements are being met. Audits will verify compliance with regulatory and quality system/GxP (Good Manufacturing Practices, Good Distribution Practices, etc.) requirements. GxP standards are dictated by the US FDA, European Medicines Agency (EMA), the UK Medicines and Healthcare Products Regulatory Agency (MHRA), and other regulatory bodies which rely on country-specific regulations as well as standards developed by the International Organization for Standardization (ISO). 

Audits are required regardless of device class, but audit requirements in the EU and US, along with most other markets, can be dependent on the device classification. For most medium to high-risk devices in the US and EU, the following audits take place:

  • Audits by EU Notified Bodies: Audits by EU Notified Bodies focus on compliance with MDR 2017/745 or IVDR 2017/746. Notified Bodies are also responsible for certifying quality management systems (QSR) against the requirements of ISO 13485:2016. Periodic “surveillance audits” will also be performed, based on the classification of the medical device(s).
  • FDA Inspections: The FDA will conduct inspections to ensure compliance with the quality system regulation, 21 CFR 820, and to confirm that a facility is capable of manufacturing the medical device. The FDA will conduct pre-approval inspections to verify data included in a market submission, along with periodic routine inspections, following the Quality System Inspection Technique (QSIT) as required by regulation (currently every two years for Class II and Class III USA-based device manufacturers and every five years for international device manufacturers).
  • Unannounced and “for cause” inspections: Manufacturers in the US and EU, and many other markets, are subject to different types of inspections triggered by consumer complaints, reported non-conformities, or other issues. These “for cause” inspections may be scheduled or unannounced.

How to prepare for an inspection

Audit preparation is a continuous process that should be built into your quality system and regulatory processes. Some items to consider:

Internal Quality audits

The best way to prepare for an upcoming audit or inspection is to use the internal audit program to your benefit. The FDA QSR, FDA 21 CFR 820, calls for medical device manufacturers to perform regular internal audits of their systems and to provide evidence of these audits and their effectiveness. When possible, conduct internal audits as if you’re the regulatory body and take them seriously. Internal audits should find the issues before the regulators do. Issue nonconformances and address them in a timely manner.

Performing “mock” audits is another great way to prepare for external inspections/audits from the FDA, notified bodies, and other regulatory authorities. Mock audits are a rehearsal for your team to prepare them for the real thing. They can act as try-outs to determine who is equipped to handle being audited and those that are too nervous or offer too much information when asked a question, requiring additional training. Mock audits are typically separate from the internal audit program since they are conducted based on different objectives and for training purposes.

It’s common to contract an independent third party to perform mock audits. Consider conducting unannounced mock audits to get the truest picture of your company’s preparedness. In short, the tougher medical device manufacturers are on themselves while preparing for the audit, then the less stressful the actual audit will be.

Self-identify issues as they appear and do not wait for the internal audit. If an issue is identified during the audit preparation or mock audit, implement corrective and preventive actions (CAPA) to address the issue. This is vital to demonstrate that you are aware of an issue and have begun remediation or corrective actions if and when those issues are uncovered during the real inspection or audit.

Choose the right audit host

When you have an upcoming audit or inspection, you must choose the right company representative to host the auditor(s). The person you choose will represent your company, so be deliberate about selecting those who know the company, its quality management system, and its products well. It should also be someone you’re confident can perform well under pressure and remain mission-focused in managing the audit and not necessarily answering every question immediately. The audit host can significantly impact the audit for the better or worse, so be certain that you have the right person in place who will be able to represent the organization’s values and facilitate an efficient audit.

While the person or people working directly with the auditor(s) are often from your quality team, they will need to be supported by subject matter experts (SMEs) from other functions for the duration of the audit – this will include the regulatory, engineering, operations, and marketing teams – who can answer specific questions and gather requested documents. These SMEs must be pre-identified along with alternates as part of the audit preparation. They should be comfortable facing an auditor and answering the auditor’s questions.

Gather all the necessary documents

As part of the audit process, the auditor(s) will expect access to information that they need to determine your organization’s compliance with all quality system and regulatory requirements. Based on the requirements, audit guidance, and previous audits, commonly requested documents should be known. This documentation should be pre-identified, compliant, and available before the start of an audit. This can be in the form of hard copies or electronically through files or links. The goal is to have documents readily available to avoid audit delays.

"If it takes too long to get documents to the auditor when they ask for them, you’re not making a good overall impression that everything is under control, making things more difficult for the auditor(s). Auditors have schedules to meet and follow certain audit trails. The last thing you want is your auditor getting agitated because they are spending a lot of time waiting for information." - Bruce McKean, Rimsys Director of Regulatory Affairs

It is critical that all regulatory information related to your products is readily available during an audit, such as registration status, certificates, regulatory impact assessments, and essential principles, along with submission content and post-market data. A central RIM system that stores all regulatory data and links to (or references) the current versions of records from other systems, such as PLM, eQMS, and ERP systems, can smooth the audit process significantly.

During an audit

As an organization, you will want to manage as much of the audit process as possible. Your audit host will greet the auditor(s) and give them a brief overview or presentation of your company, and most likely conduct a facility tour. After this, while the auditor(s) will direct the process, the more your host can assist and guide them, the better.

In the case of unannounced inspections/audits, there must be a procedure in place that defines how to receive and handle these types of audits. This will include who is the primary contact during such an inspection (often a Quality Management team member or representative), as well as Executive Management, and alternates when those people are not available.

Ideally, you should have more than one company representative with the auditor(s) during the audit and auditors should not be left alone at any point. Most companies have a team in the “front room” with the auditor(s) led by the audit host. The main job of this team is to transcribe every question, answer, and activity that occurs during the audit. The “front room” team will communicate with other team members in the “back room” in real-time (often via instant messaging), relaying to them any open questions, requested documents, or queuing up SMEs the auditor(s) need to speak with.

Best practices for sharing information with auditors

During an audit, employees should be cooperative and helpful, but should only share information that is specifically requested by the auditor. If information is requested that seems outside the scope of the audit, such as corporate strategic or financial documents, employees should notify the appropriate executive before providing such information.

Auditor(s) should be given access to requested information through photocopies or limited computer system access. Original documents can be presented if requested, but should never be kept by the auditor(s). All information provided should be prepared, verified, and recorded in the “back room” and then passed through to the audit host so that it can be controlled. The “back room” should mark the copies “Confidential” or “Proprietary,” as appropriate. They should also make an extra copy for the audit file, so the exact documentation given to the auditor(s) is known for future reference.

Addressing missing or incorrect information

Ideally, any potential issues with the existing quality system and related procedures are identified before an audit and corrective actions are identified and put in place. Even in cases where an issue has not been fully resolved, being able to point to awareness and appropriate actions is important.

Some findings may be able to be corrected during the audit. These findings are typically isolated issues (one-offs) that do not pose significant risks. For instance, a missing revision number, missing signature, or outdated reference. If corrected during the audit, it may negate a finding, but the auditor may want to understand why the issue occurred and what actions you have or will be, taking to ensure that it does not recur.

In cases where you are unable to produce the information requested by an auditor, or when there are questions about the validity or accuracy of the information, your internal team should acknowledge the issue but should not immediately speculate on the cause or the effect of the missing or inaccurate information. A discussion of appropriate actions under the existing quality system may be appropriate.

What to do in case of a finding

Be prepared to receive findings from any inspection. Ideally, the auditors should be working to ensure that you are compliant with regulatory requirements and that your records accurately state what you do. However, “By the nature of the beast,” says Bruce McKean, “they’re there to find instances of noncompliance.” This means that auditors will be focused on documentation that can prove or disprove adherence to your stated procedures and policies.

All findings should be disclosed before the audit closing meeting. There should be no surprises. Ensure that the findings are understood by both parties. If they are not clear, perhaps the auditor misunderstood or did not see specific objective evidence and you should discuss or review the issue with the auditor as this may negate a finding. Be sure to debrief upper management before the closing meeting. At the audit closing meeting, there should be no debate over findings. Any finding, whether major or minor, should be addressed diligently.

Audit findings or observations will result in the regulatory body in charge of the audit issuing a document that lists those findings. In most cases, you will have limited time to respond with a satisfactory plan for correcting and preventing the recurrence of the identified issues.

In the case of the FDA, multiple enforcement actions are available to the agency, ranging from warning letters to criminal prosecution. Note that many regulatory agencies will not respond further to your actions if they agree with the actions you prescribe for addressing audit observations. However, additional actions may be triggered if your response is not found to be satisfactory.

Rimsys is a holistic regulatory information management system designed for and by regulatory affairs professionals. Rimsys makes it easier to create and track submissions, keep up with product registrations and certificates, and even share pertinent data across ERP, PLM, and eQMS software platforms to ensure data integrity. Learn more about how Rimsys can help you face audits with the confidence that you have all of your regulatory ducks in a row.

‍

Similar posts

AI

From Regulatory Knowledge to Better Market Decisions: A Conversation with Steve Gens and James Gianoutsos

October 1, 2026

4 min read

AI is moving quickly from experimentation to implementation across life sciences. For regulatory teams, that raises a bigger question than where AI can automate a task:

How can organizations use AI to make better decisions while maintaining the traceability, expert oversight, and confidence regulatory work demands?

That was a central theme when James Gianoutsos, Founder and CEO of Rimsys, joined Steve Gens, Founder and Managing Partner of Gens & Associates, on the Regulatory Executive Podcast.

Their conversation explored the changing role of regulatory intelligence, the introduction of Rimsys Market Access, the importance of transparency in regulatory AI, and what these changes could mean for regulatory teams.

Regulatory knowledge has been difficult and costly to scale

For global medtech organizations, regulatory knowledge has traditionally been highly dependent on people.

A company may have specialists responsible for particular countries or regions, along with local representatives, distributors, consultants, and other experts. Those people develop deep knowledge of local regulations, pathways, and expectations.

The problem is that the knowledge often remains distributed across the organization. And maintaining that market-by-market expertise can become expensive as a company expands its portfolio and geographic footprint.

As James described during the podcast, large manufacturers may rely on dozens or even hundreds of internal employees, third-party distributors, local representatives, and other specialists to understand what it takes to bring products into different markets. Smaller organizations may rely more heavily on outside consultants and partners. Either way, the model can be costly and difficult to scale. 

But the cost isn't limited to what companies spend on expertise.

Steve described the workflow as a series of “start, stop” cycles: ask a local expert, verify whether the regulation is current, gather more information, and repeat. Each cycle consumes regulatory capacity and extends the time it takes to give the business a confident answer.

That creates another potentially more consequential expense: the cost of waiting.

When it takes months to understand what entering a market will require, commercial and product teams have less visibility into when revenue might begin, what resources will be needed, or whether the opportunity is worth pursuing. Regulatory teams spend valuable capacity assembling and validating information rather than applying their expertise to more strategic work.

Ultimately, scaling regulatory knowledge isn't simply an efficiency problem. It affects how quickly and confidently the business can decide where to invest.

The bigger question: Should we enter this market?

That challenge is central to Rimsys Market Access.

Most market-entry research begins with questions such as: How is the product classified? What's the regulatory pathway? What are the requirements?

Those questions establish whether a company can enter a market. But they don't necessarily tell the business whether it should.

Market Access is designed to help regulatory and commercial teams develop an earlier view of what market entry could involve, including classification, regulatory pathway, timeline, cost considerations, requirements and potential gaps. 

For commercial and product leaders, that provides greater predictability before committing budget and resources.

For regulatory leaders, it means starting with a structured plan rather than assembling every market assessment from scratch.

Moving regulatory knowledge from individuals into a shared system

During the conversation, James described one of AI's most significant opportunities as moving knowledge that has historically lived in people's heads into software.

The goal isn't to eliminate regulatory expertise. It's to make that expertise more scalable.

Market Access works from a curated regulatory corpus, allowing market-specific knowledge to be applied across workflows without requiring every initial question to go through the individual who happens to know that jurisdiction. Outputs are designed to trace back to the underlying regulation, guidance, or law. 

That can also change where regulatory professionals spend their time. Rather than focusing as heavily on finding information and assembling initial assessments, they can focus on reviewing the evidence, identifying gaps, applying judgment, and advising the business.

Steve described the concept as a subject-matter-expert assistant. James called it a workforce multiplier. 

Trust requires opening the regulatory AI “black box”

Speed isn't enough when the information is being used to inform regulatory decisions.

James and Steve returned repeatedly to the importance of knowing where an AI-generated answer comes from. As Steve summarized it during the discussion, transparency builds trust.

Market Access is designed around that principle. Its outputs are grounded in a curated regulatory corpus and traceable to the specific regulations, guidance, or laws supporting them. That gives regulatory professionals the ability to verify an answer rather than simply accept it. 

Human judgment remains essential as well. Market Access provides an upfront assessment, not a guarantee of a regulatory outcome. Reviewer interpretation and other factors can still affect what happens during the regulatory process.

The objective isn't to make uncertainty disappear. It's to make what is known, what the sources say, and where uncertainty remains much easier to see.

Connecting the market decision to execution

The discussion also pointed to a broader evolution in regulatory technology: moving from systems that primarily store regulatory information toward systems that help teams act on it.

That's where Market Access and Rimsys RIM play distinct but complementary roles.

Market Access is the planning layer. It helps teams determine whether, when, and how to enter a market.

Rimsys RIM is the execution layer. It manages the registrations, submissions, regulatory changes, and ongoing regulatory work required to act on that plan. 

Together, they create a path from the initial market decision through regulatory execution rather than leaving planning and execution disconnected.

A new role for AI in regulatory work

One of the most interesting themes from Steve and James's conversation was that AI's impact may ultimately be less about replacing individual tasks and more about expanding what regulatory teams can accomplish.

Better access to market-specific knowledge can help professionals work across more markets, spend less time gathering information, and devote more capacity to the judgment and strategic work that requires their expertise.

The technology is changing quickly, but the fundamentals of regulatory work remain: reliable information, domain expertise, transparency, and human judgment.

The opportunity is to apply those fundamentals at greater scale while giving the business a clearer answer to the question that comes before execution:

Not simply, “Can we enter this market?”

But, “Should we?”

Listen to the full conversation

Hear the complete discussion between Steve Gens, Founder and Managing Partner of Gens & Associates, and James Gianoutsos, Founder and CEO of Rimsys, on the Regulatory Executive Podcast for more on regulatory AI, market-entry planning, transparency, and the changing role of regulatory teams.

‍

AI

How Market Access Builds Trust in Regulatory Intelligence

By

Bethaney Lentz

September 18, 2026

4 min read

Why source traceability, regulatory specificity, and human judgment matter when AI informs a market-entry decision.

Regulatory professionals are not short on AI options. General-purpose tools can answer questions quickly and often sound convincing. But in medtech, a convincing answer is not the same thing as a decision you can defend.

That distinction matters most before a company commits to a new market. Teams need to understand how a product is classified, which pathway applies, how long entry may take, what it may cost, where the important gaps are, and what uncertainty still remains. Commercial leaders need that visibility to decide whether the opportunity is worth pursuing. Regulatory leaders need to know that the assessment is grounded in sources they can verify.

That is the trust standard behind Rimsys Market Access. It is not simply about producing regulatory information faster. It is about helping medtech teams decide whether, when, and how to enter a market before committing budget and resources, with outputs linked back to the regulations, guidance, and laws behind them.

Why fluency is not enough for regulatory work

A general AI tool can be useful for brainstorming, summarizing, and exploring a topic. The challenge comes when an answer has real regulatory consequences. A plausible response may still be incomplete, based on the wrong designation, or disconnected from the current source that should govern the decision.

Medtech makes this especially difficult because the same product can be treated differently across markets and product designations. Medical device, IVD, cosmetic, consumer, OTC, and combination-product considerations can change the regulatory read. A market-entry assessment has to account for that context rather than assume one answer travels cleanly from one jurisdiction to another.

For regulatory teams, the natural next question is: Where did this answer come from? If the system cannot show the underlying authority, the expert still has to rebuild the research before trusting the result. That limits the value of the speed AI was supposed to create in the first place.

Trust starts with a decision you can verify

Market Access is designed around a different outcome. The goal is not to give teams another stream of regulatory information to interpret. It is to provide a source-linked market plan that helps them answer the bigger business question: should we enter this market, not just can we?

For a market assessment, that can include classification, regulatory pathway, timeline, cost considerations, key requirements and potential gaps. The output is designed to give commercial and product leaders an earlier, more defensible view of the opportunity while giving regulatory professionals the detail they need to review the basis for the assessment.

Trust comes from being able to inspect that basis. Market Access works from a curated regulatory corpus spanning more than 200 markets rather than relying on the open web. Answers are linked to the specific regulation, guidance, or law behind them, so a regulatory professional can verify the source instead of being asked to accept an AI-generated conclusion on faith.

Three things make regulatory intelligence more defensible

1. Source traceability

A regulatory answer is more useful when the person reviewing it can follow the evidence. Market Access links outputs back to their underlying regulatory sources. That changes the review process from “Do I trust the AI?” to a more practical question: “Does the cited source support this conclusion for our product and situation?”

That is an important shift for regulatory teams. Instead of assembling every market assessment from a blank page, they can review a structured, sourced plan, challenge assumptions, and focus their expertise on the places where judgment matters most.

2. Regulatory specificity

Trust also depends on context. Market Access is purpose-built for medtech regulatory complexity and supports cross-designation classification. The objective is to evaluate the product in the context of the market and the applicable regulatory framework, then carry that context into the pathway, requirements, timeline, cost considerations, and gaps that shape the market-entry decision.

This is where a purpose-built market-planning platform differs from a generic chat experience. The value is not an impressive answer in isolation. It is a connected assessment that helps the team understand what entering the market is likely to require and whether the opportunity still makes sense once those requirements are visible.

3. Human judgment stays in control

No system can remove every area of uncertainty from regulatory work. Reviewer interpretation and expert judgment still affect outcomes, and Market Access is not designed to promise approval.

Instead, Market Access is designed to make uncertainty more manageable. Regulatory professionals review and validate the work before it moves into execution. They can verify sources, assess the gaps, and apply company and product context that software cannot fully replace. The technology accelerates the research and planning. The regulatory professional remains responsible for the judgment.

What trust looks like in a market-entry decision

Imagine a team evaluating several markets for a new product. The commercial question is not simply whether registration is technically possible. The team needs to know which markets deserve investment now, which may require more evidence or resources, and which could put the revenue window at risk because the pathway is longer or more complex than expected.

Regulatory has a different, but connected, need. They have to understand the classification and pathway, identify the requirements most likely to slow the submission, and confirm that the plan reflects the relevant regulatory sources.

Market Access brings those views together. Commercial and product leaders get earlier visibility into timing, cost considerations, requirements, and risk so they can build a stronger business case. Regulatory teams get a source-linked plan they can review rather than a collection of unsupported conclusions. Both groups are working from the same market-entry picture before significant resources are committed.

Trust is not the end goal. Better decisions are.

It is easy for a conversation about regulatory AI to become a conversation about models, agents, or architecture. Those details matter, but they are not the business outcome.

The real value of trustworthy regulatory intelligence is what it lets a team do next. It can help a commercial leader defend a market investment. It can help regulatory surface issues earlier. It can give both groups a more predictable view of the work ahead before budget, R&D capacity, or launch timing is locked in.

And once the decision is made, the work does not have to stop at an intelligence report. Market Access is the market-planning layer. Rimsys RIM is the execution layer for registrations, submissions, regulatory changes, and ongoing regulatory work. Each can stand on its own, and together they create a path from the market-entry decision into controlled execution.

A more useful standard for regulatory AI

The question for medtech teams should not be whether AI can produce a regulatory answer. It can. The better question is whether that answer is specific enough, traceable enough, and reviewable enough to support a real decision.

Market Access is built around that standard: a defensible view of whether, when, and how to enter a market, grounded in curated regulatory sources and kept under human review. Because when the decision affects revenue, resources, and regulatory strategy, sounding right is not enough. Teams need to see why the answer is right, where uncertainty remains, and what it means for the decision in front of them.

Company

Product Updates

Rimsys Announces AI-Native Market Access for Regulated Products

By

Bethaney Lentz

September 1, 2026

4 min read

New software gives regulatory and commercial leaders a source-linked, actionable view of market viability, requirements, timing, and risk before committing resources

PITTSBURGH, PA, September 1, 2026 - Rimsys, the heart of medtech regulatory operations, today announced Market Access, a new AI-native product designed to help companies determine whether a market is worth entering and what it will take to get there. Market Access will be available in beta beginning September 1, 2026, with general availability to be announced separately.

Market Access replaces what can be a 6-to-9-month research and assessment process with a same-day, actionable read on market viability. It gives regulatory and commercial leaders the classification, pathway, timeline, cost considerations, and potential regulatory gaps they need to make a more defensible investment decision before committing budget and resources.

Predictability is at the core of Market Access. Rather than simply identifying a regulatory pathway, it helps teams understand what to expect, which requirements or gaps are most likely to slow down a submission, and where uncertainty remains. Every answer traces back to the specific regulation, guidance, or law behind it, allowing regulatory professionals to verify the underlying source rather than simply trust an AI-generated response.

Key features and benefits include:

  • Same-day market assessment: Provides an actionable read on whether a market is worth entering, replacing research and assessment that can take 6 to 9 months.
  • Cross-designation classification: Evaluates products across medical device, IVD, cosmetic, consumer, and OTC designations, so the same product receives the appropriate regulatory read regardless of how a given market classifies it.
  • Clear market-entry requirements: Provides classification, regulatory pathway, timeline, cost considerations, and key requirements before teams commit resources.
  • Earlier visibility into regulatory gaps: Surfaces the specific requirements and gaps most likely to slow down a submission and helps teams understand their potential severity.
  • Greater predictability: Provides visibility into anticipated regulatory response windows, risks, and areas of uncertainty so teams can better plan for what lies ahead.
  • Source-linked intelligence: Traces answers to the underlying regulations, guidance, and laws so regulatory professionals can verify the basis for an assessment.
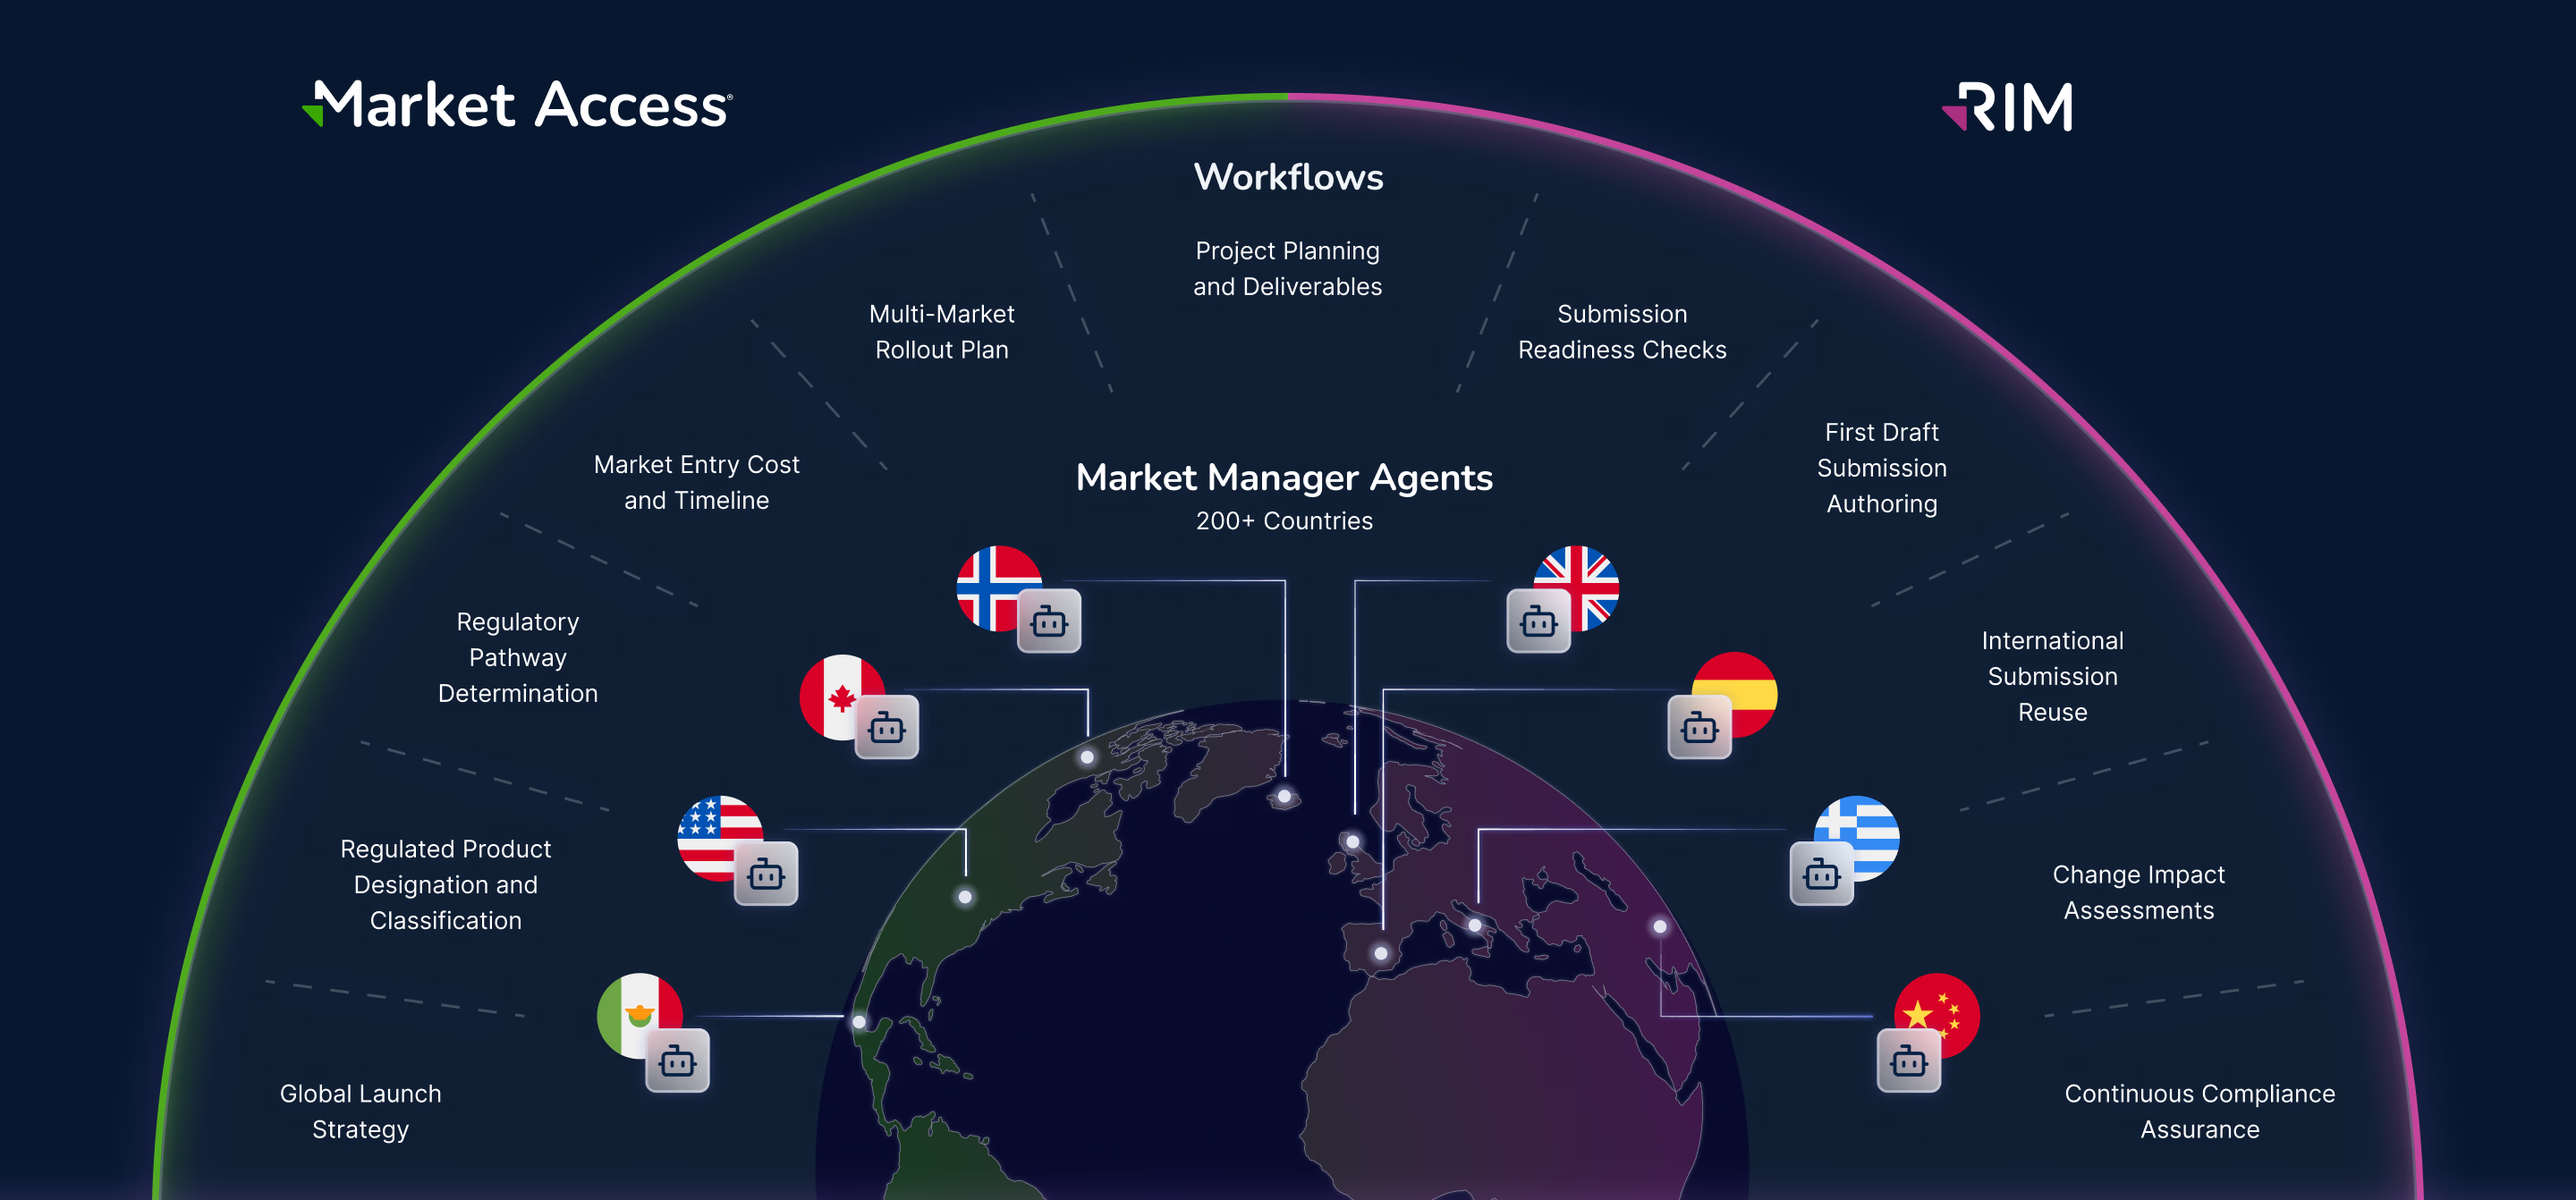
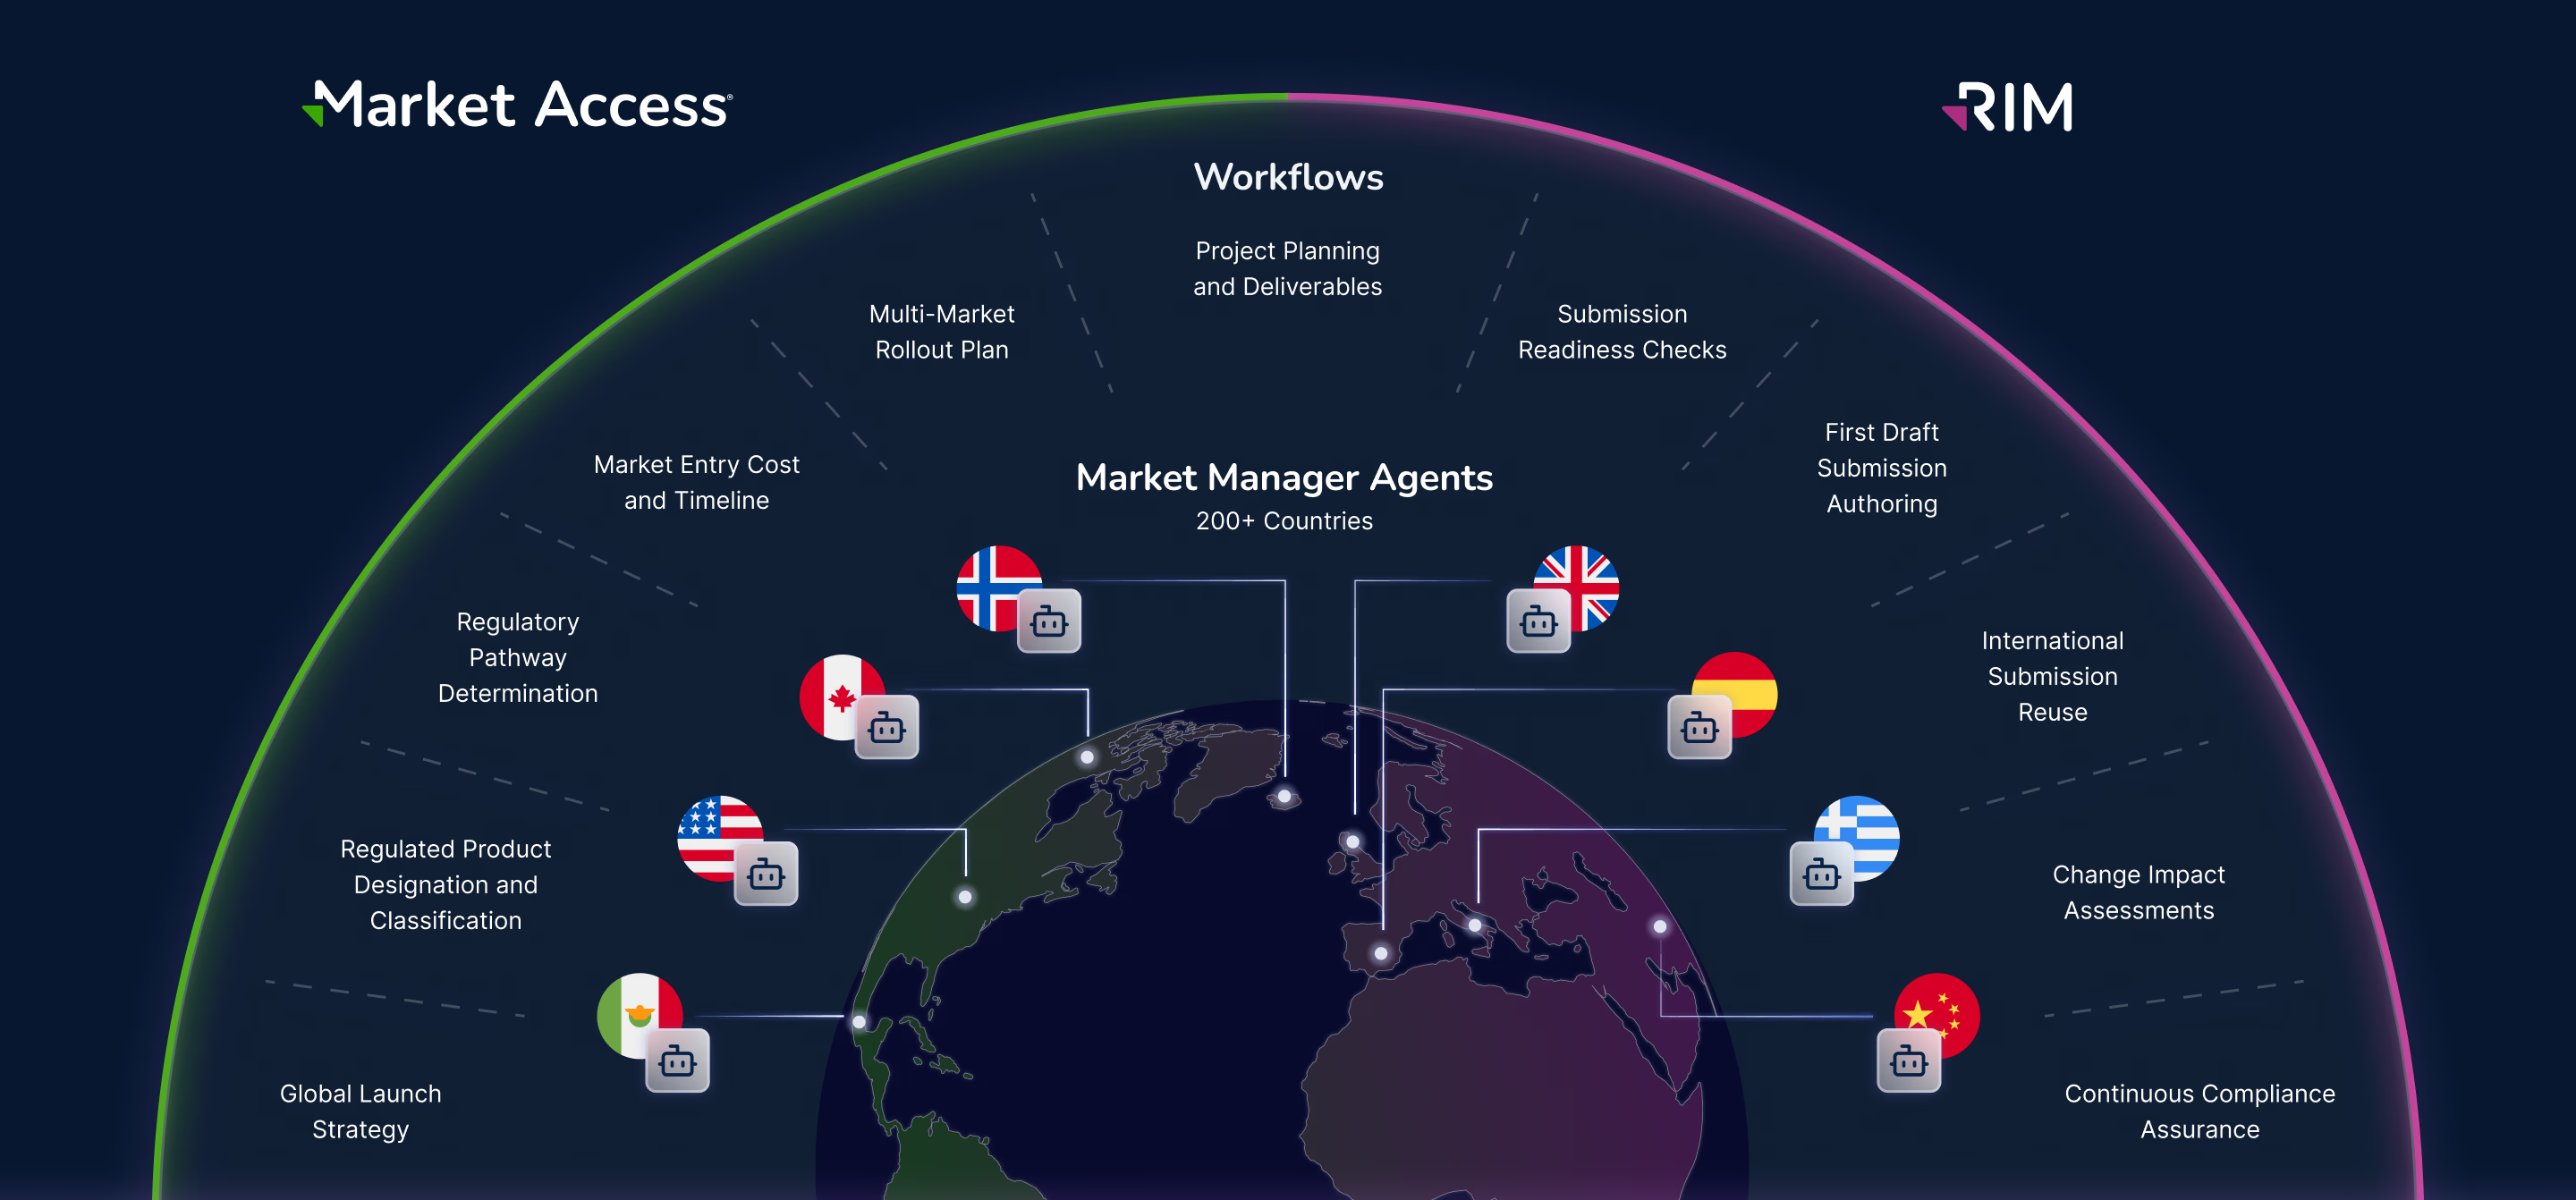
  • AI with human oversight: Uses Rimsys AI and Market Manager Agents to accelerate complex regulatory research while keeping regulatory professionals in control of review and validation.
  • Connected planning and execution: Complements RIM, connecting market-entry decisions with the registrations, submissions, regulatory changes, and ongoing compliance required to execute them.

“Regulatory knowledge has traditionally been highly dependent on individual expertise and market-by-market research,” said James Gianoutsos, Founder and CEO of Rimsys. “Market Access gives companies greater predictability before they make a market investment. Teams can understand whether an opportunity is worth pursuing, what it will take to enter, where regulatory challenges are likely to emerge, and what uncertainty remains. And because every answer is tied to its regulatory source, experts can verify the recommendation rather than simply trust the AI.”

Market Access is powered by Rimsys AI, which works from a curated regulatory corpus spanning more than 200 markets, not the open web. Its Market Manager Agents are AI agents with market-specific regulatory knowledge, including pathways, requirements, and documentation, that support market assessments and global launch planning. The product is also designed to acknowledge areas where reviewer interpretation and expert judgment can affect regulatory outcomes rather than presenting those outcomes as more predictable than they are.

Market Access expands the Rimsys product portfolio from regulatory execution into market planning. Market Access helps regulatory and commercial leaders determine where to compete, while RIM manages the registrations, submissions, regulatory changes, product data, and ongoing compliance required to enter and remain in those markets. Together, they turn regulatory intelligence into coordinated action, helping regulatory teams accomplish more in every hour while giving business leaders greater visibility into opportunities, risk, and the work required to protect and grow revenue.

Beta Availability

Market Access will be available in beta beginning September 1, 2026. During the beta period, Rimsys will continue validating product capabilities and refining workflows ahead of broader commercial availability. 

Organizations interested in learning more about Market Access or participating in the beta program can visit rimsys.io/products/market-access.

About Rimsys

Rimsys is the heart of regulatory operations, connecting AI-native market planning with AI-embedded regulatory execution, giving medtech manufacturers one living system from market-entry decision to execution. 

Rimsys Market Access is the planning layer, answering not just whether a market can be entered, but whether it should be, returning classification, pathway, timeline, and cost before a team commits budget or resources to finding out. Rimsys RIM is the execution layer, managing registrations, submissions, regulatory changes, and compliance so teams can act on that plan with confidence.

Trusted by 6 of the top 12 global MedTech manufacturers and certified to ISO 27001 and SOC 2, with ISO 42001 certification for AI management systems underway, Rimsys keeps regulatory intelligence, product data, approvals, and change management continuously connected so teams can enter new markets with speed and assurance. Learn more at rimsys.io.

Media Contact
letschat@rimsys.io

I agree to the privacy policy including to Rimsys using my contact details to contact me for marketing purposes.
Thank you! Your submission has been received!
Oops! Something went wrong while submitting the form.
Hand holding smartphone showing email app with 12 unread messages notification.