Announcing AI-Native Market Access Product for Regulated Products

Learn More

Blogs

MedTech

The ultimate guide to the EU MDR/IVDR UDI

By

Bethaney Lentz

June 22, 2021

4 min read

This article is an excerpt from The ultimate guide to the EU MDR/IVDR UDI ebook.

Table of contents

Overview

The EU Medical Device Regulation (2017/745) (“MDR”) and EU In Vitro Diagnosis Regulation (2017/746) (“IVDR”) introduce two new systems for information exchange: UDI (Unique Device Identifier) for device identification and EUDAMED (European Databank on Medical Devices) to centralize and disseminate information. UDI is a specific code assigned to all devices and higher levels of packaging. This will allow for devices being sold in the European market to be identified and traced through a globally harmonized approach. EUDAMED is the IT system developed by the European Commission to replace the EUDAMED2 database previously in place under the Medical Device Directives (MDD). EUDAMED is a multi-functional system that will be used to coordinate device registration, provide information about devices to industry professionals and the public, and highlight necessary safety details.

UDI basics and benefits

The EU MDR and IVDR UDI system is based upon the guidance of the International Medical Device Regulators Forum (IMDRF). It’s a globally harmonized system that’s designed to increase patient safety and optimize care.

UDI system goals

Increase patient safety

  • Improve tracing of devices
  • Reduce the presence of counterfeit devices

Ensure access to accurate information

  • Unambiguous identification of devices throughout distribution and use

Improve post-market surveillance

  • Improve accessibility of adverse event reports

Enhance supply chain Management

  • Streamline supply chain process and inventory management
  • Simplify medical device documentation processes

The UDI system has four key elements

Element 1 Element 2 Element 3 Element 4
Assignment of a UDI consisting of:
- Basic UDI-DI
- UDI-DI and UDI-PI
- Packaging UDI
Placing UDI on Device or Packaging through UDI Carrier Storage of UDI information by Economic Operators UDI Database to Access Information

Element 1: Assignment of UDI (UDI Components)

The first element of the UDI system is the assignment of a UDI. The UDI is a code of alphanumeric characters that acts as the access key to information about a specific medical device on the market. The EU MDR and EU IVDR requires that a UDI be assigned to all medical devices except for custom-made or investigational devices. There are three components of a UDI:

  • Basic UDI-DI
  • UDI (consisting of UDI-DI and UDI-PI)
  • Packaging UDI (Note: This is not an official term used in the EU MDR and IVDR, but we’re using it to help explain the concept. The Packing UDI is part of the UDI itself.)

1. Basic UDI-DI

The Basic UDI-DI identifies the device group that a particular device fits into. A device group is a group of products that all share the same intended purpose, risk class, essential design, and manufacturing characteristics. A device group is generally classified by medical device manufacturers as a “Product Family” or “Product Category,” depending on the internal nomenclature used within the company. The Basic UDI-DI functions as a parent or higher-level descriptor of a device.

NOTE: There can only be one Basic UDI-DI per UDI-DI.

The Basic UDI-DI is not printed on the product itself or on the packaging of a product, but rather it must be included in the following documents and applications:

  • Certificates (Including Certificate of Free Sale)
  • EU Declarations of Conformity
  • Techical Documentation
  • Summary of Safety and Clinical Performance

2. UDI (UDI-DI and UDI-PI)

The second component is the UDI itself, which consists of two parts:

Device Identifier (DI)

Production Identifier (PI)

The UDI-DI (Device Identifier DI, also referred to as “static”) identifies specific, detailed information about a particular device. If any of the below details should change, the device will need a new UDI-DI.

  • Name or trade name of the device
  • Device version or model
  • If labelled as a single use device
  • Packaged as sterile
  • Maximum number of uses
  • Need for sterilization before use
  • Quantity of devices provided in a package
  • Critical warnings or contra-indication
  • CMR/endocrine disruptors

NOTE: There can be several UDI-DIs for one Basic UDI-DI.

Meanwhile, the UDI-PI (Production Identifier PI, also referred to as "dynamic") contains manufacturing information (including serial number, lot/batch number, software identification, and manufacturing or expiry date or both types of dates.)

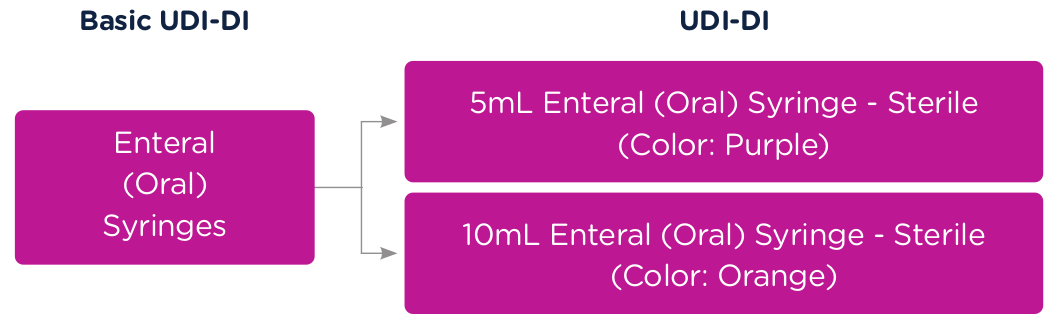
To better illustrate this concept of Basic UDI-DI and UDI (UDI-DI and UDI-PI), let’s use a syringe as an example. The Basic UDI-DI would identify the category of a syringe, for example, "Enteral (Oral) Syringe."

A 5ml Enteral (Oral) Syringe – Sterile (Color: Purple) would get a unique UDI-DI and a 10m Enteral (Oral) Syringe – Sterile (Color: Orange) would get a unique UDI-DI. Both products would be associated to the same Basic UDI-DI. In this case, the "Enteral (Oral) Syringe," which defines the category.
Each time that 5ml Enteral (Oral) Syringe – Sterile (Color: Purple) is manufactured at the same revision, it will get a new UDI-PI per lot. See the graphic below.

Each product is identical and therefore has the same UDI-DI. However, the UDI-PI changes to reflect the manufacturing date, lot number, expiry date, and serial number, as applicable.

The UDI will contain all device-specific information and have the same functions as the comparable database (GUDID) of the United States FDA. The main difference (in EUDAMED) is that the UDI data is divided into components of Basic UDI-DI, UDI, and Packaging UDI.

3. Packaging UDI

The third component of UDI is the Packaging UDI. (Note: This is not an official term used in the EU MDR and IVDR, but we’re using it to help explain the concept.)

Each level of packaging, except shipping containers, must receive its own unique UDI. Packaging UDI refers to the unique UDI assigned to higher levels of packaging instead of the device itself.

In the event of significant space constraints on the unit of use packaging, the UDI Carrier may be placed on the next higher packaging level.

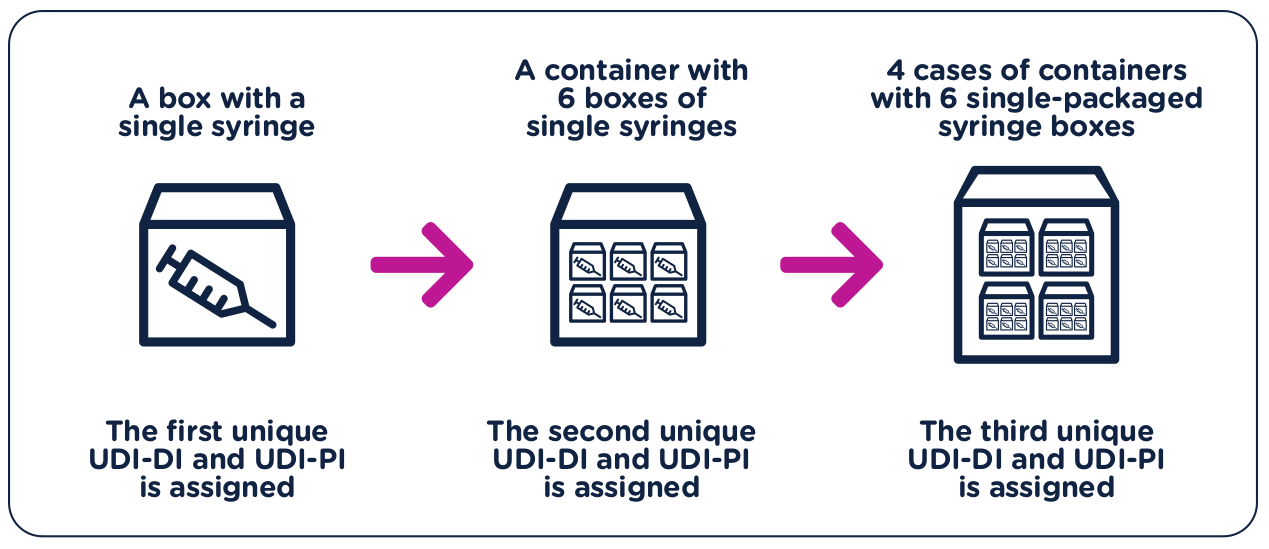
Returning to our earlier example of syringes, if a manufacturer first packages a single sellable syringe into an individual box, this package would receive its own UDI-DI and UDI-PI.

If then the manufacturer packages those individual boxes into containers of six (6), those containers would receive their own UDI-DI and UDI-PI.

And finally, if the manufacturer packages those six (6) containers into cases of four (4), those cases would receive their own UDI-DI and UDI-PI.

Each of those levels of packaging must be assigned its own UDI-DI and UDI-PI. The initial syringe did not change, but the way it is packaged did, therefore, requiring its own UDI-DI and UDI-PI.
Element 2: Placing UDI on the device and/or packaging

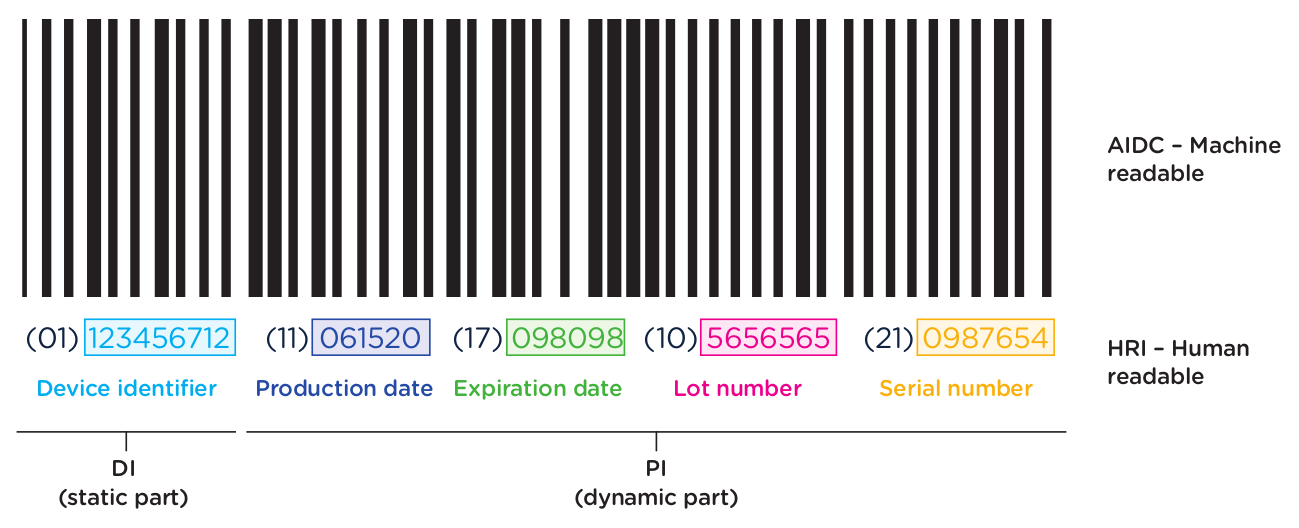
The second element to the UDI system is the placing of the UDI on the device or on its packaging through what is referred to as a “UDI Carrier.” The UDI Carrier is the part of the label that contains the UDI information that is applied directly to the device or included on the device packaging. The UDI Carrier should have both a machine-readable portion (AIDC) and a human-readable portion (HRI). (Specific details about each element of the UDI will be covered in Chapter 2.)

  • Machine-readable form – AIDC – (Automatic Identification and Data Capture) is a barcode or other machine-readable technology that can be accessed automatically by scanning the UDI information.
  • Human-readable form – HRI – (Human Readable Interpretation) is the numeric or alphanumeric code, which can be manually entered into the system for access to the UDI information.

If there are space constraints limiting the use of both the AIDC and HRI on the label, then only the AIDC is required to appear. However, on devices that are intended to be used in home-health care or other non-medical facility settings, the HRI would be required to appear.

Single-use devices may contain the UDI Carrier on its lowest level of packaging rather than on the device itself.

Reusable devices must include the UDI Carrier on the device itself, unless any type of direct marking would interfere with the safety or performance of the device, or if it is not technologically feasible to directly mark the device. If so, this should be properly documented in your design history file.

Most importantly, the UDI Carrier must be readable for the intended lifecycle of the device.

Below is an example of a GS1 AIDC and HRI barcode label.
Element 3: Storage of UDI information by Economic Operators

Storage of UDI information by "Economic Operators" is the third element of the UDI system. 2017/745 Articles 2(35), 22(1), and 22(3) define an economic operator as:

  • A manufacturer
  • An authorized representative
  • A distributor
  • An importer
  • An investigator for clinical investigations
  • A person who sterilizes systems or procedure packs

Class III, implantable device:

According to EU MDR 2017/745 Annex II, the manufacturer shall keep an updated list of all UDIs that it has assigned. Economic operators and all health institutions are required to store, preferably by electronic means, the UDI of all the devices for which they have supplied or with which they have been supplied.

For Devices Other than Class III:

Member States are encouraged, and in some cases require, health institutions to store, preferably by electronic means, the UDI of the devices with which they have been supplied. The UDI must also be included in any field safety notice for reporting serious incidents and field safety corrective actions.

The EU MDR and EU IVDR also give the European Commission authority to make additional requirements regarding the submission or maintenance of UDI information. In making those decisions, the European Commission must consider six (6) areas:

  • Confidentiality and data protection
  • Risk-based approach
  • Cost-effectiveness of the additional measures
  • The need to avoid duplications in the UDI system
  • The needs of the healthcare systems of the member states
  • Harmonization with other medical device identification systems

Element 4: The UDI Database

To continue reading this eBook including information about the EUDAMED database, UDI format requirements and issuing entities, implementation timelines, and key differences between the EU and US UDI systems, please register to download the full version
Similar posts

AI

How Market Access Builds Trust in Regulatory Intelligence

By

Bethaney Lentz

September 18, 2026

4 min read

Why source traceability, regulatory specificity, and human judgment matter when AI informs a market-entry decision.

Regulatory professionals are not short on AI options. General-purpose tools can answer questions quickly and often sound convincing. But in medtech, a convincing answer is not the same thing as a decision you can defend.

That distinction matters most before a company commits to a new market. Teams need to understand how a product is classified, which pathway applies, how long entry may take, what it may cost, where the important gaps are, and what uncertainty still remains. Commercial leaders need that visibility to decide whether the opportunity is worth pursuing. Regulatory leaders need to know that the assessment is grounded in sources they can verify.

That is the trust standard behind Rimsys Market Access. It is not simply about producing regulatory information faster. It is about helping medtech teams decide whether, when, and how to enter a market before committing budget and resources, with outputs linked back to the regulations, guidance, and laws behind them.

Why fluency is not enough for regulatory work

A general AI tool can be useful for brainstorming, summarizing, and exploring a topic. The challenge comes when an answer has real regulatory consequences. A plausible response may still be incomplete, based on the wrong designation, or disconnected from the current source that should govern the decision.

Medtech makes this especially difficult because the same product can be treated differently across markets and product designations. Medical device, IVD, cosmetic, consumer, OTC, and combination-product considerations can change the regulatory read. A market-entry assessment has to account for that context rather than assume one answer travels cleanly from one jurisdiction to another.

For regulatory teams, the natural next question is: Where did this answer come from? If the system cannot show the underlying authority, the expert still has to rebuild the research before trusting the result. That limits the value of the speed AI was supposed to create in the first place.

Trust starts with a decision you can verify

Market Access is designed around a different outcome. The goal is not to give teams another stream of regulatory information to interpret. It is to provide a source-linked market plan that helps them answer the bigger business question: should we enter this market, not just can we?

For a market assessment, that can include classification, regulatory pathway, timeline, cost considerations, key requirements and potential gaps. The output is designed to give commercial and product leaders an earlier, more defensible view of the opportunity while giving regulatory professionals the detail they need to review the basis for the assessment.

Trust comes from being able to inspect that basis. Market Access works from a curated regulatory corpus spanning more than 200 markets rather than relying on the open web. Answers are linked to the specific regulation, guidance, or law behind them, so a regulatory professional can verify the source instead of being asked to accept an AI-generated conclusion on faith.

Three things make regulatory intelligence more defensible

1. Source traceability

A regulatory answer is more useful when the person reviewing it can follow the evidence. Market Access links outputs back to their underlying regulatory sources. That changes the review process from “Do I trust the AI?” to a more practical question: “Does the cited source support this conclusion for our product and situation?”

That is an important shift for regulatory teams. Instead of assembling every market assessment from a blank page, they can review a structured, sourced plan, challenge assumptions, and focus their expertise on the places where judgment matters most.

2. Regulatory specificity

Trust also depends on context. Market Access is purpose-built for medtech regulatory complexity and supports cross-designation classification. The objective is to evaluate the product in the context of the market and the applicable regulatory framework, then carry that context into the pathway, requirements, timeline, cost considerations, and gaps that shape the market-entry decision.

This is where a purpose-built market-planning platform differs from a generic chat experience. The value is not an impressive answer in isolation. It is a connected assessment that helps the team understand what entering the market is likely to require and whether the opportunity still makes sense once those requirements are visible.

3. Human judgment stays in control

No system can remove every area of uncertainty from regulatory work. Reviewer interpretation and expert judgment still affect outcomes, and Market Access is not designed to promise approval.

Instead, Market Access is designed to make uncertainty more manageable. Regulatory professionals review and validate the work before it moves into execution. They can verify sources, assess the gaps, and apply company and product context that software cannot fully replace. The technology accelerates the research and planning. The regulatory professional remains responsible for the judgment.

What trust looks like in a market-entry decision

Imagine a team evaluating several markets for a new product. The commercial question is not simply whether registration is technically possible. The team needs to know which markets deserve investment now, which may require more evidence or resources, and which could put the revenue window at risk because the pathway is longer or more complex than expected.

Regulatory has a different, but connected, need. They have to understand the classification and pathway, identify the requirements most likely to slow the submission, and confirm that the plan reflects the relevant regulatory sources.

Market Access brings those views together. Commercial and product leaders get earlier visibility into timing, cost considerations, requirements, and risk so they can build a stronger business case. Regulatory teams get a source-linked plan they can review rather than a collection of unsupported conclusions. Both groups are working from the same market-entry picture before significant resources are committed.

Trust is not the end goal. Better decisions are.

It is easy for a conversation about regulatory AI to become a conversation about models, agents, or architecture. Those details matter, but they are not the business outcome.

The real value of trustworthy regulatory intelligence is what it lets a team do next. It can help a commercial leader defend a market investment. It can help regulatory surface issues earlier. It can give both groups a more predictable view of the work ahead before budget, R&D capacity, or launch timing is locked in.

And once the decision is made, the work does not have to stop at an intelligence report. Market Access is the market-planning layer. Rimsys RIM is the execution layer for registrations, submissions, regulatory changes, and ongoing regulatory work. Each can stand on its own, and together they create a path from the market-entry decision into controlled execution.

A more useful standard for regulatory AI

The question for medtech teams should not be whether AI can produce a regulatory answer. It can. The better question is whether that answer is specific enough, traceable enough, and reviewable enough to support a real decision.

Market Access is built around that standard: a defensible view of whether, when, and how to enter a market, grounded in curated regulatory sources and kept under human review. Because when the decision affects revenue, resources, and regulatory strategy, sounding right is not enough. Teams need to see why the answer is right, where uncertainty remains, and what it means for the decision in front of them.

Company

Product Updates

Rimsys Announces AI-Native Market Access for Regulated Products

By

Bethaney Lentz

September 1, 2026

4 min read

New software gives regulatory and commercial leaders a source-linked, actionable view of market viability, requirements, timing, and risk before committing resources

PITTSBURGH, PA, September 1, 2026 - Rimsys, the heart of medtech regulatory operations, today announced Market Access, a new AI-native product designed to help companies determine whether a market is worth entering and what it will take to get there. Market Access will be available in beta beginning September 1, 2026, with general availability to be announced separately.

Market Access replaces what can be a 6-to-9-month research and assessment process with a same-day, actionable read on market viability. It gives regulatory and commercial leaders the classification, pathway, timeline, cost considerations, and potential regulatory gaps they need to make a more defensible investment decision before committing budget and resources.

Predictability is at the core of Market Access. Rather than simply identifying a regulatory pathway, it helps teams understand what to expect, which requirements or gaps are most likely to slow down a submission, and where uncertainty remains. Every answer traces back to the specific regulation, guidance, or law behind it, allowing regulatory professionals to verify the underlying source rather than simply trust an AI-generated response.

Key features and benefits include:

  • Same-day market assessment: Provides an actionable read on whether a market is worth entering, replacing research and assessment that can take 6 to 9 months.
  • Cross-designation classification: Evaluates products across medical device, IVD, cosmetic, consumer, and OTC designations, so the same product receives the appropriate regulatory read regardless of how a given market classifies it.
  • Clear market-entry requirements: Provides classification, regulatory pathway, timeline, cost considerations, and key requirements before teams commit resources.
  • Earlier visibility into regulatory gaps: Surfaces the specific requirements and gaps most likely to slow down a submission and helps teams understand their potential severity.
  • Greater predictability: Provides visibility into anticipated regulatory response windows, risks, and areas of uncertainty so teams can better plan for what lies ahead.
  • Source-linked intelligence: Traces answers to the underlying regulations, guidance, and laws so regulatory professionals can verify the basis for an assessment.
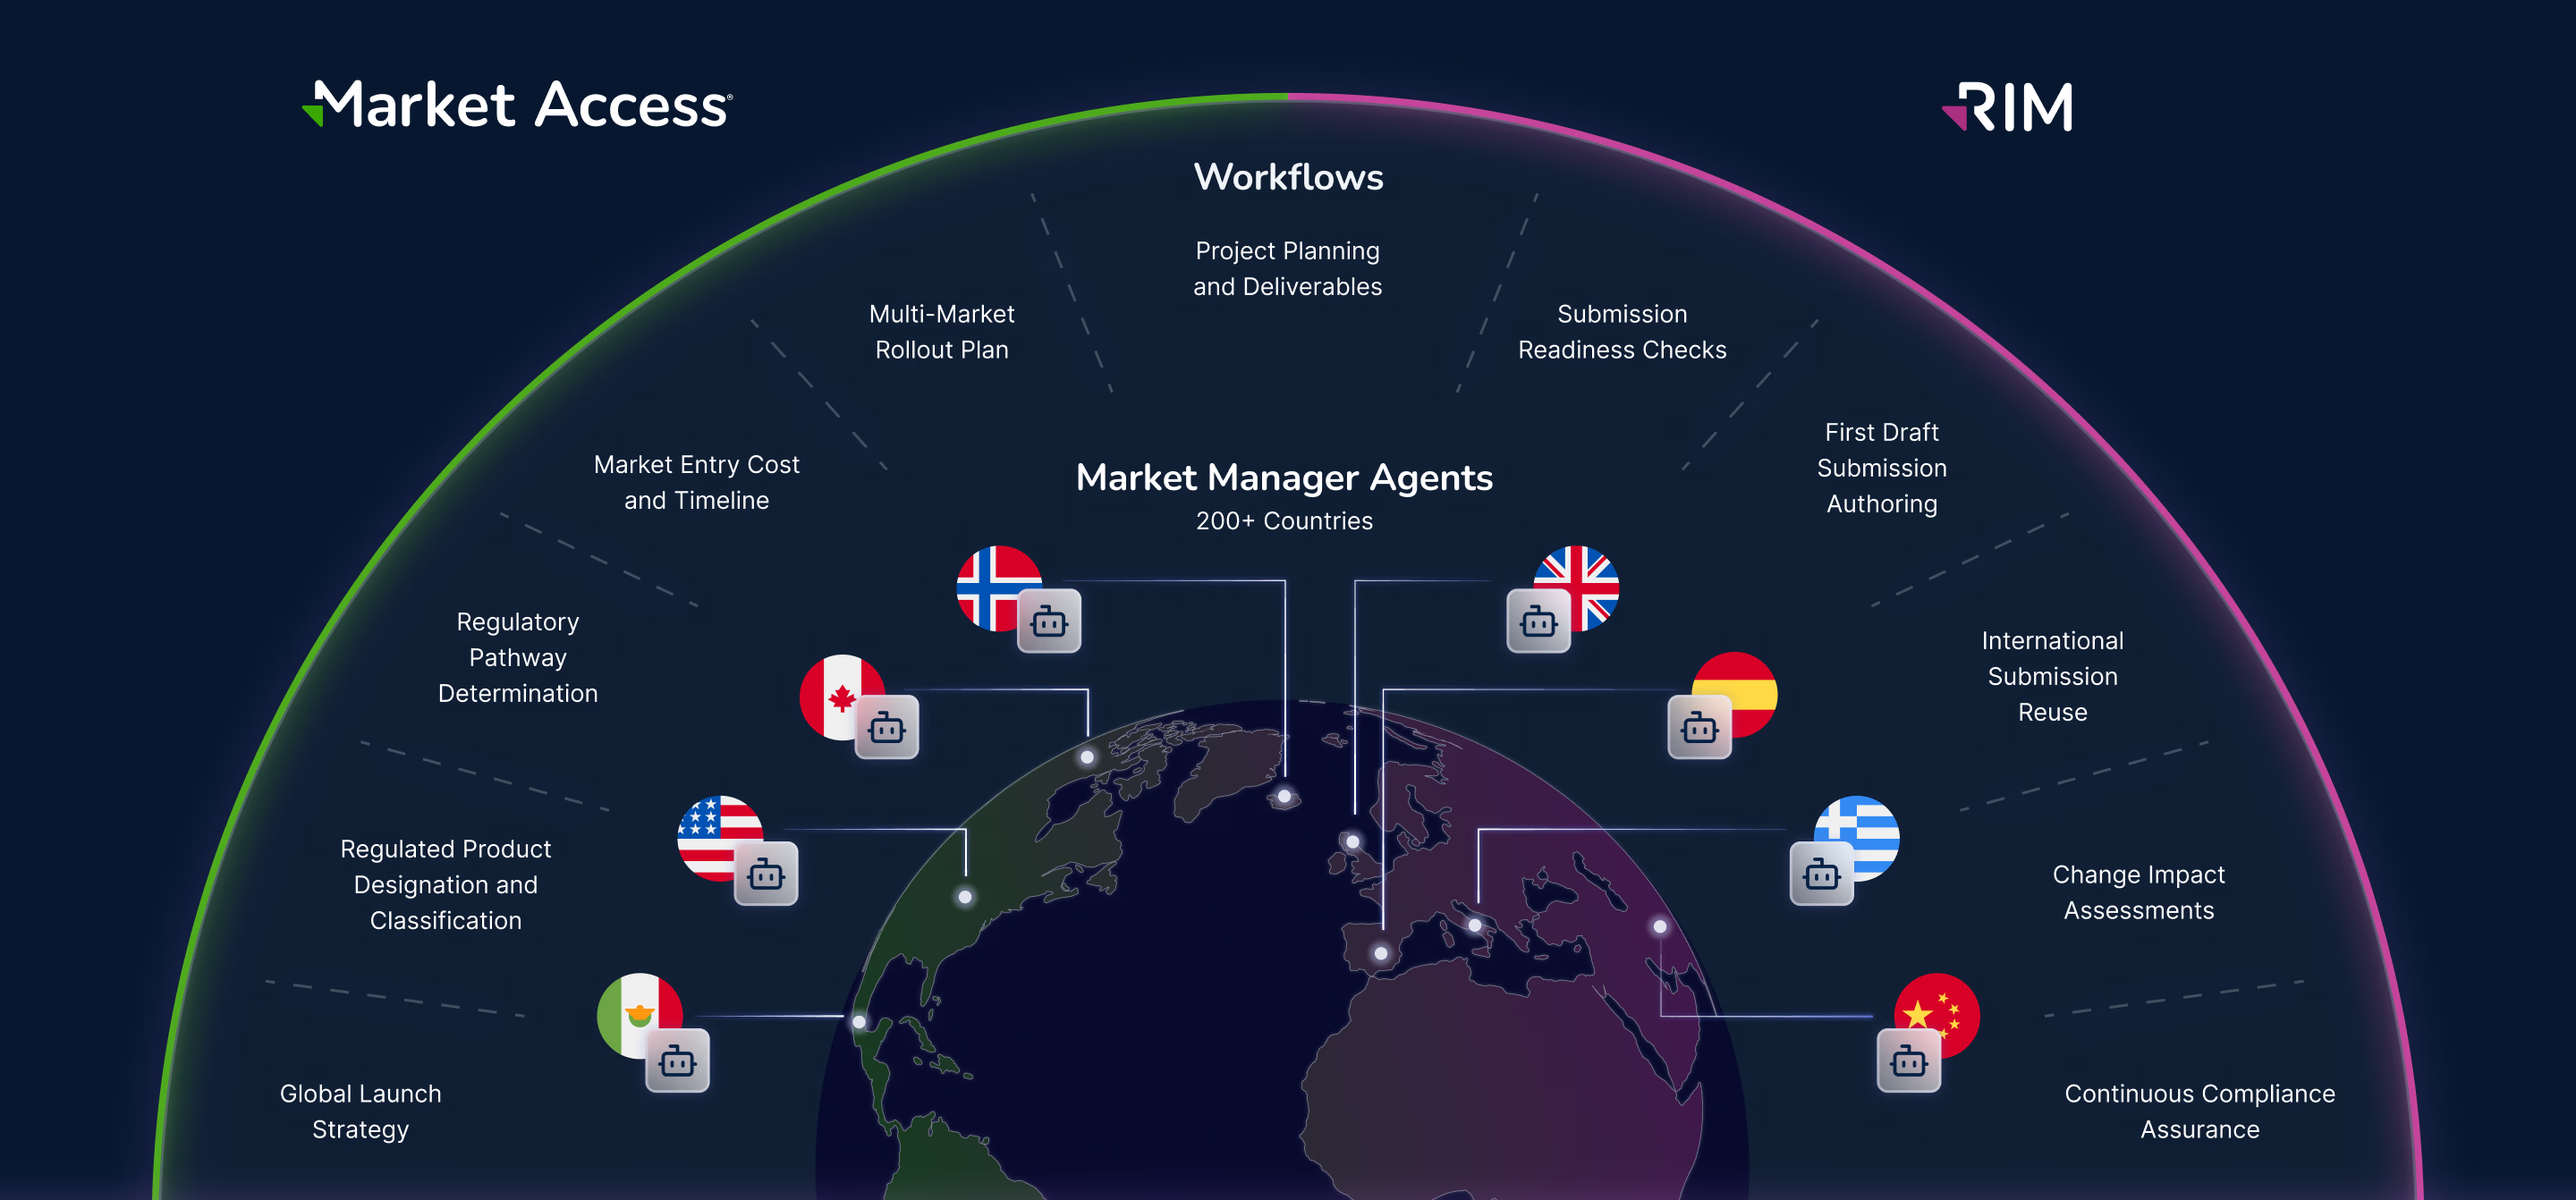
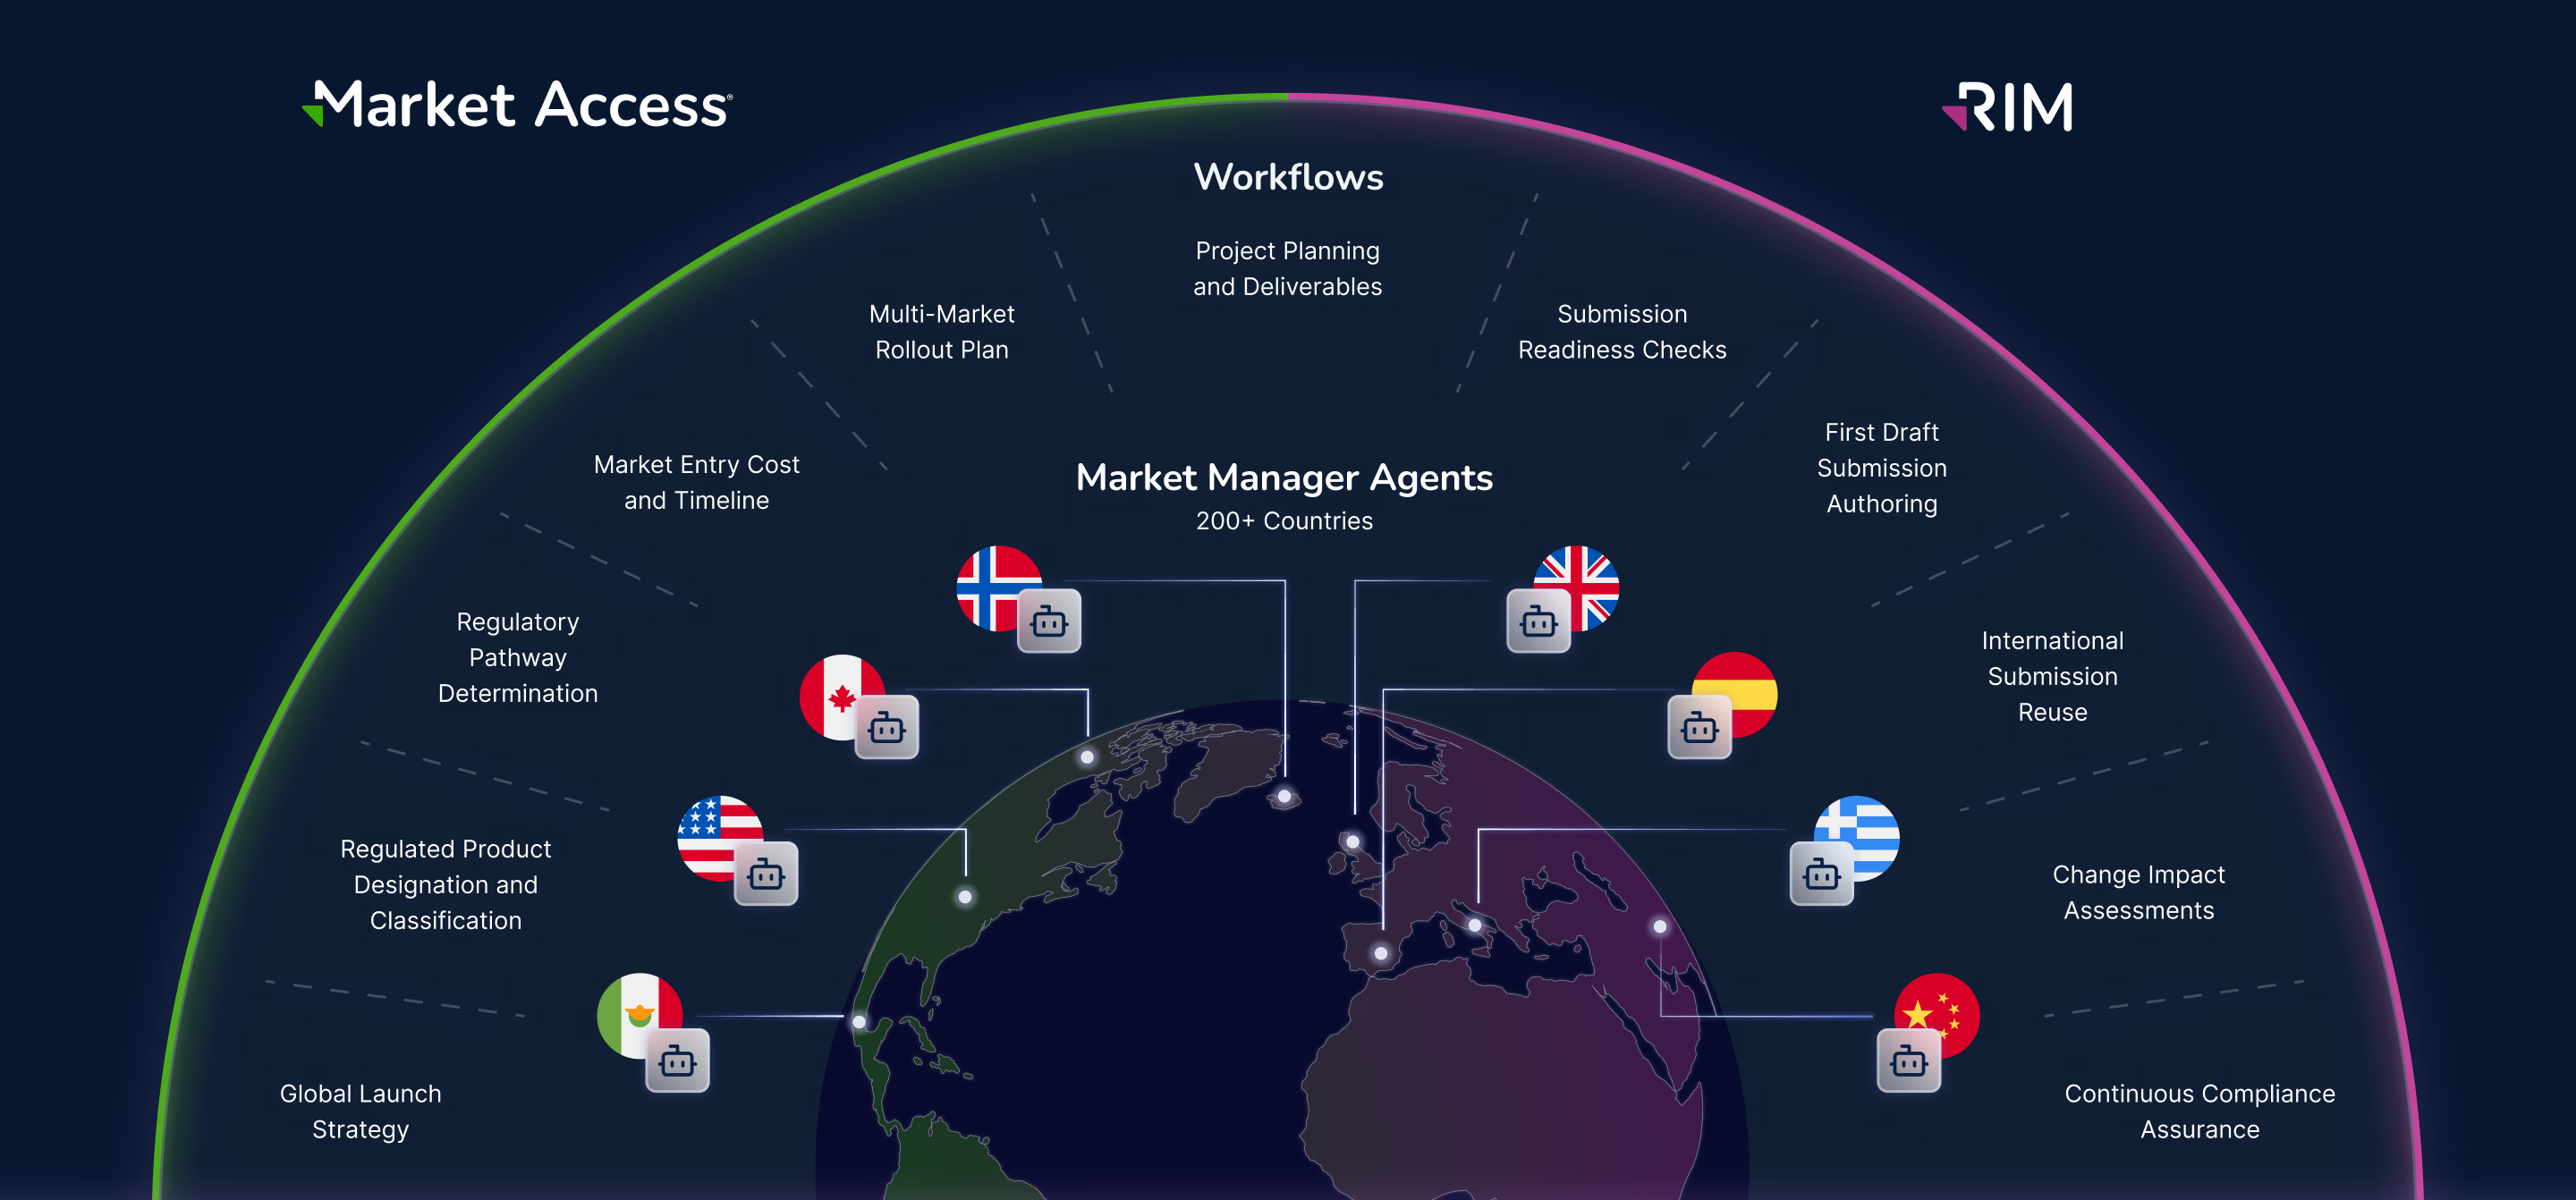
  • AI with human oversight: Uses Rimsys AI and Market Manager Agents to accelerate complex regulatory research while keeping regulatory professionals in control of review and validation.
  • Connected planning and execution: Complements RIM, connecting market-entry decisions with the registrations, submissions, regulatory changes, and ongoing compliance required to execute them.

“Regulatory knowledge has traditionally been highly dependent on individual expertise and market-by-market research,” said James Gianoutsos, Founder and CEO of Rimsys. “Market Access gives companies greater predictability before they make a market investment. Teams can understand whether an opportunity is worth pursuing, what it will take to enter, where regulatory challenges are likely to emerge, and what uncertainty remains. And because every answer is tied to its regulatory source, experts can verify the recommendation rather than simply trust the AI.”

Market Access is powered by Rimsys AI, which works from a curated regulatory corpus spanning more than 200 markets, not the open web. Its Market Manager Agents are AI agents with market-specific regulatory knowledge, including pathways, requirements, and documentation, that support market assessments and global launch planning. The product is also designed to acknowledge areas where reviewer interpretation and expert judgment can affect regulatory outcomes rather than presenting those outcomes as more predictable than they are.

Market Access expands the Rimsys product portfolio from regulatory execution into market planning. Market Access helps regulatory and commercial leaders determine where to compete, while RIM manages the registrations, submissions, regulatory changes, product data, and ongoing compliance required to enter and remain in those markets. Together, they turn regulatory intelligence into coordinated action, helping regulatory teams accomplish more in every hour while giving business leaders greater visibility into opportunities, risk, and the work required to protect and grow revenue.

Beta Availability

Market Access will be available in beta beginning September 1, 2026. During the beta period, Rimsys will continue validating product capabilities and refining workflows ahead of broader commercial availability. 

Organizations interested in learning more about Market Access or participating in the beta program can visit rimsys.io/products/market-access.

About Rimsys

Rimsys is the heart of regulatory operations, connecting AI-native market planning with AI-embedded regulatory execution, giving medtech manufacturers one living system from market-entry decision to execution. 

Rimsys Market Access is the planning layer, answering not just whether a market can be entered, but whether it should be, returning classification, pathway, timeline, and cost before a team commits budget or resources to finding out. Rimsys RIM is the execution layer, managing registrations, submissions, regulatory changes, and compliance so teams can act on that plan with confidence.

Trusted by 6 of the top 12 global MedTech manufacturers and certified to ISO 27001 and SOC 2, with ISO 42001 certification for AI management systems underway, Rimsys keeps regulatory intelligence, product data, approvals, and change management continuously connected so teams can enter new markets with speed and assurance. Learn more at rimsys.io.

Media Contact
letschat@rimsys.io

Registration deadlines don't have to be a risk. See how centralized data, automated alerts, and standardized workflows keep global registrations on track

MedTech

RIM

Product Updates

How to Prevent Registration Deadline Misses in 2026

By

Bethaney Lentz

August 4, 2026

4 min read

A missed registration deadline can halt sales, trigger regulatory penalties, and damage your organization's credibility with health authorities. For MedTech regulatory affairs teams managing hundreds or thousands of product registrations across dozens of markets, preventing deadline misses requires more than calendar reminders. It demands structured data, clear workflows, and real-time visibility into expiration timelines.

This guide explains why registration compliance deadlines are missed and how your team can prevent delays through stronger data control, workflow visibility, and process standardization.

Key Takeaways: How to Prevent Registration Deadline Misses in 2026

  • Registration deadline misses typically stem from fragmented data, unclear ownership, and siloed communication between teams.
  • Proactive renewal management requires real-time visibility into expiration dates across your entire global product portfolio.
  • Standardized workflows with assigned accountability eliminate the confusion that leads to overlooked deadlines.
  • Rimsys centralizes registration data and automates renewal alerts, reducing the risk of missed deadlines by up to 90%.
  • Investing in a purpose-built RIM platform protects revenue continuity and maintains uninterrupted market access.

Why Do MedTech Teams Miss Registration Compliance Deadlines?

Registration deadline misses  occur because the operating model cannot scale with growing portfolio complexity. Understanding root causes is the first step toward prevention.

‍

Fragmented Data Across Systems and Regions

Many organizations track registrations using disconnected regional trackers, shared drives, and individual team member records. When product data lives in multiple locations, no single source of truth exists. This fragmentation makes it difficult to identify upcoming expirations before they become urgent.

A registration that expires in Brazil may be tracked in one system while the same product's EU approval is monitored elsewhere. Without connection between these records, teams lack portfolio-wide visibility into renewal timelines.

‍

Unclear Ownership and Accountability

When responsibilities are distributed across regional leads, in-country partners, and distributors without clear accountability structures, deadlines fall through the cracks. Team members may assume someone else is handling a renewal, leading to inaction until the deadline passes.

Organizations with high turnover face additional risk. When key personnel leave, institutional knowledge about pending renewals often leaves with them.

‍

Reactive Tracking Instead of Proactive Management

Calendar-based reminders and email notifications are inherently reactive. By the time a reminder appears, teams may not have enough runway to gather required documentation, coordinate with authorities, or address unexpected complications.

Effective deadline management requires proactive monitoring that surfaces upcoming expirations months in advance, not days before they occur.

What Happens When Registration Deadlines Are Missed?

The consequences of missed registration deadlines extend beyond administrative inconvenience. They create tangible business disruptions that affect revenue, reputation, and regulatory standing.

‍

Loss of Market Access and Revenue Disruption

An expired registration means your product cannot legally be sold in that market. Regulatory teams must halt shipments, distribution partners cannot fulfill orders, and revenue stops flowing. For high-volume products or critical markets, even brief interruptions to the supply chain can translate to significant financial impact.

Re-registering a lapsed product often takes longer than a standard renewal. Your team may face additional scrutiny, updated documentation requirements, or queue delays that extend the time to market restoration.

‍

Regulatory Penalties and Increased Scrutiny

Health authorities track compliance history. Missed deadlines signal operational weaknesses that can trigger increased inspection frequency, additional documentation requests, or enhanced scrutiny on future submissions. Building trust with regulators takes years; eroding it takes one avoidable miss.

‍

Damage to Distributor and Partner Relationships

In-country partners and distributors depend on your organization to maintain valid registrations. When market access lapses, partners bear the burden of explaining delays to healthcare providers and end customers. Repeated misses can damage long-term business relationships and competitive positioning.

How to Identify Products at Risk of Deadline Misses

Prevention starts with identifying which registrations are most vulnerable. A systematic risk assessment helps teams prioritize attention and allocate resources effectively.

‍

Create a Consolidated Registration Inventory

Bring all registration data into a single view. Include product identifiers, registration numbers, approval dates, expiration dates, responsible parties, and market-specific requirements. This inventory becomes your foundation for risk assessment.

Rimsys Registration Software centralizes this data automatically, linking registrations to products, submissions, and regulatory intelligence in one structured system.

‍

Segment by Expiration Timeline and Complexity

Not all renewals carry equal risk. Categorize registrations by time remaining until expiration and by the complexity of renewal requirements. Products expiring within the next 90 days with extensive documentation needs should receive immediate attention.

Consider country-specific factors. Some markets require local testing, updated clinical data, or notified body involvement. These requirements extend lead times and increase the risk of delays.

‍

Assess Documentation Readiness

For each at-risk registration, evaluate whether required documentation is current and accessible. Outdated technical files, missing test reports, or incomplete labeling information create bottlenecks that delay renewal submissions.

Documentation gaps discovered weeks before a deadline often cannot be addressed in time. Early assessment ensures enough runway to resolve issues.

Building a Proactive Renewal Management System

Moving from reactive tracking to proactive management requires intentional process design. The following components form the foundation of an effective renewal management system.

‍

Establish Clear Ownership for Every Registration

Assign a single accountable owner for each registration. This person is responsible for monitoring expiration timelines, coordinating renewal activities, and escalating issues that require additional support.

Ownership should be documented within your registration management system, not stored in separate organizational charts or email threads. When someone leaves the organization, ownership transfer becomes straightforward.

‍

Define Standard Lead Times by Market and Product Type

Different markets have different renewal timelines. EU MDR renewals involve notified body coordination. FDA establishment registrations follow annual cycles. ANVISA submissions may require specific local documentation.

Document the standard lead time needed for each market and product type. Use these lead times to trigger renewal workflows well before deadlines approach.

‍

Implement Automated Expiration Monitoring

Replace manual calendar tracking with automated monitoring that surfaces upcoming expirations across your entire portfolio. Alerts should trigger at multiple intervals, such as 180 days, 90 days, 60 days, and 30 days before expiration.

Automated monitoring ensures that no registration is overlooked, regardless of portfolio size or team bandwidth. Rimsys Global Regulatory Compliance capabilities include lifecycle tracking and automated alerts that reduce missed renewal risk.

Standardizing Workflows to Prevent Oversight

Consistent workflows eliminate the variability that leads to deadlines. When every renewal follows the same structured process, teams can identify and address deviations before they cause problems.

‍

Create Renewal Workflow Templates

Define the steps required for each type of renewal. Include documentation gathering, internal review, submission preparation, authority submission, and post-submission tracking. Assign responsible parties and expected durations for each step.

Templates ensure that new team members can execute renewals correctly without relying on undocumented institutional knowledge.

‍

Build Escalation Paths for Delayed Activities

Not every renewal proceeds according to plan. Documentation may be delayed. Authority responses may take longer than expected. Partners may fail to submit required local information.

Define clear escalation paths that activate when activities fall behind schedule. Escalation should be automatic and visible, not dependent on individual team members raising concerns manually.

‍

Conduct Regular Pipeline Reviews

Schedule recurring reviews of your renewal pipeline. During these reviews, examine registrations approaching expiration, identify any blocked activities, and confirm that responsible parties are on track.

Pipeline reviews surface potential issues early, when corrective action is still possible. They also reinforce accountability by making renewal status visible to leadership.

How Data Control Reduces Deadline Risk

Strong data control is the foundation of reliable deadline management. When registration data is accurate, current, and accessible, teams make better decisions and avoid preventable misses.

‍

Maintain a Single Source of Truth

Eliminate competing data sources by establishing one authoritative system for registration information. All team members, partners, and stakeholders should access the same data, ensuring consistency and reducing confusion.

A single source of truth means that when someone asks about a product's registration status in a specific market, the answer is immediate and reliable. No cross-referencing multiple trackers. No waiting for colleagues to respond to email inquiries.

‍

Connect Registration Data to Product and Submission Records

Registrations do not exist in isolation. They connect to specific products, rely on previous submissions, and may be affected by regulatory changes or product modifications. Systems that link these data types enable more accurate impact analysis.

When a product design change occurs, connected data helps teams identify which registrations may need updated. When regulations evolve, linked intelligence shows which markets and products are affected.

‍

Ensure Version Control and Audit Readiness

Regulatory authorities expect organizations to demonstrate controlled processes. Maintain version history for all registration-related documentation. Track who made changes, when changes occurred, and what was modified.

Audit readiness built into daily operations means less scrambling when inspections occur. It also shows the historical context needed to understand how registration status evolved over time.

Leveraging Technology for Visibility and Automation ‍

Technology plays a critical role in scaling deadline management across large portfolios. Purpose-built regulatory information management platforms offer capabilities that generic tools cannot match.

‍

Dashboard Visibility Across Global Markets

Executive and operational dashboards display at-a-glance visibility into registration status across your entire portfolio. Filter by market, product line, expiration timeline, or responsible party, to focus on relevant subsets.

Dashboard visibility enables leadership to ask and answer questions quickly:

  • How many registrations will expire in the next quarter?
  • Which markets have the highest concentration of upcoming renewals?
  • Are any renewals currently blocked?

‍

Automated Alerts and Notifications

Configure alerts that notify responsible parties when action is required. Notifications should be specific, actionable, and timely. Rather than generic reminders, alerts should identify the specific registration, required action, and deadline.

Effective notification systems reduce the cognitive burden on team members. Instead of tracking deadlines mentally, they receive prompts when attention is needed.

‍

Reporting for Continuous Improvement

Track metrics that indicate process health. Monitor the number of registrations renewed on time, average lead time utilization, and frequency of expedited renewals. Use these metrics to identify process weaknesses and drive improvement.

Organizations that measure renewal performance can demonstrate compliance maturity to regulators, partners, and investors. Metrics also help justify investments in process improvement.

How Rimsys Helps Prevent Registration Deadline Misses

Rimsys is the first and only holistic Regulatory Information Management software purpose-built for MedTech. It centralizes registrations, submissions, regulatory intelligence, and UDI data in one connected platform, giving teams the visibility and automation needed to prevent deadline misses.

‍

Centralized Global Registration Tracking

Rimsys maintains a structured, product-centric data model that connects registrations to products, markets, and submissions. Teams gain real-time visibility into where every product can be sold, which registrations are pending, and which are approaching expiration.

Six of the world's top 12 MedTech manufacturers trust Rimsys to manage global regulatory operations, achieving up to 90% reduction in reporting effort and elimination of manual tracking risk.

‍

Automated Lifecycle Alerts

Rimsys monitors registration lifecycles and automatically alerts teams to upcoming expirations, renewals, and information requests. Automated alerts ensure that no registration is overlooked, regardless of the portfolio size.

With Rimsys AI, teams can accelerate regulatory work through AI-assisted workflows that reduce repetitive tasks while keeping human judgment in control.

‍

Connected Regulatory Intelligence

Registration deadlines do not exist in isolation from regulatory change. New requirements can affect renewal timelines, documentation needs, or market access conditions.

Rimsys connects registration data to regulatory intelligence and impact assessment workflows, helping teams understand how changes affect their portfolio before deadlines become urgent.

‍

Creating a Culture of Deadline Accountability

Technology and process alone do not prevent deadline misses. Organizations must also cultivate a culture where deadline accountability is valued and reinforced.

‍

Make Renewal Performance Visible

Share renewal metrics with teams and leadership regularly. Celebrate on-time renewals and analyze near-misses to identify improvement opportunities. Visibility creates accountability without requiring punitive measures.

‍

Invest in Team Training and Development

Ensure that team members understand the importance of deadline management and have the skills to execute renewal workflows effectively. Training should cover both procedural requirements and the business impact of missed deadlines.

‍

Support Cross-Functional Collaboration

Registrations often require input from quality, engineering, labeling, and commercial teams. Foster collaboration across functions to ensure that dependencies are identified early and addressed proactively.

When regulatory affairs teams operate in silos, they lack the information needed to anticipate complications. Cross-functional visibility enables earlier intervention.

In Conclusion: How to Prevent Registration Deadline Misses

Preventing registration deadline misses requires intentional effort across three dimensions: data control, workflow visibility, and process standardization. Organizations that invest in these areas protect revenue continuity, maintain regulatory standing, and preserve partner relationships.

The complexity of global regulatory operations is not slowing down. As portfolios expand and requirements evolve, teams need infrastructure that scales. Rimsys transforms regulatory operations from administrative tracking into strategic enablement, bringing speed, visibility, and confidence to global expansion.

Ready to eliminate deadline risk from your registration management? Speak with the Rimsys team to see how leading MedTech manufacturers manage global registrations with precision and control.

‍

References:

I agree to the privacy policy including to Rimsys using my contact details to contact me for marketing purposes.
Thank you! Your submission has been received!
Oops! Something went wrong while submitting the form.
Hand holding smartphone showing email app with 12 unread messages notification.