Announcing AI-Native Market Access Product for Regulated Products

Learn More

Blogs

MedTech

ISO 10993: Standards for the biologic evaluation of medical devices

By

Dell Lundy

January 6, 2023

4 min read

The International Organization for Standardization (ISO) is the largest body in the world publishing standards. In fact, it is a conglomeration of standards bodies from over 160 countries working together to harmonize standards. As such, ISO 10993 is the international standard that is practically used globally for testing and determining the biocompatibility of medical devices. So it’s critical for medical device manufacturers to understand all 23 parts of ISO 10993 for the success of 510(k), pre-market authorization (PMA), and other device submission projects for regulatory authorities worldwide. As an example, the FDA has issued guidance on the Use of International Standard ISO 10993-1.

What is biocompatibility?

According to ISO 10993-1:2018, the current version of part 1 of the standard, biocompatibility is the ability of a medical device or material to perform with an appropriate host response in a specific application. Any device that comes into direct or indirect contact with the skin must be tested for biocompatibility. A medical device that makes indirect contact with the skin is one that encounters a liquid, gas, or another medium, that makes direct contact with the patient or user.

Categorizations for medical devices according to ISO 10993

When testing the biocompatibility of a device, it is broken down into two categories; one based on its type of contact with humans, and the other based on the duration of contact.

The categorizations for types of contact are:

  • Non-contacting medical devices: These are medical devices that do not make direct or indirect contact with patients. Examples include in-vitro diagnostics devices, blood collection tubes, and petri dishes.
  • Surface-contacting devices: Surface-contacting medical devices are ones that touch the skin, in-tact mucous membranes, and breached or compromised surfaces. Examples of these devices are catheters, contact lenses, and bronchoscopes.
  • Externally communicating devices: Externally communicating devices are those that are partially or wholly external and come into contact with bodily fluids. These devices are usually intended to deliver or draw fluids to or from the body and are attached to an external delivery or withdrawal system. Examples include dialyzers and dialysis tubing accessories, transfer and transfusion sets, and arthroscopes.
  • Implantable devices: Implantable devices are the riskiest type for medical devices because they are embedded within human tissue. Pacemakers, artificial larynxes, and heart valves are all implantable devices.

The categorizations for times of duration are:

  • Limited exposure – Medical devices whose cumulative sum of single, multiple, or repeated duration of contact is up to 24 hours.
  • Prolonged exposure – Medical devices whose cumulative sum of single, multiple, or repeated contact time is likely to exceed 24 hours but does not exceed 30 days.
  • Long-term exposure – Medical devices whose cumulative sum of single, multiple, or repeated contact time exceeds 30 days.

Determining biocompatibility

Medical devices are most commonly made of metals, plastics, and fabrics, which are composed of chemicals with varying properties. Manufacturers must gather physical and chemical information about the device, which is vital to its biological and material evaluation and characterization.

For devices with components that are made of or utilize novel chemicals or materials, or those known to cause adverse effects, ISO 10993 requires rigorous risk assessment and management according to the standards of ISO 14971. Furthermore, there are prescribed data endpoints that set the foundation for determining the biocompatibility of medical devices and their intended uses and components.

The main things manufacturers must consider when determining the biocompatibility of medical devices and their components are listed below:

  • Complete chemical characterization – ISO 10993 requires manufacturers to describe the chemical and material makeup of the medical device and its components, as well as the use of chemicals in the manufacturing of the device. Sometimes, a test of extractable and leachable chemicals is required to determine the safety of the medical device.
  • Toxicological assessment – Toxicological assessment serves to determine and mitigate the risk of medical devices when they come into contact with patients and users. There are four pillars of toxicology assessment: hazard identification, hazard characterization, exposure assessment, and risk characterization.
  • Biocompatibility testing – Biocompatibility testing is the process of testing the local and systemic effects of a medical device on the tissues it comes into contact with. Oftentimes a favorable toxicological assessment by a qualified individual, based on the facts of the thorough chemical characterization, can rule out the possibility of adverse effects and the need for biocompatibility testing.

ISO 10993 compliance

Biocompatibility assessment is a vital part of risk management according to ISO 14971. Ensuring compliance with risk management and biocompatibility assessment standards requires buy-in from all departments, from marketing and design to quality assurance and regulatory affairs.

It is vital that you begin considering ISO 10993-1:2018 in the early stages of product design. Part 1 of the standard will refer to additional parts, as listed in the following section. Completing your complete chemical characterization and toxicology assessment early in the process will help ensure the biocompatibility of your medical device during the design phase and expedite your device registration and time to market.

Also, it’s important to note that many regulatory authorities around the world have their own variation of ISO 10993. While these varying standards have the same foundation and are similar in many ways, you must understand their nuances if you plan to offer your medical device internationally.

ISO 10993 sections

ISO 10993 is made up of 23 different sections or parts, each of which is maintained and updated separately. Previews of the standard can be viewed on the ISO website, but full versions of the standard need to be purchased.

  • ISO 10993-1:2018 – Evaluation and testing within a risk management system
  • ISO 10993-2:2022 – Animal welfare requirements
  • ISO 10993-3:2014 – Tests for genotoxicity, carcinogenicity, and reproductive toxicity
  • ISO 10993-4:2017 – Selection of tests for interactions with blood
  • ISO 10993-5:2009 – Tests for in vitro cytotoxicity
  • ISO 10993-6:2016 – Tests for local effects after implantation
  • ISO 10993-7:2008 – Ethylene oxide sterilization residuals
  • ISO 10993-8: - Withdrawn (Selection of reference materials for biologic tests)
  • ISO 10993-9:2019 – Framework for identification and quantification of potential degradation products
  • ISO 10993-10:2021 – Tests for skin sensitization
  • ISO 10993-11:2017 – Tests for systemic toxicity
  • ISO 10993-12:2021 – Sample preparation and reference materials
  • ISO 10993-13:2010 – Identification and quantification of degradation products from polymeric medical devices
  • ISO 10993-14:2001 – Identification and quantification of degradation products from ceramics
  • ISO 10993-15:2019 – Identification and quantification of degradation products from metals and alloys
  • ISO 10993-16:2017 – Toxicokinetic study design for degradation products and leachables
  • ISO 10993-17:2002 – Establishment of allowable limits for leachable substances
  • ISO 10993-18:2020 – Chemical characterization of medical device materials within a risk management process
  • ISO 10993-19:2020 – Physico-chemical, morphological, and topographical characterization of materials
  • ISO 10993-20:2006 – Principles and methods for immunotoxicology testing of medical devices
  • ISO 10993-22:2017 – Guidance on nanomaterials
  • ISO 10993-23:2021 – Tests for irritation

How can we help?

Many manufacturers endure longer and more costly paths to market than necessary because they do not have systems and tools designed specifically for their regulatory teams. Furthermore, a lack of visibility and collaboration from departments that see regulatory teams traditionally as the “department of saying no” leaves ample room for human error in regulatory, quality management, and even marketing processes and activities. Read more about why we believe regulatory teams need to be considered revenue functions, not cost centers.

The resulting inefficiencies lead to problems such as marketing products with expired certificates, missing certificates, inaccurate and/or incomplete submissions, and even non-compliance with current regulatory requirements. Having a holistic RIM system is central to staying in compliance with standards, regulations, and guidance in the many markets around the world. Rimsys is the only RIM system of its kind built specifically for the medtech industry.

To learn how Rimsys can help your company get its regulatory ducks in a row, click here to schedule a demo.

‍

Similar posts

AI

From Regulatory Knowledge to Better Market Decisions: A Conversation with Steve Gens and James Gianoutsos

October 1, 2026

4 min read

AI is moving quickly from experimentation to implementation across life sciences. For regulatory teams, that raises a bigger question than where AI can automate a task:

How can organizations use AI to make better decisions while maintaining the traceability, expert oversight, and confidence regulatory work demands?

That was a central theme when James Gianoutsos, Founder and CEO of Rimsys, joined Steve Gens, Founder and Managing Partner of Gens & Associates, on the Regulatory Executive Podcast.

Their conversation explored the changing role of regulatory intelligence, the introduction of Rimsys Market Access, the importance of transparency in regulatory AI, and what these changes could mean for regulatory teams.

Regulatory knowledge has been difficult and costly to scale

For global medtech organizations, regulatory knowledge has traditionally been highly dependent on people.

A company may have specialists responsible for particular countries or regions, along with local representatives, distributors, consultants, and other experts. Those people develop deep knowledge of local regulations, pathways, and expectations.

The problem is that the knowledge often remains distributed across the organization. And maintaining that market-by-market expertise can become expensive as a company expands its portfolio and geographic footprint.

As James described during the podcast, large manufacturers may rely on dozens or even hundreds of internal employees, third-party distributors, local representatives, and other specialists to understand what it takes to bring products into different markets. Smaller organizations may rely more heavily on outside consultants and partners. Either way, the model can be costly and difficult to scale. 

But the cost isn't limited to what companies spend on expertise.

Steve described the workflow as a series of “start, stop” cycles: ask a local expert, verify whether the regulation is current, gather more information, and repeat. Each cycle consumes regulatory capacity and extends the time it takes to give the business a confident answer.

That creates another potentially more consequential expense: the cost of waiting.

When it takes months to understand what entering a market will require, commercial and product teams have less visibility into when revenue might begin, what resources will be needed, or whether the opportunity is worth pursuing. Regulatory teams spend valuable capacity assembling and validating information rather than applying their expertise to more strategic work.

Ultimately, scaling regulatory knowledge isn't simply an efficiency problem. It affects how quickly and confidently the business can decide where to invest.

The bigger question: Should we enter this market?

That challenge is central to Rimsys Market Access.

Most market-entry research begins with questions such as: How is the product classified? What's the regulatory pathway? What are the requirements?

Those questions establish whether a company can enter a market. But they don't necessarily tell the business whether it should.

Market Access is designed to help regulatory and commercial teams develop an earlier view of what market entry could involve, including classification, regulatory pathway, timeline, cost considerations, requirements and potential gaps. 

For commercial and product leaders, that provides greater predictability before committing budget and resources.

For regulatory leaders, it means starting with a structured plan rather than assembling every market assessment from scratch.

Moving regulatory knowledge from individuals into a shared system

During the conversation, James described one of AI's most significant opportunities as moving knowledge that has historically lived in people's heads into software.

The goal isn't to eliminate regulatory expertise. It's to make that expertise more scalable.

Market Access works from a curated regulatory corpus, allowing market-specific knowledge to be applied across workflows without requiring every initial question to go through the individual who happens to know that jurisdiction. Outputs are designed to trace back to the underlying regulation, guidance, or law. 

That can also change where regulatory professionals spend their time. Rather than focusing as heavily on finding information and assembling initial assessments, they can focus on reviewing the evidence, identifying gaps, applying judgment, and advising the business.

Steve described the concept as a subject-matter-expert assistant. James called it a workforce multiplier. 

Trust requires opening the regulatory AI “black box”

Speed isn't enough when the information is being used to inform regulatory decisions.

James and Steve returned repeatedly to the importance of knowing where an AI-generated answer comes from. As Steve summarized it during the discussion, transparency builds trust.

Market Access is designed around that principle. Its outputs are grounded in a curated regulatory corpus and traceable to the specific regulations, guidance, or laws supporting them. That gives regulatory professionals the ability to verify an answer rather than simply accept it. 

Human judgment remains essential as well. Market Access provides an upfront assessment, not a guarantee of a regulatory outcome. Reviewer interpretation and other factors can still affect what happens during the regulatory process.

The objective isn't to make uncertainty disappear. It's to make what is known, what the sources say, and where uncertainty remains much easier to see.

Connecting the market decision to execution

The discussion also pointed to a broader evolution in regulatory technology: moving from systems that primarily store regulatory information toward systems that help teams act on it.

That's where Market Access and Rimsys RIM play distinct but complementary roles.

Market Access is the planning layer. It helps teams determine whether, when, and how to enter a market.

Rimsys RIM is the execution layer. It manages the registrations, submissions, regulatory changes, and ongoing regulatory work required to act on that plan. 

Together, they create a path from the initial market decision through regulatory execution rather than leaving planning and execution disconnected.

A new role for AI in regulatory work

One of the most interesting themes from Steve and James's conversation was that AI's impact may ultimately be less about replacing individual tasks and more about expanding what regulatory teams can accomplish.

Better access to market-specific knowledge can help professionals work across more markets, spend less time gathering information, and devote more capacity to the judgment and strategic work that requires their expertise.

The technology is changing quickly, but the fundamentals of regulatory work remain: reliable information, domain expertise, transparency, and human judgment.

The opportunity is to apply those fundamentals at greater scale while giving the business a clearer answer to the question that comes before execution:

Not simply, “Can we enter this market?”

But, “Should we?”

Listen to the full conversation

Hear the complete discussion between Steve Gens, Founder and Managing Partner of Gens & Associates, and James Gianoutsos, Founder and CEO of Rimsys, on the Regulatory Executive Podcast for more on regulatory AI, market-entry planning, transparency, and the changing role of regulatory teams.

‍

AI

How Market Access Builds Trust in Regulatory Intelligence

By

Bethaney Lentz

September 18, 2026

4 min read

Why source traceability, regulatory specificity, and human judgment matter when AI informs a market-entry decision.

Regulatory professionals are not short on AI options. General-purpose tools can answer questions quickly and often sound convincing. But in medtech, a convincing answer is not the same thing as a decision you can defend.

That distinction matters most before a company commits to a new market. Teams need to understand how a product is classified, which pathway applies, how long entry may take, what it may cost, where the important gaps are, and what uncertainty still remains. Commercial leaders need that visibility to decide whether the opportunity is worth pursuing. Regulatory leaders need to know that the assessment is grounded in sources they can verify.

That is the trust standard behind Rimsys Market Access. It is not simply about producing regulatory information faster. It is about helping medtech teams decide whether, when, and how to enter a market before committing budget and resources, with outputs linked back to the regulations, guidance, and laws behind them.

Why fluency is not enough for regulatory work

A general AI tool can be useful for brainstorming, summarizing, and exploring a topic. The challenge comes when an answer has real regulatory consequences. A plausible response may still be incomplete, based on the wrong designation, or disconnected from the current source that should govern the decision.

Medtech makes this especially difficult because the same product can be treated differently across markets and product designations. Medical device, IVD, cosmetic, consumer, OTC, and combination-product considerations can change the regulatory read. A market-entry assessment has to account for that context rather than assume one answer travels cleanly from one jurisdiction to another.

For regulatory teams, the natural next question is: Where did this answer come from? If the system cannot show the underlying authority, the expert still has to rebuild the research before trusting the result. That limits the value of the speed AI was supposed to create in the first place.

Trust starts with a decision you can verify

Market Access is designed around a different outcome. The goal is not to give teams another stream of regulatory information to interpret. It is to provide a source-linked market plan that helps them answer the bigger business question: should we enter this market, not just can we?

For a market assessment, that can include classification, regulatory pathway, timeline, cost considerations, key requirements and potential gaps. The output is designed to give commercial and product leaders an earlier, more defensible view of the opportunity while giving regulatory professionals the detail they need to review the basis for the assessment.

Trust comes from being able to inspect that basis. Market Access works from a curated regulatory corpus spanning more than 200 markets rather than relying on the open web. Answers are linked to the specific regulation, guidance, or law behind them, so a regulatory professional can verify the source instead of being asked to accept an AI-generated conclusion on faith.

Three things make regulatory intelligence more defensible

1. Source traceability

A regulatory answer is more useful when the person reviewing it can follow the evidence. Market Access links outputs back to their underlying regulatory sources. That changes the review process from “Do I trust the AI?” to a more practical question: “Does the cited source support this conclusion for our product and situation?”

That is an important shift for regulatory teams. Instead of assembling every market assessment from a blank page, they can review a structured, sourced plan, challenge assumptions, and focus their expertise on the places where judgment matters most.

2. Regulatory specificity

Trust also depends on context. Market Access is purpose-built for medtech regulatory complexity and supports cross-designation classification. The objective is to evaluate the product in the context of the market and the applicable regulatory framework, then carry that context into the pathway, requirements, timeline, cost considerations, and gaps that shape the market-entry decision.

This is where a purpose-built market-planning platform differs from a generic chat experience. The value is not an impressive answer in isolation. It is a connected assessment that helps the team understand what entering the market is likely to require and whether the opportunity still makes sense once those requirements are visible.

3. Human judgment stays in control

No system can remove every area of uncertainty from regulatory work. Reviewer interpretation and expert judgment still affect outcomes, and Market Access is not designed to promise approval.

Instead, Market Access is designed to make uncertainty more manageable. Regulatory professionals review and validate the work before it moves into execution. They can verify sources, assess the gaps, and apply company and product context that software cannot fully replace. The technology accelerates the research and planning. The regulatory professional remains responsible for the judgment.

What trust looks like in a market-entry decision

Imagine a team evaluating several markets for a new product. The commercial question is not simply whether registration is technically possible. The team needs to know which markets deserve investment now, which may require more evidence or resources, and which could put the revenue window at risk because the pathway is longer or more complex than expected.

Regulatory has a different, but connected, need. They have to understand the classification and pathway, identify the requirements most likely to slow the submission, and confirm that the plan reflects the relevant regulatory sources.

Market Access brings those views together. Commercial and product leaders get earlier visibility into timing, cost considerations, requirements, and risk so they can build a stronger business case. Regulatory teams get a source-linked plan they can review rather than a collection of unsupported conclusions. Both groups are working from the same market-entry picture before significant resources are committed.

Trust is not the end goal. Better decisions are.

It is easy for a conversation about regulatory AI to become a conversation about models, agents, or architecture. Those details matter, but they are not the business outcome.

The real value of trustworthy regulatory intelligence is what it lets a team do next. It can help a commercial leader defend a market investment. It can help regulatory surface issues earlier. It can give both groups a more predictable view of the work ahead before budget, R&D capacity, or launch timing is locked in.

And once the decision is made, the work does not have to stop at an intelligence report. Market Access is the market-planning layer. Rimsys RIM is the execution layer for registrations, submissions, regulatory changes, and ongoing regulatory work. Each can stand on its own, and together they create a path from the market-entry decision into controlled execution.

A more useful standard for regulatory AI

The question for medtech teams should not be whether AI can produce a regulatory answer. It can. The better question is whether that answer is specific enough, traceable enough, and reviewable enough to support a real decision.

Market Access is built around that standard: a defensible view of whether, when, and how to enter a market, grounded in curated regulatory sources and kept under human review. Because when the decision affects revenue, resources, and regulatory strategy, sounding right is not enough. Teams need to see why the answer is right, where uncertainty remains, and what it means for the decision in front of them.

Company

Product Updates

Rimsys Announces AI-Native Market Access for Regulated Products

By

Bethaney Lentz

September 1, 2026

4 min read

New software gives regulatory and commercial leaders a source-linked, actionable view of market viability, requirements, timing, and risk before committing resources

PITTSBURGH, PA, September 1, 2026 - Rimsys, the heart of medtech regulatory operations, today announced Market Access, a new AI-native product designed to help companies determine whether a market is worth entering and what it will take to get there. Market Access will be available in beta beginning September 1, 2026, with general availability to be announced separately.

Market Access replaces what can be a 6-to-9-month research and assessment process with a same-day, actionable read on market viability. It gives regulatory and commercial leaders the classification, pathway, timeline, cost considerations, and potential regulatory gaps they need to make a more defensible investment decision before committing budget and resources.

Predictability is at the core of Market Access. Rather than simply identifying a regulatory pathway, it helps teams understand what to expect, which requirements or gaps are most likely to slow down a submission, and where uncertainty remains. Every answer traces back to the specific regulation, guidance, or law behind it, allowing regulatory professionals to verify the underlying source rather than simply trust an AI-generated response.

Key features and benefits include:

  • Same-day market assessment: Provides an actionable read on whether a market is worth entering, replacing research and assessment that can take 6 to 9 months.
  • Cross-designation classification: Evaluates products across medical device, IVD, cosmetic, consumer, and OTC designations, so the same product receives the appropriate regulatory read regardless of how a given market classifies it.
  • Clear market-entry requirements: Provides classification, regulatory pathway, timeline, cost considerations, and key requirements before teams commit resources.
  • Earlier visibility into regulatory gaps: Surfaces the specific requirements and gaps most likely to slow down a submission and helps teams understand their potential severity.
  • Greater predictability: Provides visibility into anticipated regulatory response windows, risks, and areas of uncertainty so teams can better plan for what lies ahead.
  • Source-linked intelligence: Traces answers to the underlying regulations, guidance, and laws so regulatory professionals can verify the basis for an assessment.
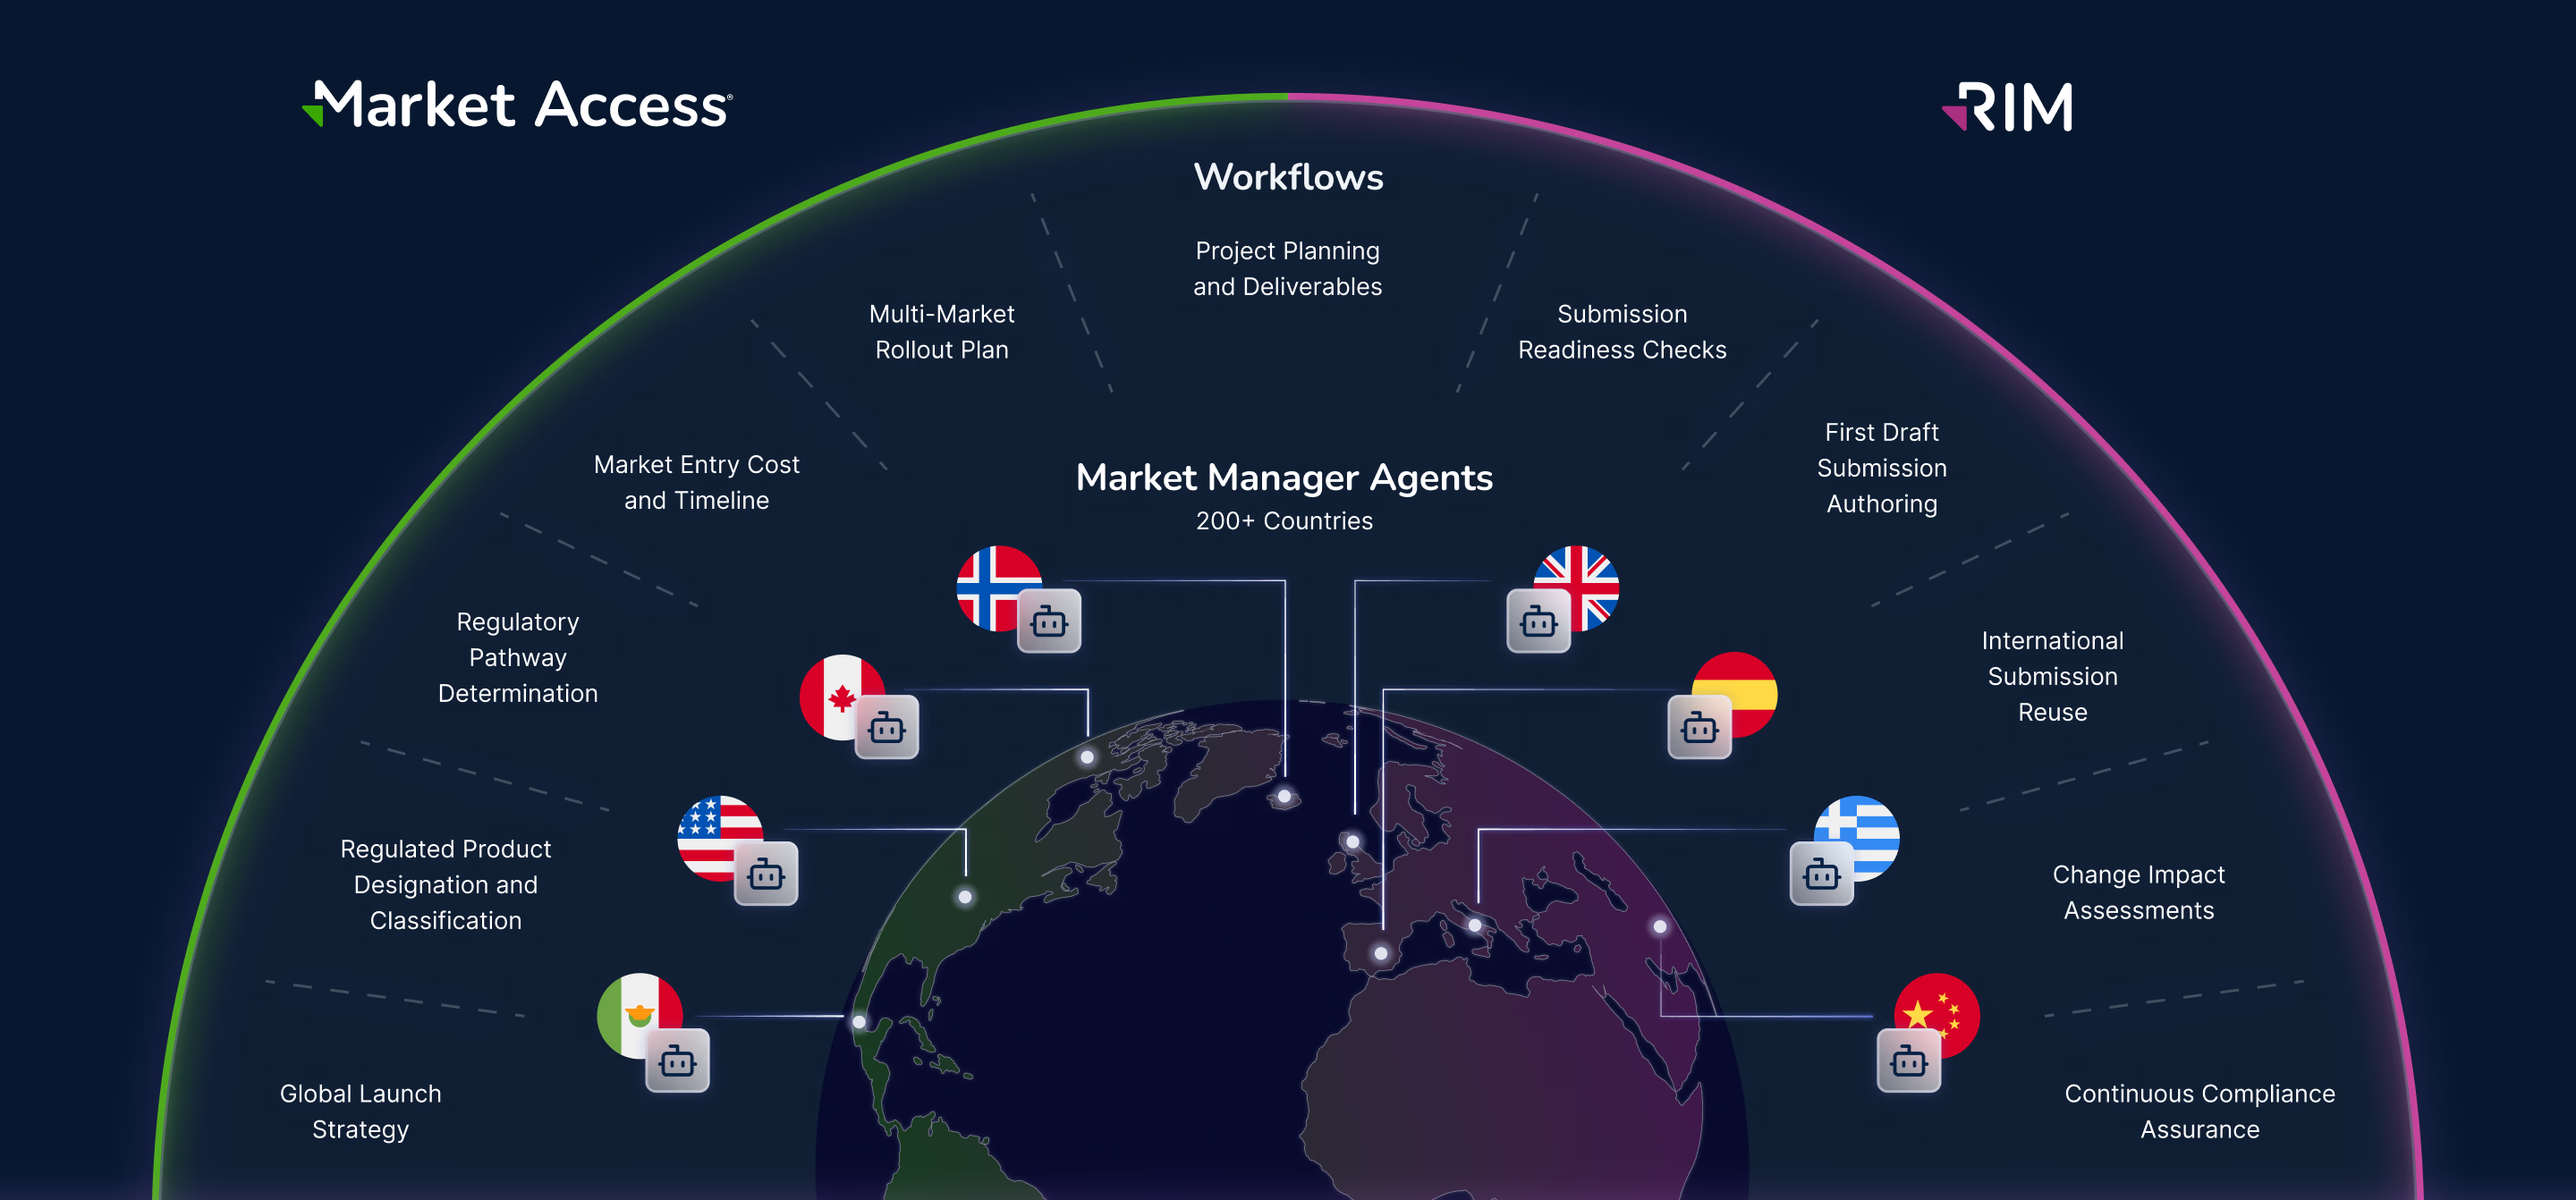
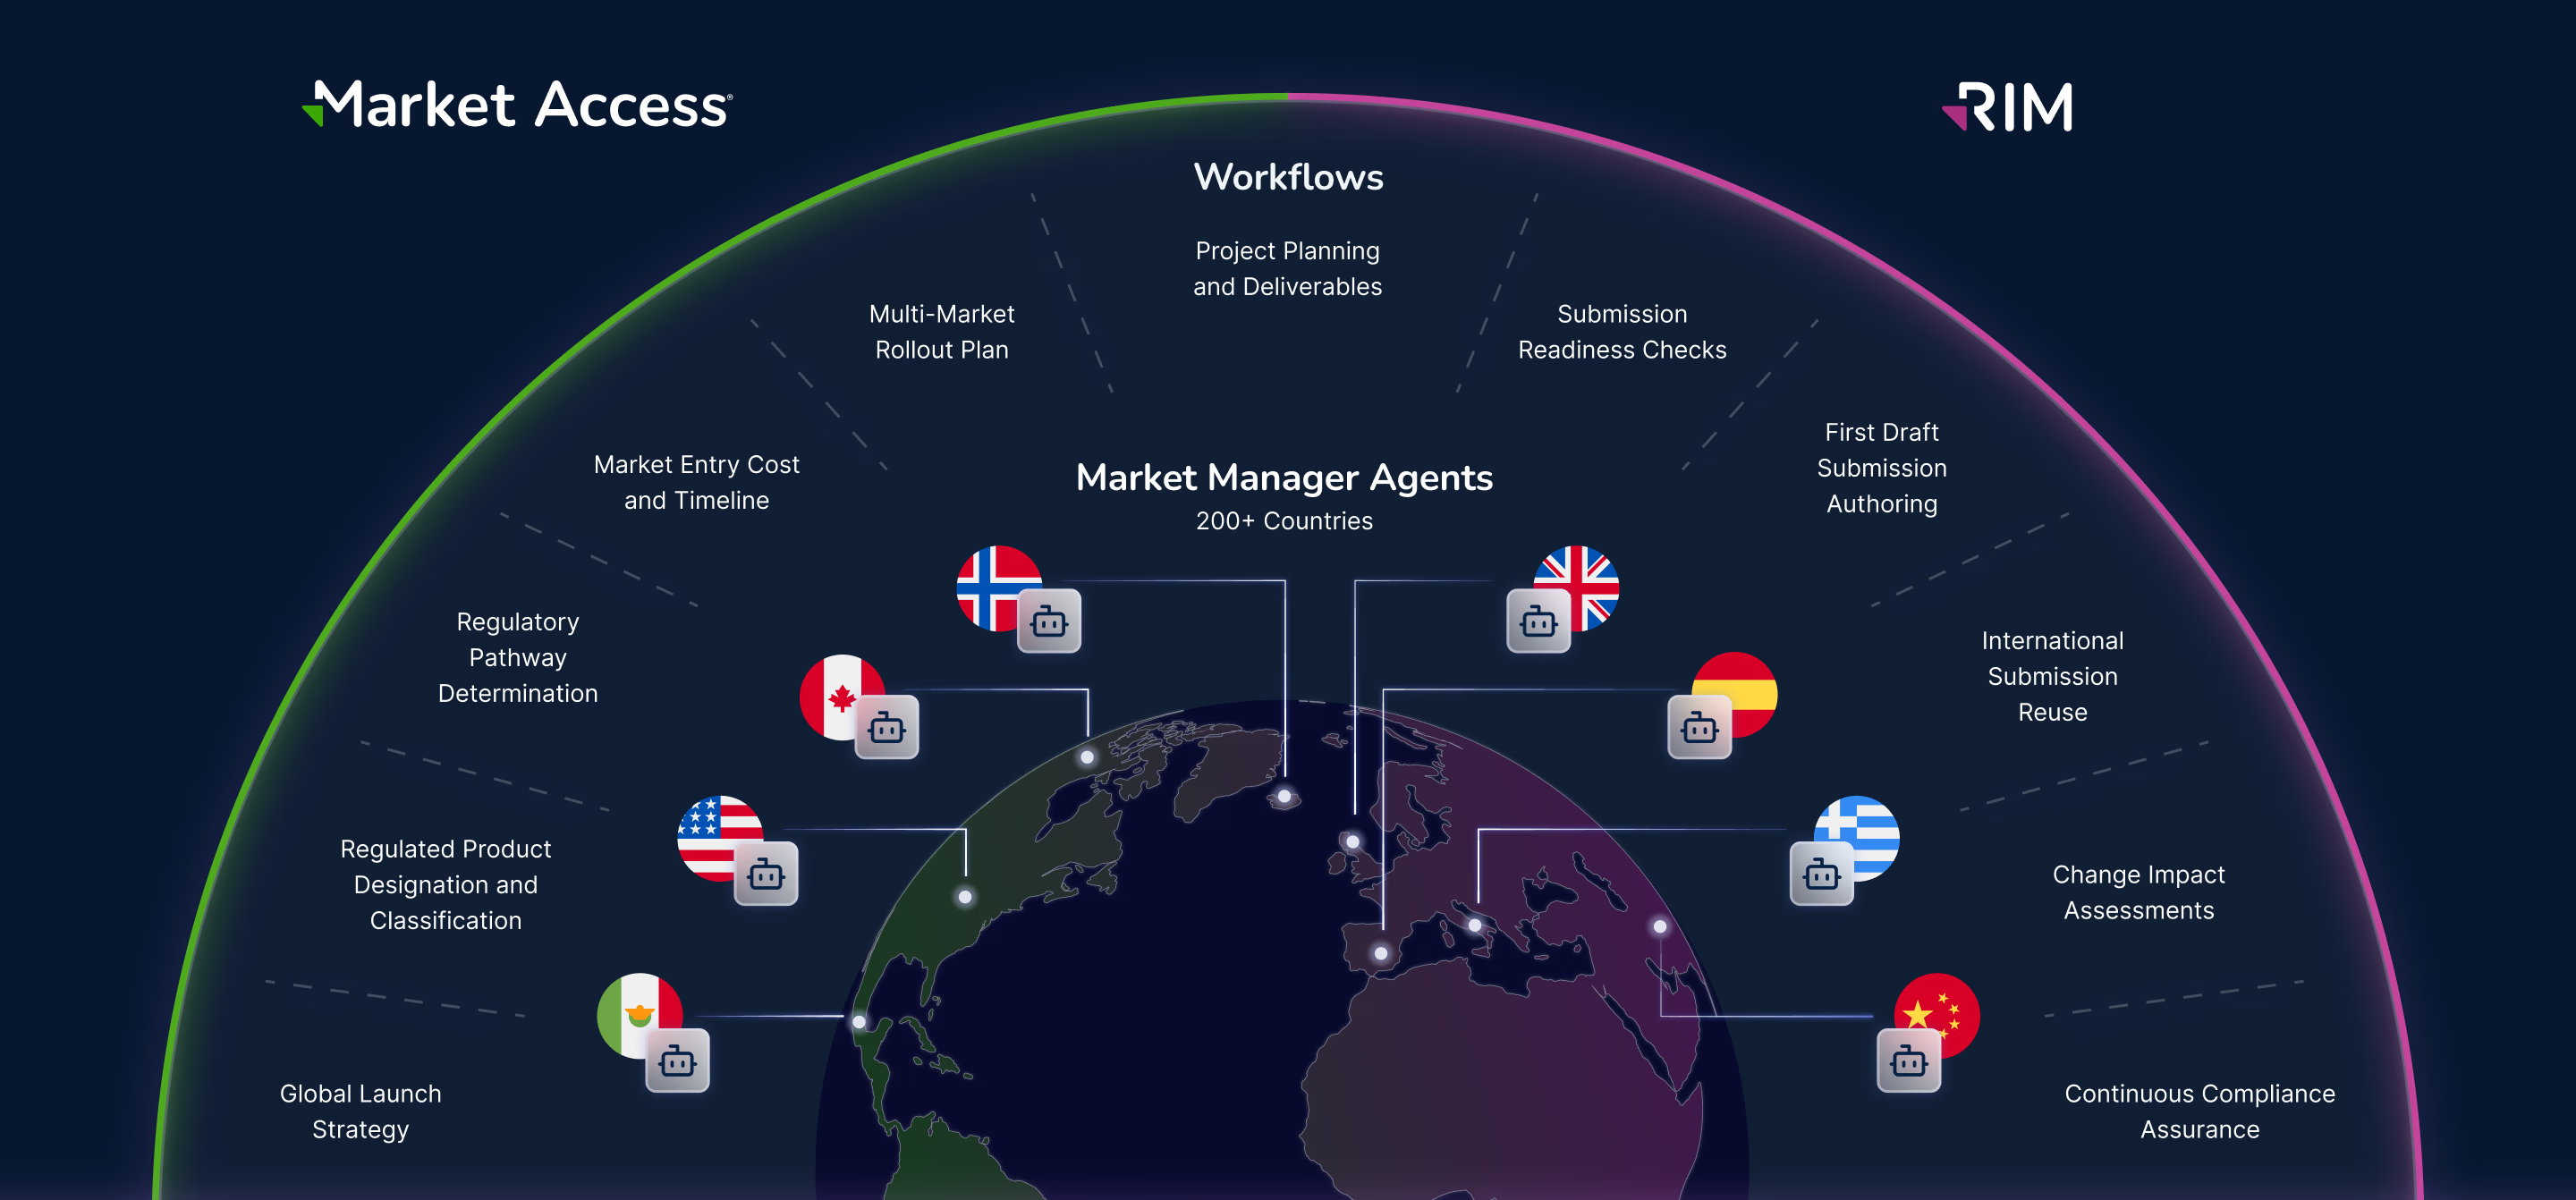
  • AI with human oversight: Uses Rimsys AI and Market Manager Agents to accelerate complex regulatory research while keeping regulatory professionals in control of review and validation.
  • Connected planning and execution: Complements RIM, connecting market-entry decisions with the registrations, submissions, regulatory changes, and ongoing compliance required to execute them.

“Regulatory knowledge has traditionally been highly dependent on individual expertise and market-by-market research,” said James Gianoutsos, Founder and CEO of Rimsys. “Market Access gives companies greater predictability before they make a market investment. Teams can understand whether an opportunity is worth pursuing, what it will take to enter, where regulatory challenges are likely to emerge, and what uncertainty remains. And because every answer is tied to its regulatory source, experts can verify the recommendation rather than simply trust the AI.”

Market Access is powered by Rimsys AI, which works from a curated regulatory corpus spanning more than 200 markets, not the open web. Its Market Manager Agents are AI agents with market-specific regulatory knowledge, including pathways, requirements, and documentation, that support market assessments and global launch planning. The product is also designed to acknowledge areas where reviewer interpretation and expert judgment can affect regulatory outcomes rather than presenting those outcomes as more predictable than they are.

Market Access expands the Rimsys product portfolio from regulatory execution into market planning. Market Access helps regulatory and commercial leaders determine where to compete, while RIM manages the registrations, submissions, regulatory changes, product data, and ongoing compliance required to enter and remain in those markets. Together, they turn regulatory intelligence into coordinated action, helping regulatory teams accomplish more in every hour while giving business leaders greater visibility into opportunities, risk, and the work required to protect and grow revenue.

Beta Availability

Market Access will be available in beta beginning September 1, 2026. During the beta period, Rimsys will continue validating product capabilities and refining workflows ahead of broader commercial availability. 

Organizations interested in learning more about Market Access or participating in the beta program can visit rimsys.io/products/market-access.

About Rimsys

Rimsys is the heart of regulatory operations, connecting AI-native market planning with AI-embedded regulatory execution, giving medtech manufacturers one living system from market-entry decision to execution. 

Rimsys Market Access is the planning layer, answering not just whether a market can be entered, but whether it should be, returning classification, pathway, timeline, and cost before a team commits budget or resources to finding out. Rimsys RIM is the execution layer, managing registrations, submissions, regulatory changes, and compliance so teams can act on that plan with confidence.

Trusted by 6 of the top 12 global MedTech manufacturers and certified to ISO 27001 and SOC 2, with ISO 42001 certification for AI management systems underway, Rimsys keeps regulatory intelligence, product data, approvals, and change management continuously connected so teams can enter new markets with speed and assurance. Learn more at rimsys.io.

Media Contact
letschat@rimsys.io

I agree to the privacy policy including to Rimsys using my contact details to contact me for marketing purposes.
Thank you! Your submission has been received!
Oops! Something went wrong while submitting the form.
Hand holding smartphone showing email app with 12 unread messages notification.