Announcing AI-Native Market Access Product for Regulated Products

Learn More

Blogs

MedTech

Software as a medical device (SAMD) - classification overview

By

Wendy Levine

February 7, 2022

4 min read

What is software as a medical device (SaMD)?

As the pace of technological innovation continues to increase, the definition of what constitutes a medical device also continues to evolve as countries update regulations.  In 2013, the International Medical Device Regulators Forum (IMDRF) created the Software as a Medical Device working group. Currently chaired by the U.S. FDA, the working group is chartered with developing guidance that encourages innovation while assuring safe and effective products. While SaMD is regulated differently in different countries, this article will focus on the many similarities, and some differences, between the FDA regulations in the U.S. and the MDR regulations in the EU.

In general, medical device software falls into 3 different categories:

  • Software as a Medical Device (SaMD): The IMDRF defines SaMD as “software intended to be used for one or more medical purposes that perform these purposes without being part of a hardware medical device.” We list specific examples below, but typically the software classified as SaMD isdesigned to run on generally available hardware, such as Windows computers or mobile devices, or online in the “cloud”. While they may be utilizing data from another medical device, SaMD performs its function independently of any medical equipment or hardware.  
  • Software in a Medical Device: Sometimes referred to as SiMD, Software in a medical device cannot operate separately from its device, or perform its primary function without the device. For example, the software used to program and run an MRI machine would be useless without the MRI machine. Software in this category is regulated together, and as part of, the whole device.
  • Software as an Accessory to a Medical Device: Similarly to Software in a Medical Device, software in this category cannot fulfill a medical purpose on their own. In some cases, a manufacturer may be able to bundle or embed the software in the medical device (making it SiMD) and/or also sell the software separately, making it an accessory.

Similarly, in Europe, the European Commission’s Medical Device Group (MDCG) defines Medical Device Software (MDSW) as a having its own intended purpose. Software which controls a medical device or is otherwise part of a medical device and does not serve a separate medical purpose does not qualify as MDSW, but is regulated by the MDR.

Software in the SaMD category is both a medical device AND software - with relevant regulatory and quality considerations that are specific and unique to each category, yet which must work in tandem. For example, software development best-practices, referenced by the IMDRF SaMD working group, call for iterative feedback loops allowing for quick turnaround of feature requests and bug fixes. While the ability to provide the latest technology and features to the market is an important advantage with SaMD, it does not supersede applicable medical device regulations governing patient safety and efficacy.  

Examples of software as a medical device (SaMD)

Because SaMD software, by definition, is capable of running independently of any specific medical device or hardware, there is a relatively clear line between Software as a Medical Device and other medical software:

Software as Medical Device (SaMD) Not SaMD
Software that evaluates MRI images and makes diagnosis recommendations. Software that controls an MRI machine (SiMD).
A mobile app that monitors a patient’s heart rate or glucose levels and makes treatment recommendations to the patient and/or patient’s doctor. Software that allows a patient to control their insulin pump based on glucose readings (SiMD).
An application, based on machine learning algorithms, that reviews patient health data and makes treatment recommendations. Software that securely stores patient health and treatment history (Digital Health Records or Medical Information System).

SaMD regulatory overview and framework

Why does SaMD have independent regulatory considerations?

Over the past decade, the IMDRF, FDA, and other regulatory bodies have worked to better align regulations with the quickly evolving capabilities and nature of digital devices. Software-only devices (SaMD) generate unique opportunities and considerations:

Because SaMD software typically runs on publicly available hardware:

  • SaMD software must not only be designed to work on specific platforms (usually multiple), but must also be tested and updated frequently as new hardware and operating systems become available.  
  • Agile software development methodologies can provide an environment in which fast product feedback loops are supported within the required regulatory framework.

Because SaMD software can be made readily available to the general public, or specific patient groups, using their own devices:

  • SaMD software can generate faster user/patient feedback - both for medical professionals and for the device manufacturer. Given this environment, regulatory bodies generally want to enable the market to safely take advantage of new innovations as quickly as possible.
  • Product testing needs to take into account the unique and varied environments in which the software may be used, potentially in environments the product is not intended for. The software developer cannot control updates to operating systems or internet browsers, other software that may be running on a user’s device, or the ability for the user to potentially share software or data with others.
  • The advantages of quickly delivering product fixes and updates need to be measured against potentially introducing new, possibly misunderstood or unwanted features, to all users at once.  

SaMD and MDSW risk-based categories

As with all medical devices, the FDA and European MDR classify SaMD and MDSW, respectively, based on the potential impact to patient or public health. The SaMD categorization framework from the IMDRF is an effort “to introduce a foundational approach, harmonized vocabulary, and general and specific considerations, for manufacturers, regulators, and users alike to address the unique challenges associated with the use of SaMD...” This framework provides guidance only for today’s medical software developers, as well as regulatory bodies, such as the FDA. The chart below lists these risk categories by the state of the healthcare situation or condition and by the significance of the information provided by the SAMD:

SaMD categories

State of healthcare situation or condition Treat or diagnose Drive clinical management Inform clinical management
Critical IV III II
Serious III II I
Non-serious II I I

‍‍SaMD Category I:  

SaMD Category I software can provide information for both Serious and Non-Serious health conditions or diseases. Software dealing with serious conditions can be classified in Category I only if it is providing information to inform, not drive, clinical management and is of low impact. Otherwise, Category I SaMD software provides information related to non-serious diseases or health conditions.

Example: Software that collects exercise-related data, such as heart rate, number of steps, and distance walked. If the information is stored and/or transmitted for use by a qualified professional the software is considered to be in Category I, as long as the information is not used by the software to make treatment recommendations directly to the patient or healthcare provider.‍

SaMD Category II:‍

SaMD software in Category II may be used to provide information relevant to a non-serious, serious, or critical healthcare condition or disease, depending on the significance of the information it provides to the healthcare decision. Software that is used to treat or diagnose a health condition is only classified in Category II for non-serious health conditions. Software that provides information regarding serious or critical health conditions or diseases will be classified in Category II only if it provides information to drive or inform, respectively, clinical management decisions (as opposed to providing treatment or diagnosis recommendations directly).‍

Example: SaMD that monitors a diabetic patient’s carbohydrate intake and blood glucose level to calculate recommended insulin dose.‍

SaMD Category III:

SaMD software that provides information to treat or diagnose serious conditions - or which drives clinical management for a critical condition - is classified in Category III.‍

Example: SaMD that analyzes individual data from at-risk populations for a specific type of cancer, and is used to develop preventative intervention strategies. ‍

SaMD Category IV:

SaMD Category IV software provides information used to treat or diagnose a critical health condition and is considered to be very high impact.‍

Example: SaMD that evaluates images of skin lesions in order to determine malignancy.  ‍

For SaMD intended to be used in multiple healthcare situations, the software will be categorized at the highest category according to the SaMD definition statement. ‍

SaMD within the FDA and MDR regulatory framework

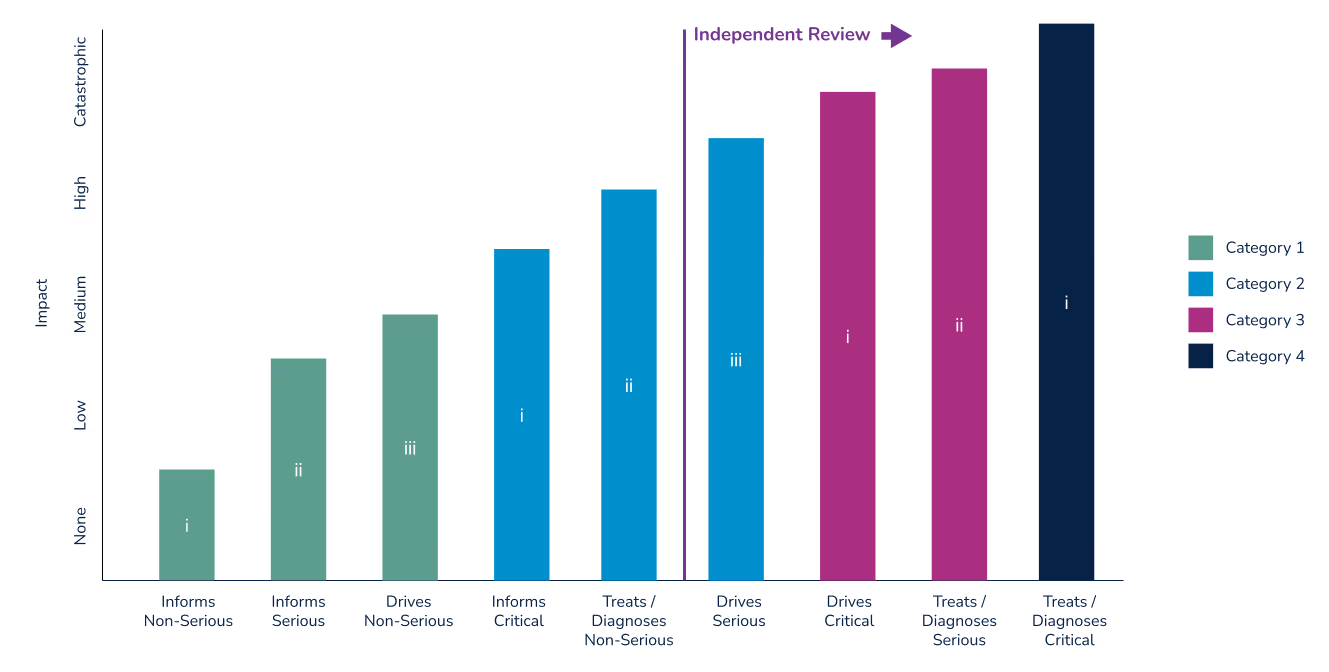
The graph below relates SaMD categories with FDA medical device classes, and further identifies where an independent review is recommended by the FDA. For additional information see Software as a Medical Device (SaMD): Clinical Evaluation.  Guidance for Industry and Food and Drug Administration Staff.‍

Note that FDA regulatory classifications more closely align with the IMDF’s SaMD categories than do those defined by under the European MDR. In the U.S., a device is classified by identifying similar, predicate devices. The simpler 510(k) pre-market submission can be used if a manufacturer can show that their device is substantially equivalent to a Class I or II device. With few exceptions, devices that are Class III are subject to the more rigorous Pre-Market Authorization (PMA) process.  Devices where no substantial equivalent can be shown are subject to to the PMA or De Novo regulatory submissions processes.  The De Novo process was implemented in 2010 to provide a pathway for novel devices with lower risk profiles.

Medical Device Software is classified by the European Commission’s Medical Device Coordination Group (MDCG) into Class I, II, or III. For devices falling under the IVDR, the MDR defines four classes using letters; A, B, C, and D.

Intervention type Treat or diagnose Drive clinical management Informs clinical management
Critical III IIb IIa
Serious IIb IIa IIa
Non-serious IIa IIa IIa

In the EU, a rules-based framework is used to classify devices. MDSW classification is relatively complex, using a series of 22 “waterfalling” rules with yes/no questions that lead to a final classification of each device. While we won’t go into detail in this article on each of the rules, Rule 11 does deserve discussion. (You can read the Guidance on Classification of Software for MDR and IVDR here).

The MDR’s “rule 11” regarding MDSW classification was released in 2017. This rule  states, in part, that “Software intended to provide information which is used to take decisions with diagnosis or therapeutic purposes is classified as class IIa.” The rule then provides 2 exceptions that increase the MDSW classification in cases where the decisions could cause “a serious deterioration of a person’s state of health or a surgical intervention” (class IIb), or could cause “death or an irreversible deterioration of a person’s state of health” (class III).

As a result of this rule, very few MDSW products can be classified as Class I, meaning that the majority of MDSW is subject to conformity assessments by a Notified Body. Medical software manufacturers marketing their product in Europe should therefore not rely on previous product classifications as a guide.

The future of regulatory approval for software as a medical device (SaMD)

The FDA recently released a new draft guidance document addressing the Content of Premarket Submissions for Device Software Functions. This is a major update of the 2005 guidance on pre-market submissions for software in medical devices and is also long overdue, given the pace of change in the software industry (the FDA originally committed to delivering this update in their fiscal year 2019 as part of the MDUFA IV agreement). This guidance sets out requirements for all pre-market submissions, including 510(k), DeNovo, and PMA submissions.

The recent draft document provides additional guidance around the documentation required for premarket submissions. It defines an “enhanced” level of documentation required for software that is a Class III device, combination product, Blood Establishment Computer software, or when the failure of the software would present probable risk of death or serious injury.  

Staying on top of medical device classifications

Classifying a medical device and gaining regulatory clearance can be a complex process, especially if the device contains software or other emerging technology where some regulatory bodies may be further along than others in developing applicable regulations. For additional information about some of the common pathways to market for new products in the U.S., read our Beginner’s Guide to the FDA 510(k) and Beginner’s Guide to the FDA De Novo Classification Process.

Interested in learning how Rimsys can automate your submission process? Request a custom demo of our RIM platform.
Similar posts

AI

From Regulatory Knowledge to Better Market Decisions: A Conversation with Steve Gens and James Gianoutsos

October 1, 2026

4 min read

AI is moving quickly from experimentation to implementation across life sciences. For regulatory teams, that raises a bigger question than where AI can automate a task:

How can organizations use AI to make better decisions while maintaining the traceability, expert oversight, and confidence regulatory work demands?

That was a central theme when James Gianoutsos, Founder and CEO of Rimsys, joined Steve Gens, Founder and Managing Partner of Gens & Associates, on the Regulatory Executive Podcast.

Their conversation explored the changing role of regulatory intelligence, the introduction of Rimsys Market Access, the importance of transparency in regulatory AI, and what these changes could mean for regulatory teams.

Regulatory knowledge has been difficult and costly to scale

For global medtech organizations, regulatory knowledge has traditionally been highly dependent on people.

A company may have specialists responsible for particular countries or regions, along with local representatives, distributors, consultants, and other experts. Those people develop deep knowledge of local regulations, pathways, and expectations.

The problem is that the knowledge often remains distributed across the organization. And maintaining that market-by-market expertise can become expensive as a company expands its portfolio and geographic footprint.

As James described during the podcast, large manufacturers may rely on dozens or even hundreds of internal employees, third-party distributors, local representatives, and other specialists to understand what it takes to bring products into different markets. Smaller organizations may rely more heavily on outside consultants and partners. Either way, the model can be costly and difficult to scale. 

But the cost isn't limited to what companies spend on expertise.

Steve described the workflow as a series of “start, stop” cycles: ask a local expert, verify whether the regulation is current, gather more information, and repeat. Each cycle consumes regulatory capacity and extends the time it takes to give the business a confident answer.

That creates another potentially more consequential expense: the cost of waiting.

When it takes months to understand what entering a market will require, commercial and product teams have less visibility into when revenue might begin, what resources will be needed, or whether the opportunity is worth pursuing. Regulatory teams spend valuable capacity assembling and validating information rather than applying their expertise to more strategic work.

Ultimately, scaling regulatory knowledge isn't simply an efficiency problem. It affects how quickly and confidently the business can decide where to invest.

The bigger question: Should we enter this market?

That challenge is central to Rimsys Market Access.

Most market-entry research begins with questions such as: How is the product classified? What's the regulatory pathway? What are the requirements?

Those questions establish whether a company can enter a market. But they don't necessarily tell the business whether it should.

Market Access is designed to help regulatory and commercial teams develop an earlier view of what market entry could involve, including classification, regulatory pathway, timeline, cost considerations, requirements and potential gaps. 

For commercial and product leaders, that provides greater predictability before committing budget and resources.

For regulatory leaders, it means starting with a structured plan rather than assembling every market assessment from scratch.

Moving regulatory knowledge from individuals into a shared system

During the conversation, James described one of AI's most significant opportunities as moving knowledge that has historically lived in people's heads into software.

The goal isn't to eliminate regulatory expertise. It's to make that expertise more scalable.

Market Access works from a curated regulatory corpus, allowing market-specific knowledge to be applied across workflows without requiring every initial question to go through the individual who happens to know that jurisdiction. Outputs are designed to trace back to the underlying regulation, guidance, or law. 

That can also change where regulatory professionals spend their time. Rather than focusing as heavily on finding information and assembling initial assessments, they can focus on reviewing the evidence, identifying gaps, applying judgment, and advising the business.

Steve described the concept as a subject-matter-expert assistant. James called it a workforce multiplier. 

Trust requires opening the regulatory AI “black box”

Speed isn't enough when the information is being used to inform regulatory decisions.

James and Steve returned repeatedly to the importance of knowing where an AI-generated answer comes from. As Steve summarized it during the discussion, transparency builds trust.

Market Access is designed around that principle. Its outputs are grounded in a curated regulatory corpus and traceable to the specific regulations, guidance, or laws supporting them. That gives regulatory professionals the ability to verify an answer rather than simply accept it. 

Human judgment remains essential as well. Market Access provides an upfront assessment, not a guarantee of a regulatory outcome. Reviewer interpretation and other factors can still affect what happens during the regulatory process.

The objective isn't to make uncertainty disappear. It's to make what is known, what the sources say, and where uncertainty remains much easier to see.

Connecting the market decision to execution

The discussion also pointed to a broader evolution in regulatory technology: moving from systems that primarily store regulatory information toward systems that help teams act on it.

That's where Market Access and Rimsys RIM play distinct but complementary roles.

Market Access is the planning layer. It helps teams determine whether, when, and how to enter a market.

Rimsys RIM is the execution layer. It manages the registrations, submissions, regulatory changes, and ongoing regulatory work required to act on that plan. 

Together, they create a path from the initial market decision through regulatory execution rather than leaving planning and execution disconnected.

A new role for AI in regulatory work

One of the most interesting themes from Steve and James's conversation was that AI's impact may ultimately be less about replacing individual tasks and more about expanding what regulatory teams can accomplish.

Better access to market-specific knowledge can help professionals work across more markets, spend less time gathering information, and devote more capacity to the judgment and strategic work that requires their expertise.

The technology is changing quickly, but the fundamentals of regulatory work remain: reliable information, domain expertise, transparency, and human judgment.

The opportunity is to apply those fundamentals at greater scale while giving the business a clearer answer to the question that comes before execution:

Not simply, “Can we enter this market?”

But, “Should we?”

Listen to the full conversation

Hear the complete discussion between Steve Gens, Founder and Managing Partner of Gens & Associates, and James Gianoutsos, Founder and CEO of Rimsys, on the Regulatory Executive Podcast for more on regulatory AI, market-entry planning, transparency, and the changing role of regulatory teams.

‍

AI

How Market Access Builds Trust in Regulatory Intelligence

By

Bethaney Lentz

September 18, 2026

4 min read

Why source traceability, regulatory specificity, and human judgment matter when AI informs a market-entry decision.

Regulatory professionals are not short on AI options. General-purpose tools can answer questions quickly and often sound convincing. But in medtech, a convincing answer is not the same thing as a decision you can defend.

That distinction matters most before a company commits to a new market. Teams need to understand how a product is classified, which pathway applies, how long entry may take, what it may cost, where the important gaps are, and what uncertainty still remains. Commercial leaders need that visibility to decide whether the opportunity is worth pursuing. Regulatory leaders need to know that the assessment is grounded in sources they can verify.

That is the trust standard behind Rimsys Market Access. It is not simply about producing regulatory information faster. It is about helping medtech teams decide whether, when, and how to enter a market before committing budget and resources, with outputs linked back to the regulations, guidance, and laws behind them.

Why fluency is not enough for regulatory work

A general AI tool can be useful for brainstorming, summarizing, and exploring a topic. The challenge comes when an answer has real regulatory consequences. A plausible response may still be incomplete, based on the wrong designation, or disconnected from the current source that should govern the decision.

Medtech makes this especially difficult because the same product can be treated differently across markets and product designations. Medical device, IVD, cosmetic, consumer, OTC, and combination-product considerations can change the regulatory read. A market-entry assessment has to account for that context rather than assume one answer travels cleanly from one jurisdiction to another.

For regulatory teams, the natural next question is: Where did this answer come from? If the system cannot show the underlying authority, the expert still has to rebuild the research before trusting the result. That limits the value of the speed AI was supposed to create in the first place.

Trust starts with a decision you can verify

Market Access is designed around a different outcome. The goal is not to give teams another stream of regulatory information to interpret. It is to provide a source-linked market plan that helps them answer the bigger business question: should we enter this market, not just can we?

For a market assessment, that can include classification, regulatory pathway, timeline, cost considerations, key requirements and potential gaps. The output is designed to give commercial and product leaders an earlier, more defensible view of the opportunity while giving regulatory professionals the detail they need to review the basis for the assessment.

Trust comes from being able to inspect that basis. Market Access works from a curated regulatory corpus spanning more than 200 markets rather than relying on the open web. Answers are linked to the specific regulation, guidance, or law behind them, so a regulatory professional can verify the source instead of being asked to accept an AI-generated conclusion on faith.

Three things make regulatory intelligence more defensible

1. Source traceability

A regulatory answer is more useful when the person reviewing it can follow the evidence. Market Access links outputs back to their underlying regulatory sources. That changes the review process from “Do I trust the AI?” to a more practical question: “Does the cited source support this conclusion for our product and situation?”

That is an important shift for regulatory teams. Instead of assembling every market assessment from a blank page, they can review a structured, sourced plan, challenge assumptions, and focus their expertise on the places where judgment matters most.

2. Regulatory specificity

Trust also depends on context. Market Access is purpose-built for medtech regulatory complexity and supports cross-designation classification. The objective is to evaluate the product in the context of the market and the applicable regulatory framework, then carry that context into the pathway, requirements, timeline, cost considerations, and gaps that shape the market-entry decision.

This is where a purpose-built market-planning platform differs from a generic chat experience. The value is not an impressive answer in isolation. It is a connected assessment that helps the team understand what entering the market is likely to require and whether the opportunity still makes sense once those requirements are visible.

3. Human judgment stays in control

No system can remove every area of uncertainty from regulatory work. Reviewer interpretation and expert judgment still affect outcomes, and Market Access is not designed to promise approval.

Instead, Market Access is designed to make uncertainty more manageable. Regulatory professionals review and validate the work before it moves into execution. They can verify sources, assess the gaps, and apply company and product context that software cannot fully replace. The technology accelerates the research and planning. The regulatory professional remains responsible for the judgment.

What trust looks like in a market-entry decision

Imagine a team evaluating several markets for a new product. The commercial question is not simply whether registration is technically possible. The team needs to know which markets deserve investment now, which may require more evidence or resources, and which could put the revenue window at risk because the pathway is longer or more complex than expected.

Regulatory has a different, but connected, need. They have to understand the classification and pathway, identify the requirements most likely to slow the submission, and confirm that the plan reflects the relevant regulatory sources.

Market Access brings those views together. Commercial and product leaders get earlier visibility into timing, cost considerations, requirements, and risk so they can build a stronger business case. Regulatory teams get a source-linked plan they can review rather than a collection of unsupported conclusions. Both groups are working from the same market-entry picture before significant resources are committed.

Trust is not the end goal. Better decisions are.

It is easy for a conversation about regulatory AI to become a conversation about models, agents, or architecture. Those details matter, but they are not the business outcome.

The real value of trustworthy regulatory intelligence is what it lets a team do next. It can help a commercial leader defend a market investment. It can help regulatory surface issues earlier. It can give both groups a more predictable view of the work ahead before budget, R&D capacity, or launch timing is locked in.

And once the decision is made, the work does not have to stop at an intelligence report. Market Access is the market-planning layer. Rimsys RIM is the execution layer for registrations, submissions, regulatory changes, and ongoing regulatory work. Each can stand on its own, and together they create a path from the market-entry decision into controlled execution.

A more useful standard for regulatory AI

The question for medtech teams should not be whether AI can produce a regulatory answer. It can. The better question is whether that answer is specific enough, traceable enough, and reviewable enough to support a real decision.

Market Access is built around that standard: a defensible view of whether, when, and how to enter a market, grounded in curated regulatory sources and kept under human review. Because when the decision affects revenue, resources, and regulatory strategy, sounding right is not enough. Teams need to see why the answer is right, where uncertainty remains, and what it means for the decision in front of them.

Company

Product Updates

Rimsys Announces AI-Native Market Access for Regulated Products

By

Bethaney Lentz

September 1, 2026

4 min read

New software gives regulatory and commercial leaders a source-linked, actionable view of market viability, requirements, timing, and risk before committing resources

PITTSBURGH, PA, September 1, 2026 - Rimsys, the heart of medtech regulatory operations, today announced Market Access, a new AI-native product designed to help companies determine whether a market is worth entering and what it will take to get there. Market Access will be available in beta beginning September 1, 2026, with general availability to be announced separately.

Market Access replaces what can be a 6-to-9-month research and assessment process with a same-day, actionable read on market viability. It gives regulatory and commercial leaders the classification, pathway, timeline, cost considerations, and potential regulatory gaps they need to make a more defensible investment decision before committing budget and resources.

Predictability is at the core of Market Access. Rather than simply identifying a regulatory pathway, it helps teams understand what to expect, which requirements or gaps are most likely to slow down a submission, and where uncertainty remains. Every answer traces back to the specific regulation, guidance, or law behind it, allowing regulatory professionals to verify the underlying source rather than simply trust an AI-generated response.

Key features and benefits include:

  • Same-day market assessment: Provides an actionable read on whether a market is worth entering, replacing research and assessment that can take 6 to 9 months.
  • Cross-designation classification: Evaluates products across medical device, IVD, cosmetic, consumer, and OTC designations, so the same product receives the appropriate regulatory read regardless of how a given market classifies it.
  • Clear market-entry requirements: Provides classification, regulatory pathway, timeline, cost considerations, and key requirements before teams commit resources.
  • Earlier visibility into regulatory gaps: Surfaces the specific requirements and gaps most likely to slow down a submission and helps teams understand their potential severity.
  • Greater predictability: Provides visibility into anticipated regulatory response windows, risks, and areas of uncertainty so teams can better plan for what lies ahead.
  • Source-linked intelligence: Traces answers to the underlying regulations, guidance, and laws so regulatory professionals can verify the basis for an assessment.
  • AI with human oversight: Uses Rimsys AI and Market Manager Agents to accelerate complex regulatory research while keeping regulatory professionals in control of review and validation.
  • Connected planning and execution: Complements RIM, connecting market-entry decisions with the registrations, submissions, regulatory changes, and ongoing compliance required to execute them.

“Regulatory knowledge has traditionally been highly dependent on individual expertise and market-by-market research,” said James Gianoutsos, Founder and CEO of Rimsys. “Market Access gives companies greater predictability before they make a market investment. Teams can understand whether an opportunity is worth pursuing, what it will take to enter, where regulatory challenges are likely to emerge, and what uncertainty remains. And because every answer is tied to its regulatory source, experts can verify the recommendation rather than simply trust the AI.”

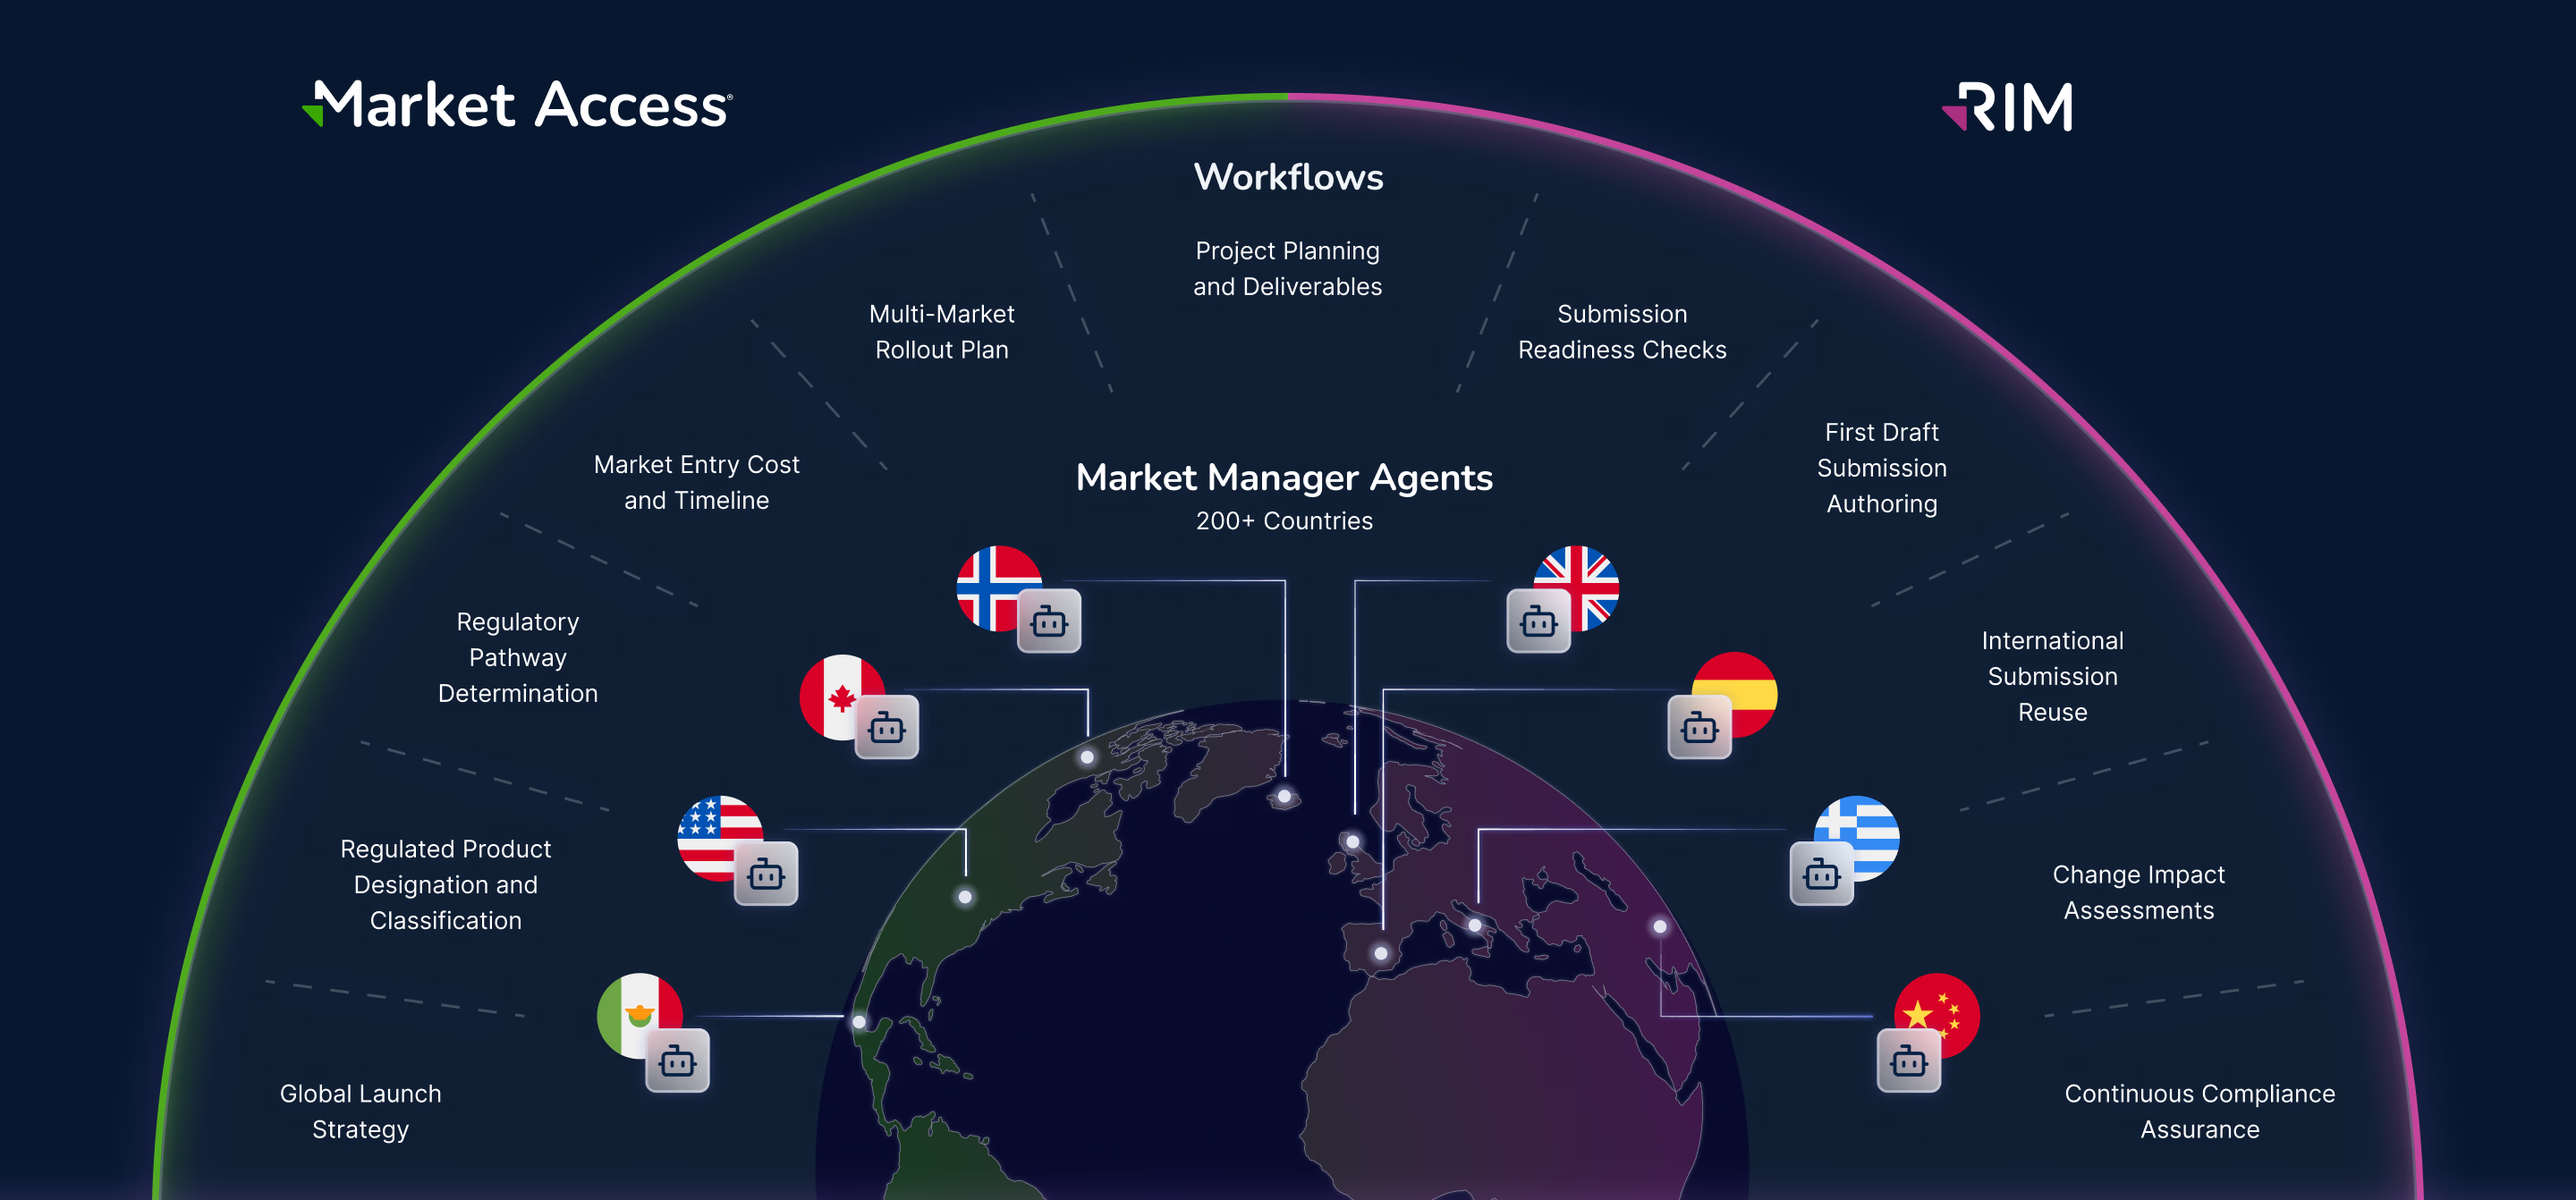
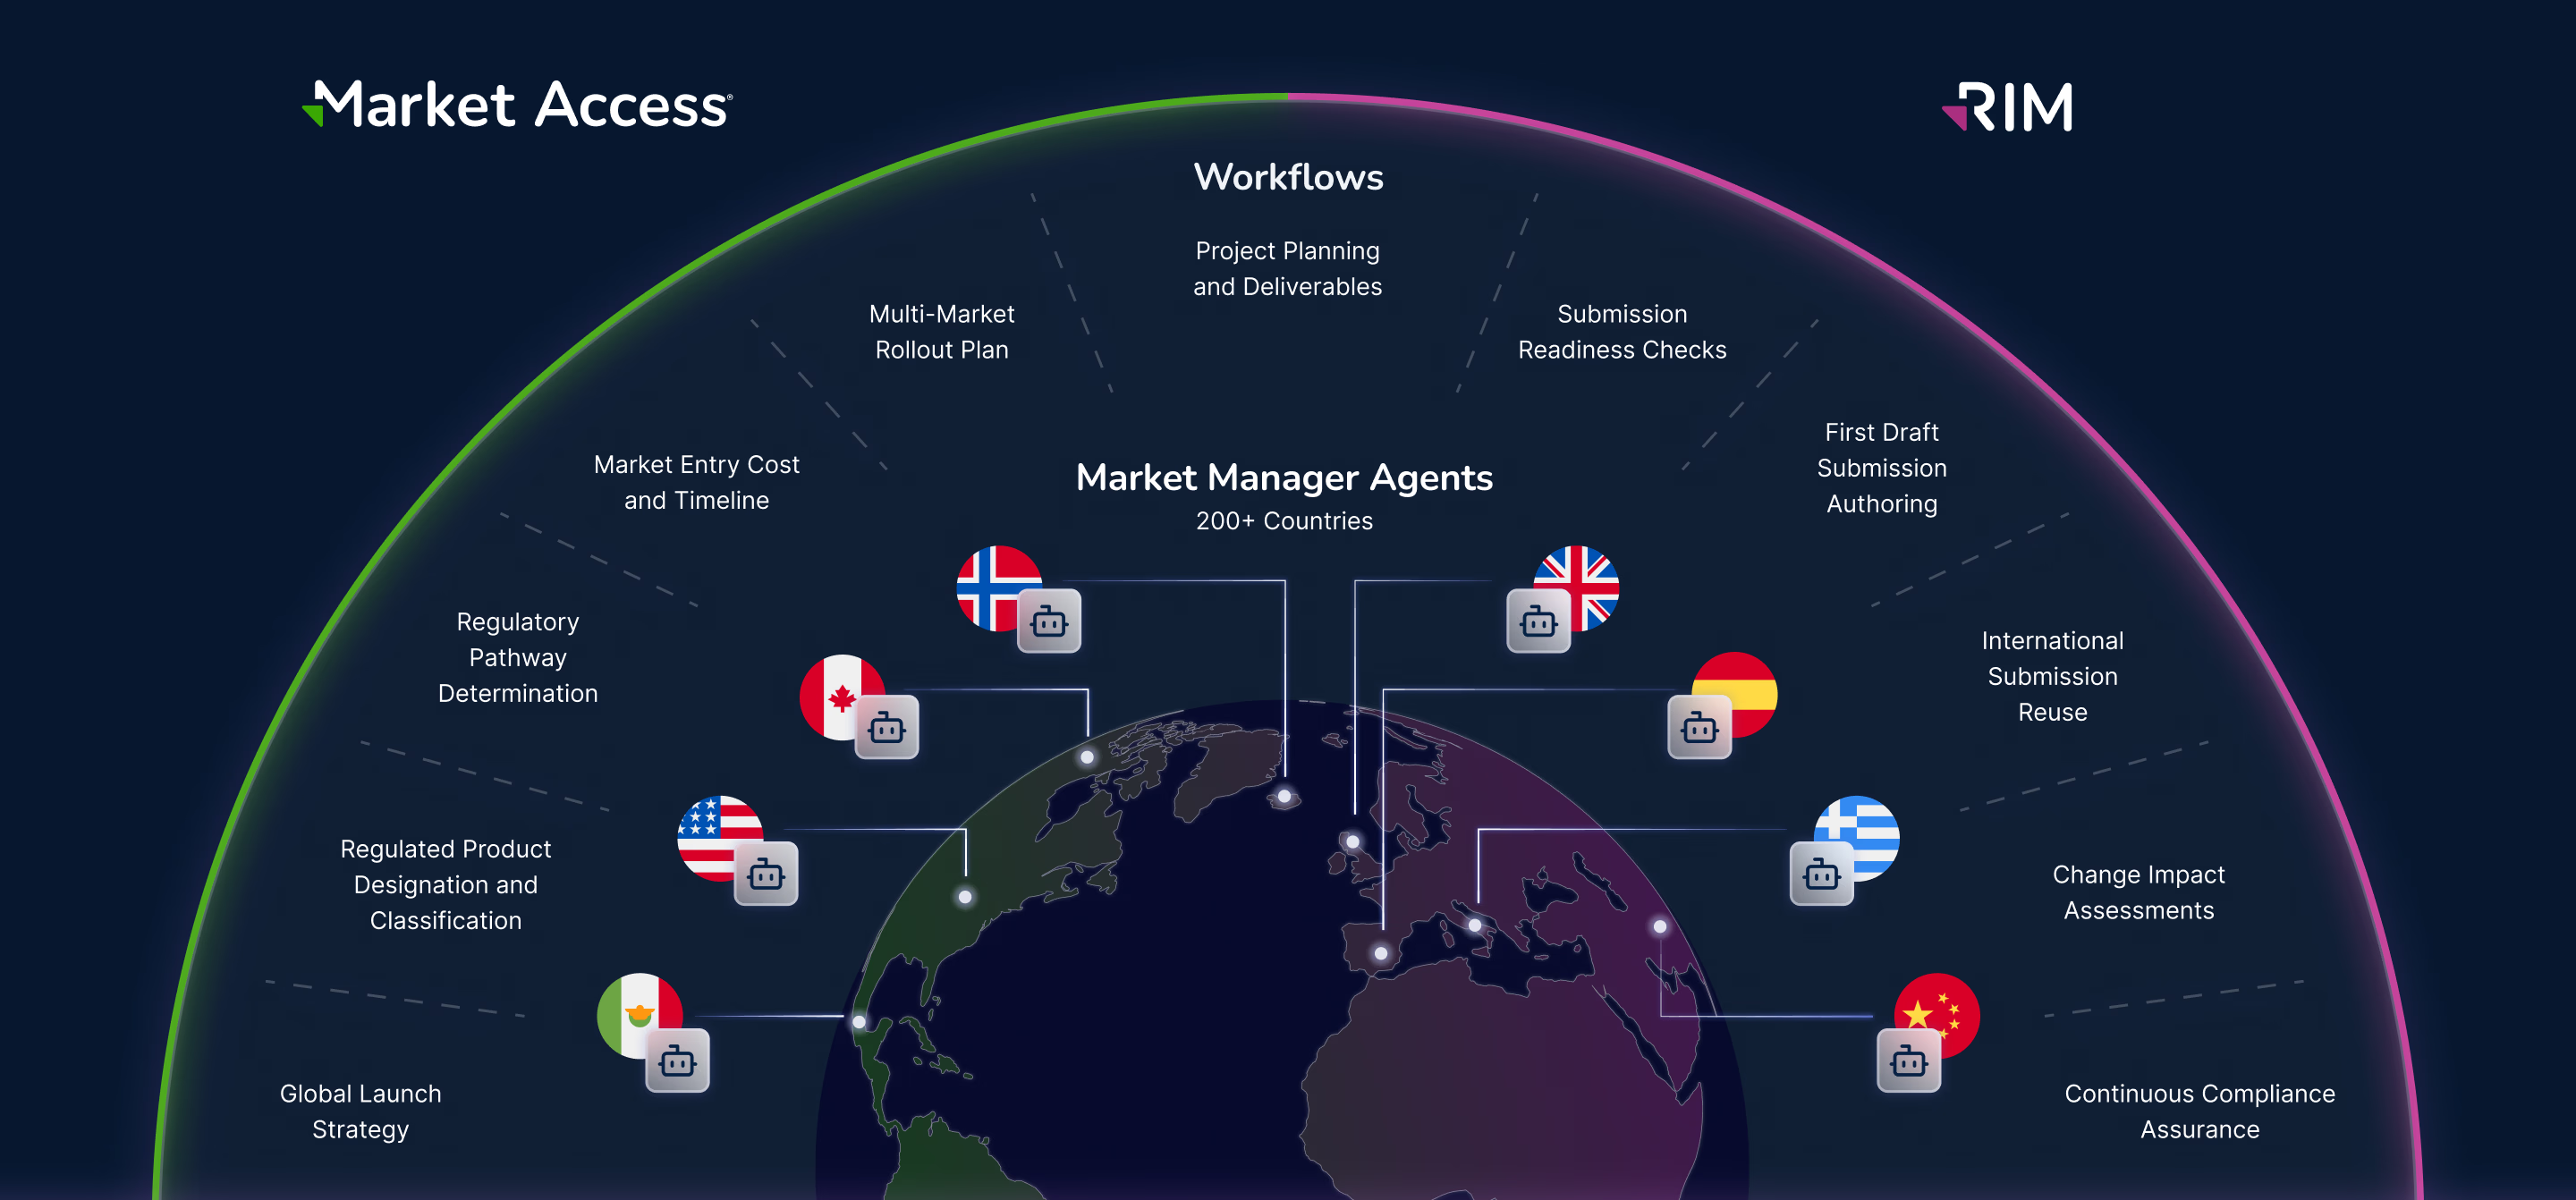
Market Access is powered by Rimsys AI, which works from a curated regulatory corpus spanning more than 200 markets, not the open web. Its Market Manager Agents are AI agents with market-specific regulatory knowledge, including pathways, requirements, and documentation, that support market assessments and global launch planning. The product is also designed to acknowledge areas where reviewer interpretation and expert judgment can affect regulatory outcomes rather than presenting those outcomes as more predictable than they are.

Market Access expands the Rimsys product portfolio from regulatory execution into market planning. Market Access helps regulatory and commercial leaders determine where to compete, while RIM manages the registrations, submissions, regulatory changes, product data, and ongoing compliance required to enter and remain in those markets. Together, they turn regulatory intelligence into coordinated action, helping regulatory teams accomplish more in every hour while giving business leaders greater visibility into opportunities, risk, and the work required to protect and grow revenue.

Beta Availability

Market Access will be available in beta beginning September 1, 2026. During the beta period, Rimsys will continue validating product capabilities and refining workflows ahead of broader commercial availability. 

Organizations interested in learning more about Market Access or participating in the beta program can visit rimsys.io/products/market-access.

About Rimsys

Rimsys is the heart of regulatory operations, connecting AI-native market planning with AI-embedded regulatory execution, giving medtech manufacturers one living system from market-entry decision to execution. 

Rimsys Market Access is the planning layer, answering not just whether a market can be entered, but whether it should be, returning classification, pathway, timeline, and cost before a team commits budget or resources to finding out. Rimsys RIM is the execution layer, managing registrations, submissions, regulatory changes, and compliance so teams can act on that plan with confidence.

Trusted by 6 of the top 12 global MedTech manufacturers and certified to ISO 27001 and SOC 2, with ISO 42001 certification for AI management systems underway, Rimsys keeps regulatory intelligence, product data, approvals, and change management continuously connected so teams can enter new markets with speed and assurance. Learn more at rimsys.io.

Media Contact
letschat@rimsys.io

I agree to the privacy policy including to Rimsys using my contact details to contact me for marketing purposes.
Thank you! Your submission has been received!
Oops! Something went wrong while submitting the form.
Hand holding smartphone showing email app with 12 unread messages notification.