Announcing AI-Native Market Access Product for Regulated Products

Learn More

Blogs

MedTech

Your eSTAR submission questions answered by FDA experts

By

Bruce McKean

September 8, 2023

4 min read

Your eSTAR submission questions answered

Starting on October 1, 2023, all 510(k) submissions, unless exempted, will need to be submitted electronically using eSTAR. eSTAR, which is short for Electronic Submission Template and Resource, is a dynamic PDF submission template that contains automation, guides, integrated databases, integrated policies and procedures, and automatic verification to help users prepare a comprehensive medical device submission.

Rimsys recently hosted a webinar, eSTAR submissions overview and live Q&A with FDA, to help the medtech community prepare for the quickly approaching eSTAR deadline. Patrick Axtell, Assistant Director for Tools and Templates Team, and Sajjad Syed, Software Engineer for Tools and Templates Team, from FDA joined our panel of Rimsys regulatory experts to provide an overview of eSTAR, demo, and live question and answer session. If you're interested in watching the webinar replay, you can find it here.

Due to high participation, we couldn't answer all questions live. This blog post provides Patrick's and Sajjad's answers to questions we couldn't get to during the webinar. Read below to learn what other industry professionals want to know about eSTAR!

Q: In the past, the sections were numbered Section 10 - Device Description.  How should section(s) be numbered in the eSTAR?

You may number attachments how you prefer, this includes numbering attachments according to the previous section numbers of the Format for Traditional and Abbreviated guidance document.  However, the numbering hierarchy in the IMDRF documents, which the attachment questions in eSTAR correlate to, may be better, since that numbering scheme is internationally harmonized.  So for example, if you are using the nIVD eSTAR, the IMDRF document specifies the Comprehensive Device Description would be numbered 2.04.01: https://www.imdrf.org/sites/default/files/docs/imdrf/final/technical/imdrf-tech-190321-nivd-dma-toc-n9.pdf.

Q: After October 1, 2023, will the In Vitro Diagnostics eSTAR Version be acceptable to use for a Dual 510(k) and CLIA Waiver by Application (Dual Submission)?

Yes, the next IVD eSTAR update will fully support Dual submissions, and the workaround we were previously implementing will no longer be needed. CLIA waiver submissions will continue to be submitted via current methods, and are not required to use eSTAR after Oct 1, 2023.

Q. It is not possible to view eSTAR online from cloud storage platforms. eSTAR like IFU Form and CDRH Form cannot be viewed online without users downloading these files. I will highly appreciate if  someone from the panel can give a solution to this. I understand these are special pre-programmed PDFs but it will immensely help if these can be shared for online viewing without having users download these files.

You need to choose to open eSTAR in the default application from within the cloud application you use, you can’t just click on it, since it will open in the browser, and browsers can’t open dynamic PDFs like eSTAR, Form 3881, Form 3514, and other forms FDA uses.

We can’t say that all cloud applications will work, but using Box does work. Once eSTAR is loaded in, click on the three dots, then choose the application you have installed, see screenshots below. I have Adobe Acrobat 2017 and Adobe Acrobat Pro both installed.

Be aware that every time you save the form, you upload it back into the cloud, so depending on how large it is and how fast your internet connection is, it may take many seconds to save each time, though you will see a progress bar.

Q: Does the eSTAR submission need to be submitted by only the US Agent or can this be done by the Foreign Manufacturer?

510(k) eSTARs, like 510(k) eCopies, can be submitted by foreign applicants.

Q: Will it be mandatory to use the eStar template for PMA supplement submissions?

Yes, by the end of 2023, eSTAR for PMA will be released. Similar to eSTAR for 510(k), there will be a pilot, guidance, and a transition period before it becomes mandatory.

Q: When do you expect to release the next version or sub-version of the non-IVD eSTAR template (4.x or 5.0) and will it be more elaborate on cybersecurity?

We are at the mercy of updating when policies are updated, which is irregular and unpredictable. Please email CDRH management and recommend a more consistent (e.g. quarterly) policy deployment schedule so that eSTAR can also have a more predictable deployment schedule. We are updating approximately one to every two months currently. Please be sure to read the Version History on page 1 of eSTAR regarding versioning and which version to use.

Q: How should a device categorized as a breathing gas pathway device in accordance with ISO 18562 be handled in eSTAR? One cannot find ISO 18562 categorization for a device in the eSTAR form.

This is a regulatory and policy question, and we are not sure what is meant by the statement “be handled.” Please reach out to OPEQ Submission Support OPEQSubmissionSupport@fda.hhs.gov with this question and they will direct your question appropriately.

Q: How often does FDA anticipate revising the eSTAR form?

eSTAR updates are almost exclusively timed according to policy updates, which the Tools and Templates Team has no control over. The T&T Team, as well as other groups in CDRH, are pushing for a quarterly update, where any policy updates in a quarter have an implementation date set for the beginning of the next quarter (e.g. Oct 1st, Jan 1st, Apr 1st). Please email Center management and promote this.

Q: The guidance Format for Traditional and Abbreviated 510(k)s: Guidance for Industry and FDA staff recommends a specific format for submissions. Biocompatibility, for example, is section 15. Should attachments in the eSTAR submission cross-reference the sections cited in the guidance? (So still using biocompatibility as an example, attachment 15.1 would be Cytotoxicity, attachment 15.2 would be Sensitization and so on)

The Format guidance, as well as the eCopy guidance, RTA guidance, and many other guidances, no longer apply for eSTARs.  Eliminating the applicability of guidances - simplifying the preparation process - is something we are proud of doing. You do not need to follow the numbering of this guidance.  See response above, where we recommend using the IMDRF numbering scheme instead, which is internationally harmonized.

Q: Where is information for risk assessments and management best included?

Depending on the type of submission you have selected (e.g. 510(k), De Novo, then Traditional, Special, Abbreviated), certain sections will become visible and active and some of them will be disabled. However, the bookmark pane will show all the sections in the entire eSTAR. Hence, please fill the sections that are visible to you since they are based on your selections. For example, if you have selected a Traditional 510k, the “Benefits, Risks and Mitigation Measures” section is not applicable. Hence it is not displayed and clicking on it leads a user back to the first page of the eSTAR. However, if you select “Special” as the 510k you are submitting, the “Benefits, Risks and Mitigation Measures” section does become applicable, and is displayed if you click on the bookmark. Also note that if you select De Novo, the section does become active as well. Another point to note is that for a Traditional 510(k) submission, if you do want to submit a Risks, Mitigations, Benefits report, you can attach it in the Performance Testing --> Bench Testing section of the eSTAR and identify it as “Other Non-Clinical Evidence”.

Q: Under Biocompatibility any idea when there will be endpoint questions for hemocompatibility or coagulation?

Hemocompatibility endpoint information, including Hemolysis, Complement Activation, and Thrombogenicity information, will appear depending on the tissue contact type and duration you choose. If you choose Implant Device >30 days, you will see this tab appear for example. The tabs that show are based on our Biocompatibility guidance.

Q: Guidance and Special Controls Adherence Section: According to the 510(k) electronic submission guidance document, identification of any applicable device-specific guidance documents or special controls should be included, which was done in the past; however, eSTAR requires each specific special control regulation or guidance recommendation to be listed. Can you please specify to what level of detail needs to be included within this section? The text box contains a character limitation, so it is unclear how specific information needs to be.

There are four textboxes like this in eSTAR. The specifications, characteristics, etc., of the device should not be provided in these textboxes. Instead, these textboxes should be used to refer to the attachments where these controls or recommendations are found, per each specific recommendation. These textboxes do expand as text is typed in, if needed. Keep in mind that the special controls, device-specific guidance citations would be divided among these four textboxes.

Q: Why are Advertisements included within the Labeling section, when they should be considered promotional materials? Could reference to advertisements be removed from this section?

Advertisements and promotional materials are included under the Other Labeling, and within the sub-attachment type "Other Labeling and promotional material" since this is consistent with Internationally harmonized document N9 and N13.  As the help text states, whether you actually need to attach it at all (it is an optional attachment) is dependent on the jurisdiction. For FDA, you would need to reach out regarding your device and current policy at opeqsubmissionsupport@fda.hhs.gov.

Q: I need to know how eSTAR would promote efficiency.

There are many efficiency gains with eSTAR, this is not an exhaustive list. The eSTAR website also includes some of this information.

-eSTAR complements the reviewer's internal smart review template used to review the submission (i.e. the questions correlate).  Therefore the reviewer is getting what they are expecting

-eSTAR provides a standardized format so that the reviewer (and applicant) always know where to find certain information

-eSTAR auto-updates many aspects of the submission, most notably it ensures the content is present negating the need for an RTA review by the reviewer, and therefore RTA holds

-eSTAR autocompletes information that you enter, ensuring you never need to enter the same information twice

-eSTAR includes built-in databases that ensure you see the information pertinent to your device (e.g. device-specific guidances), classification information automation, and the standards database, for auto-filling standards info accurately

-eSTAR has many forms built in, so that you don’t need to attach or upload them (e.g. T&A statement, Form 3514, 510(k) Summary, Declaration of Conformity, Form 3881)

-eSTAR guides the applicant through what is needed throughout, ensuring we collect everything that needs to be provided, and you know how to do it correctly

-eSTAR is collecting submission data in a structured format, which will help automate many aspects of our processing.  This will provide applicants and reviewers many benefits (e.g. automating the submission log-in process, therefore permitting reviewers to receive a submission within minutes of when the submission is uploaded in the CDRH Portal)

-eSTAR is intended to be used as a resource also, since it consolidates all the information needed (e.g., regulation links, guidance links, submission process information) to prepare a submission
‍

Similar posts

AI

From Regulatory Knowledge to Better Market Decisions: A Conversation with Steve Gens and James Gianoutsos

October 1, 2026

4 min read

‍

AI is moving quickly from experimentation to implementation across life sciences. For regulatory teams, that raises a bigger question than where AI can automate a task:

How can organizations use AI to make better decisions while maintaining the traceability, expert oversight, and confidence regulatory work demands?

That was a central theme when James Gianoutsos, Founder and CEO of Rimsys, joined Steve Gens, Founder and Managing Partner of Gens & Associates, on the Regulatory Executive Podcast.

Their conversation explored the changing role of regulatory intelligence, the introduction of Rimsys Market Access, the importance of transparency in regulatory AI, and what these changes could mean for regulatory teams.

Regulatory knowledge has been difficult and costly to scale

For global medtech organizations, regulatory knowledge has traditionally been highly dependent on people.

A company may have specialists responsible for particular countries or regions, along with local representatives, distributors, consultants, and other experts. Those people develop deep knowledge of local regulations, pathways, and expectations.

The problem is that the knowledge often remains distributed across the organization. And maintaining that market-by-market expertise can become expensive as a company expands its portfolio and geographic footprint.

As James described during the podcast, large manufacturers may rely on dozens or even hundreds of internal employees, third-party distributors, local representatives, and other specialists to understand what it takes to bring products into different markets. Smaller organizations may rely more heavily on outside consultants and partners. Either way, the model can be costly and difficult to scale. 

But the cost isn't limited to what companies spend on expertise.

Steve described the workflow as a series of “start, stop” cycles: ask a local expert, verify whether the regulation is current, gather more information, and repeat. Each cycle consumes regulatory capacity and extends the time it takes to give the business a confident answer.

That creates another potentially more consequential expense: the cost of waiting.

When it takes months to understand what entering a market will require, commercial and product teams have less visibility into when revenue might begin, what resources will be needed, or whether the opportunity is worth pursuing. Regulatory teams spend valuable capacity assembling and validating information rather than applying their expertise to more strategic work.

Ultimately, scaling regulatory knowledge isn't simply an efficiency problem. It affects how quickly and confidently the business can decide where to invest.

The bigger question: Should we enter this market?

That challenge is central to Rimsys Market Access.

Most market-entry research begins with questions such as: How is the product classified? What's the regulatory pathway? What are the requirements?

Those questions establish whether a company can enter a market. But they don't necessarily tell the business whether it should.

Market Access is designed to help regulatory and commercial teams develop an earlier view of what market entry could involve, including classification, regulatory pathway, timeline, cost considerations, requirements and potential gaps. 

For commercial and product leaders, that provides greater predictability before committing budget and resources.

For regulatory leaders, it means starting with a structured plan rather than assembling every market assessment from scratch.

Moving regulatory knowledge from individuals into a shared system

During the conversation, James described one of AI's most significant opportunities as moving knowledge that has historically lived in people's heads into software.

The goal isn't to eliminate regulatory expertise. It's to make that expertise more scalable.

Market Access works from a curated regulatory corpus, allowing market-specific knowledge to be applied across workflows without requiring every initial question to go through the individual who happens to know that jurisdiction. Outputs are designed to trace back to the underlying regulation, guidance, or law. 

That can also change where regulatory professionals spend their time. Rather than focusing as heavily on finding information and assembling initial assessments, they can focus on reviewing the evidence, identifying gaps, applying judgment, and advising the business.

Steve described the concept as a subject-matter-expert assistant. James called it a workforce multiplier. 

Trust requires opening the regulatory AI “black box”

Speed isn't enough when the information is being used to inform regulatory decisions.

James and Steve returned repeatedly to the importance of knowing where an AI-generated answer comes from. As Steve summarized it during the discussion, transparency builds trust.

Market Access is designed around that principle. Its outputs are grounded in a curated regulatory corpus and traceable to the specific regulations, guidance, or laws supporting them. That gives regulatory professionals the ability to verify an answer rather than simply accept it. 

Human judgment remains essential as well. Market Access provides an upfront assessment, not a guarantee of a regulatory outcome. Reviewer interpretation and other factors can still affect what happens during the regulatory process.

The objective isn't to make uncertainty disappear. It's to make what is known, what the sources say, and where uncertainty remains much easier to see.

Connecting the market decision to execution

The discussion also pointed to a broader evolution in regulatory technology: moving from systems that primarily store regulatory information toward systems that help teams act on it.

That's where Market Access and Rimsys RIM play distinct but complementary roles.

Market Access is the planning layer. It helps teams determine whether, when, and how to enter a market.

Rimsys RIM is the execution layer. It manages the registrations, submissions, regulatory changes, and ongoing regulatory work required to act on that plan. 

Together, they create a path from the initial market decision through regulatory execution rather than leaving planning and execution disconnected.

A new role for AI in regulatory work

One of the most interesting themes from Steve and James's conversation was that AI's impact may ultimately be less about replacing individual tasks and more about expanding what regulatory teams can accomplish.

Better access to market-specific knowledge can help professionals work across more markets, spend less time gathering information, and devote more capacity to the judgment and strategic work that requires their expertise.

The technology is changing quickly, but the fundamentals of regulatory work remain: reliable information, domain expertise, transparency, and human judgment.

The opportunity is to apply those fundamentals at greater scale while giving the business a clearer answer to the question that comes before execution:

Not simply, “Can we enter this market?”

But, “Should we?”

Listen to the full conversation

Hear the complete discussion between Steve Gens, Founder and Managing Partner of Gens & Associates, and James Gianoutsos, Founder and CEO of Rimsys, on the Regulatory Executive Podcast for more on regulatory AI, market-entry planning, transparency, and the changing role of regulatory teams.

‍

AI

How Market Access Builds Trust in Regulatory Intelligence

By

Bethaney Lentz

September 18, 2026

4 min read

Why source traceability, regulatory specificity, and human judgment matter when AI informs a market-entry decision.

Regulatory professionals are not short on AI options. General-purpose tools can answer questions quickly and often sound convincing. But in medtech, a convincing answer is not the same thing as a decision you can defend.

That distinction matters most before a company commits to a new market. Teams need to understand how a product is classified, which pathway applies, how long entry may take, what it may cost, where the important gaps are, and what uncertainty still remains. Commercial leaders need that visibility to decide whether the opportunity is worth pursuing. Regulatory leaders need to know that the assessment is grounded in sources they can verify.

That is the trust standard behind Rimsys Market Access. It is not simply about producing regulatory information faster. It is about helping medtech teams decide whether, when, and how to enter a market before committing budget and resources, with outputs linked back to the regulations, guidance, and laws behind them.

Why fluency is not enough for regulatory work

A general AI tool can be useful for brainstorming, summarizing, and exploring a topic. The challenge comes when an answer has real regulatory consequences. A plausible response may still be incomplete, based on the wrong designation, or disconnected from the current source that should govern the decision.

Medtech makes this especially difficult because the same product can be treated differently across markets and product designations. Medical device, IVD, cosmetic, consumer, OTC, and combination-product considerations can change the regulatory read. A market-entry assessment has to account for that context rather than assume one answer travels cleanly from one jurisdiction to another.

For regulatory teams, the natural next question is: Where did this answer come from? If the system cannot show the underlying authority, the expert still has to rebuild the research before trusting the result. That limits the value of the speed AI was supposed to create in the first place.

Trust starts with a decision you can verify

Market Access is designed around a different outcome. The goal is not to give teams another stream of regulatory information to interpret. It is to provide a source-linked market plan that helps them answer the bigger business question: should we enter this market, not just can we?

For a market assessment, that can include classification, regulatory pathway, timeline, cost considerations, key requirements and potential gaps. The output is designed to give commercial and product leaders an earlier, more defensible view of the opportunity while giving regulatory professionals the detail they need to review the basis for the assessment.

Trust comes from being able to inspect that basis. Market Access works from a curated regulatory corpus spanning more than 200 markets rather than relying on the open web. Answers are linked to the specific regulation, guidance, or law behind them, so a regulatory professional can verify the source instead of being asked to accept an AI-generated conclusion on faith.

Three things make regulatory intelligence more defensible

1. Source traceability

A regulatory answer is more useful when the person reviewing it can follow the evidence. Market Access links outputs back to their underlying regulatory sources. That changes the review process from “Do I trust the AI?” to a more practical question: “Does the cited source support this conclusion for our product and situation?”

That is an important shift for regulatory teams. Instead of assembling every market assessment from a blank page, they can review a structured, sourced plan, challenge assumptions, and focus their expertise on the places where judgment matters most.

2. Regulatory specificity

Trust also depends on context. Market Access is purpose-built for medtech regulatory complexity and supports cross-designation classification. The objective is to evaluate the product in the context of the market and the applicable regulatory framework, then carry that context into the pathway, requirements, timeline, cost considerations, and gaps that shape the market-entry decision.

This is where a purpose-built market-planning platform differs from a generic chat experience. The value is not an impressive answer in isolation. It is a connected assessment that helps the team understand what entering the market is likely to require and whether the opportunity still makes sense once those requirements are visible.

3. Human judgment stays in control

No system can remove every area of uncertainty from regulatory work. Reviewer interpretation and expert judgment still affect outcomes, and Market Access is not designed to promise approval.

Instead, Market Access is designed to make uncertainty more manageable. Regulatory professionals review and validate the work before it moves into execution. They can verify sources, assess the gaps, and apply company and product context that software cannot fully replace. The technology accelerates the research and planning. The regulatory professional remains responsible for the judgment.

What trust looks like in a market-entry decision

Imagine a team evaluating several markets for a new product. The commercial question is not simply whether registration is technically possible. The team needs to know which markets deserve investment now, which may require more evidence or resources, and which could put the revenue window at risk because the pathway is longer or more complex than expected.

Regulatory has a different, but connected, need. They have to understand the classification and pathway, identify the requirements most likely to slow the submission, and confirm that the plan reflects the relevant regulatory sources.

Market Access brings those views together. Commercial and product leaders get earlier visibility into timing, cost considerations, requirements, and risk so they can build a stronger business case. Regulatory teams get a source-linked plan they can review rather than a collection of unsupported conclusions. Both groups are working from the same market-entry picture before significant resources are committed.

Trust is not the end goal. Better decisions are.

It is easy for a conversation about regulatory AI to become a conversation about models, agents, or architecture. Those details matter, but they are not the business outcome.

The real value of trustworthy regulatory intelligence is what it lets a team do next. It can help a commercial leader defend a market investment. It can help regulatory surface issues earlier. It can give both groups a more predictable view of the work ahead before budget, R&D capacity, or launch timing is locked in.

And once the decision is made, the work does not have to stop at an intelligence report. Market Access is the market-planning layer. Rimsys RIM is the execution layer for registrations, submissions, regulatory changes, and ongoing regulatory work. Each can stand on its own, and together they create a path from the market-entry decision into controlled execution.

A more useful standard for regulatory AI

The question for medtech teams should not be whether AI can produce a regulatory answer. It can. The better question is whether that answer is specific enough, traceable enough, and reviewable enough to support a real decision.

Market Access is built around that standard: a defensible view of whether, when, and how to enter a market, grounded in curated regulatory sources and kept under human review. Because when the decision affects revenue, resources, and regulatory strategy, sounding right is not enough. Teams need to see why the answer is right, where uncertainty remains, and what it means for the decision in front of them.

Company

Product Updates

Rimsys Announces AI-Native Market Access for Regulated Products

By

Bethaney Lentz

September 1, 2026

4 min read

New software gives regulatory and commercial leaders a source-linked, actionable view of market viability, requirements, timing, and risk before committing resources

PITTSBURGH, PA, September 1, 2026 - Rimsys, the heart of medtech regulatory operations, today announced Market Access, a new AI-native product designed to help companies determine whether a market is worth entering and what it will take to get there. Market Access will be available in beta beginning September 1, 2026, with general availability to be announced separately.

Market Access replaces what can be a 6-to-9-month research and assessment process with a same-day, actionable read on market viability. It gives regulatory and commercial leaders the classification, pathway, timeline, cost considerations, and potential regulatory gaps they need to make a more defensible investment decision before committing budget and resources.

Predictability is at the core of Market Access. Rather than simply identifying a regulatory pathway, it helps teams understand what to expect, which requirements or gaps are most likely to slow down a submission, and where uncertainty remains. Every answer traces back to the specific regulation, guidance, or law behind it, allowing regulatory professionals to verify the underlying source rather than simply trust an AI-generated response.

Key features and benefits include:

  • Same-day market assessment: Provides an actionable read on whether a market is worth entering, replacing research and assessment that can take 6 to 9 months.
  • Cross-designation classification: Evaluates products across medical device, IVD, cosmetic, consumer, and OTC designations, so the same product receives the appropriate regulatory read regardless of how a given market classifies it.
  • Clear market-entry requirements: Provides classification, regulatory pathway, timeline, cost considerations, and key requirements before teams commit resources.
  • Earlier visibility into regulatory gaps: Surfaces the specific requirements and gaps most likely to slow down a submission and helps teams understand their potential severity.
  • Greater predictability: Provides visibility into anticipated regulatory response windows, risks, and areas of uncertainty so teams can better plan for what lies ahead.
  • Source-linked intelligence: Traces answers to the underlying regulations, guidance, and laws so regulatory professionals can verify the basis for an assessment.
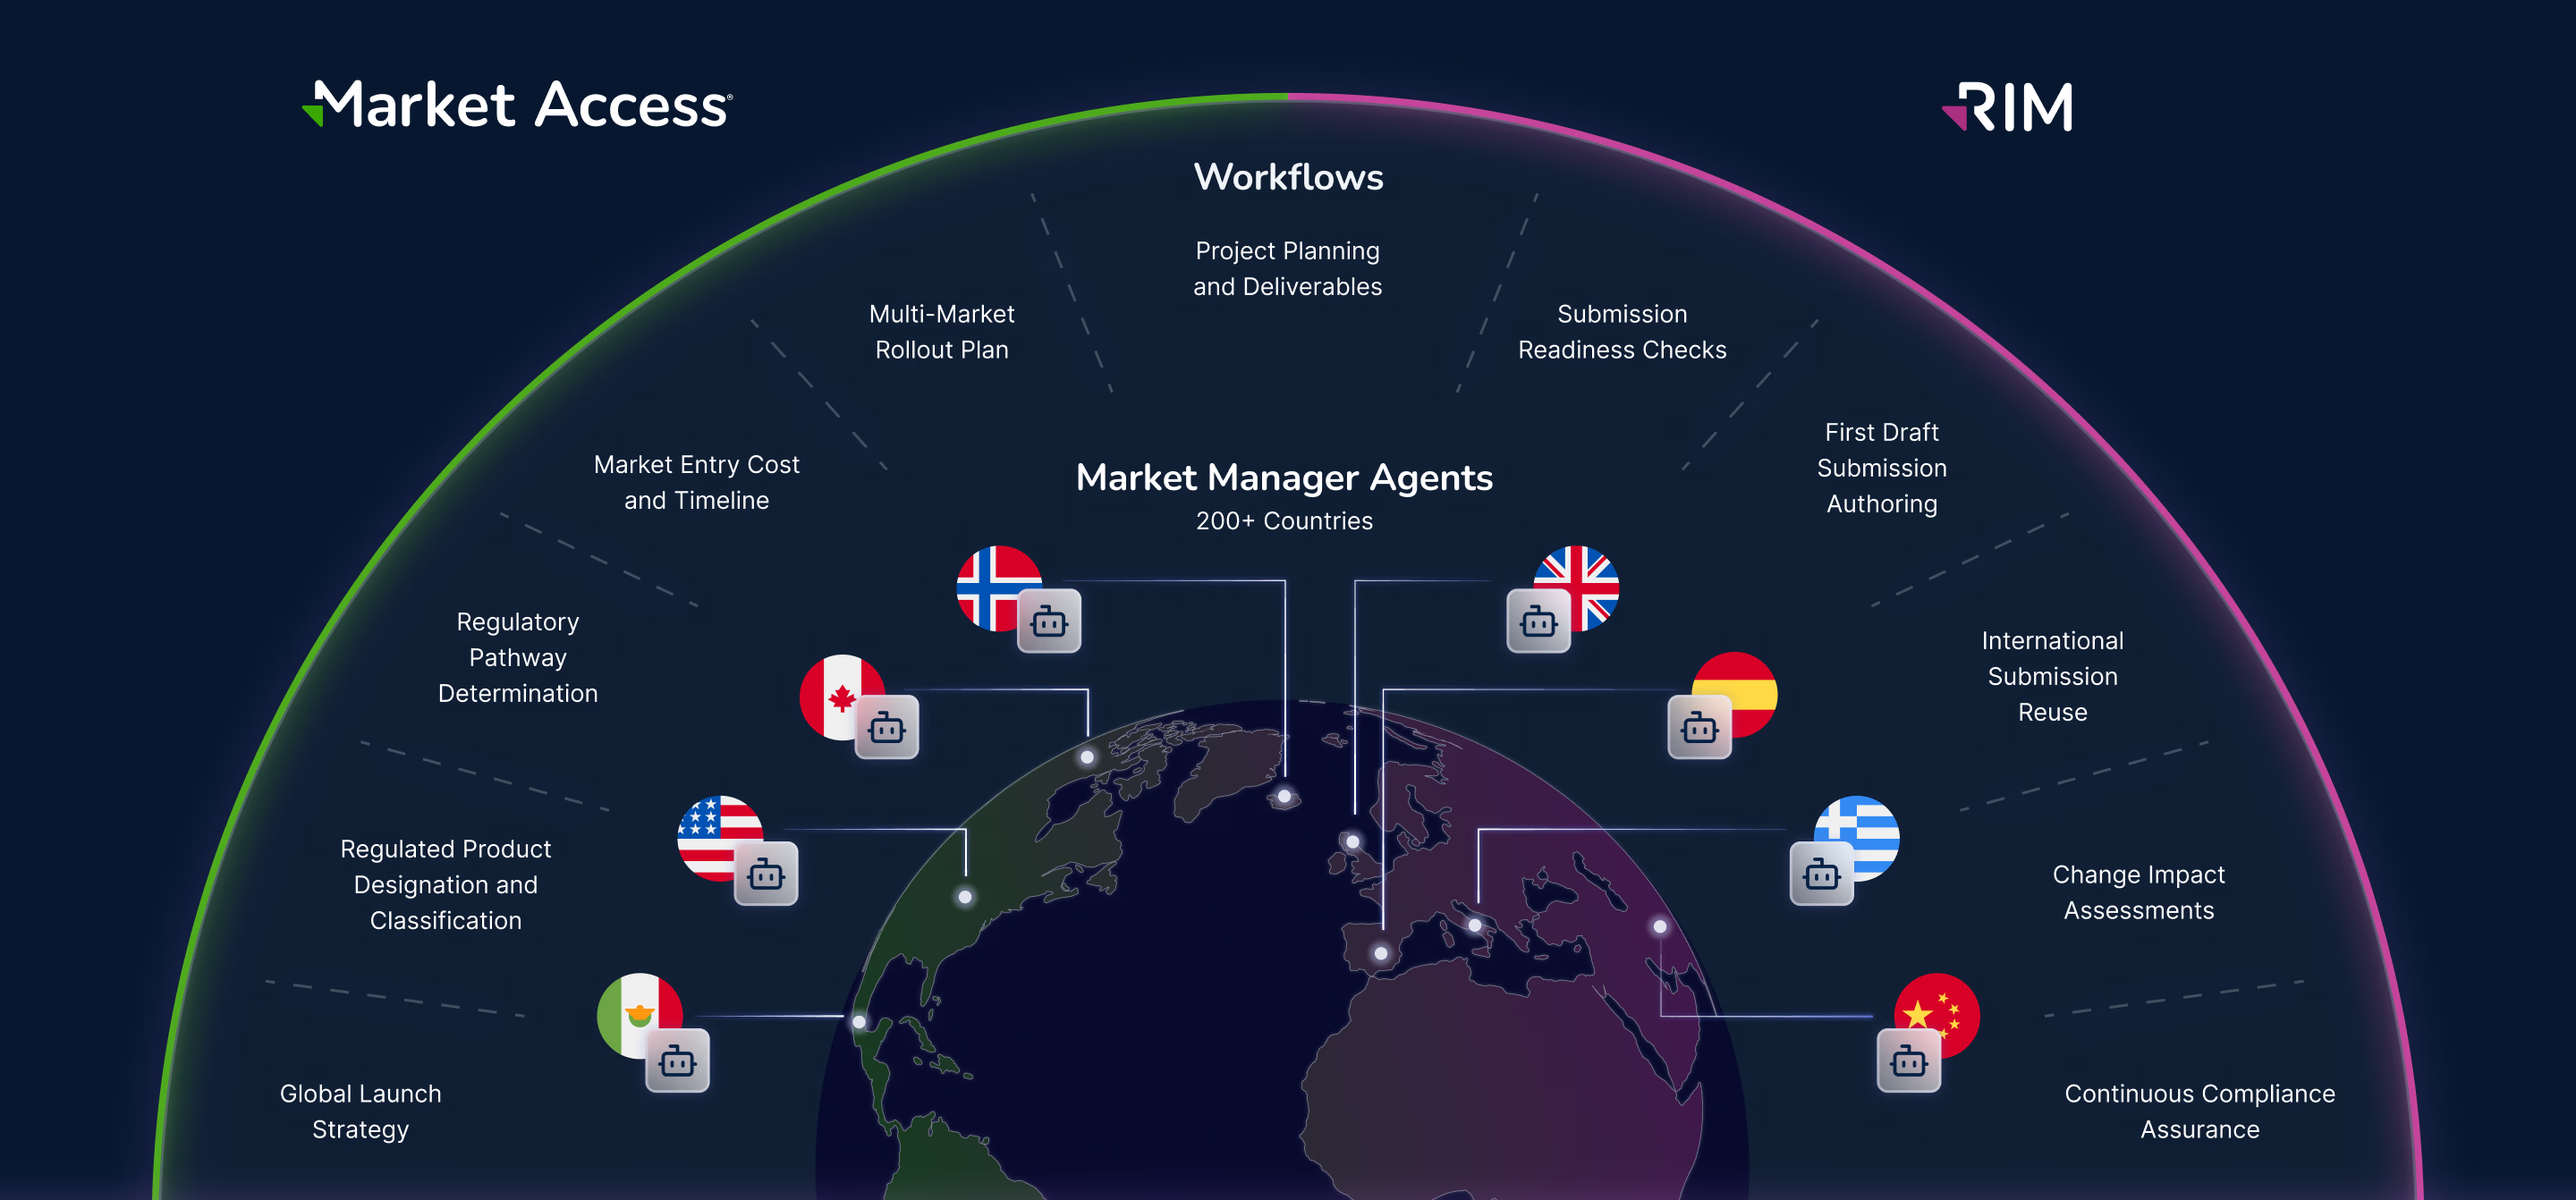
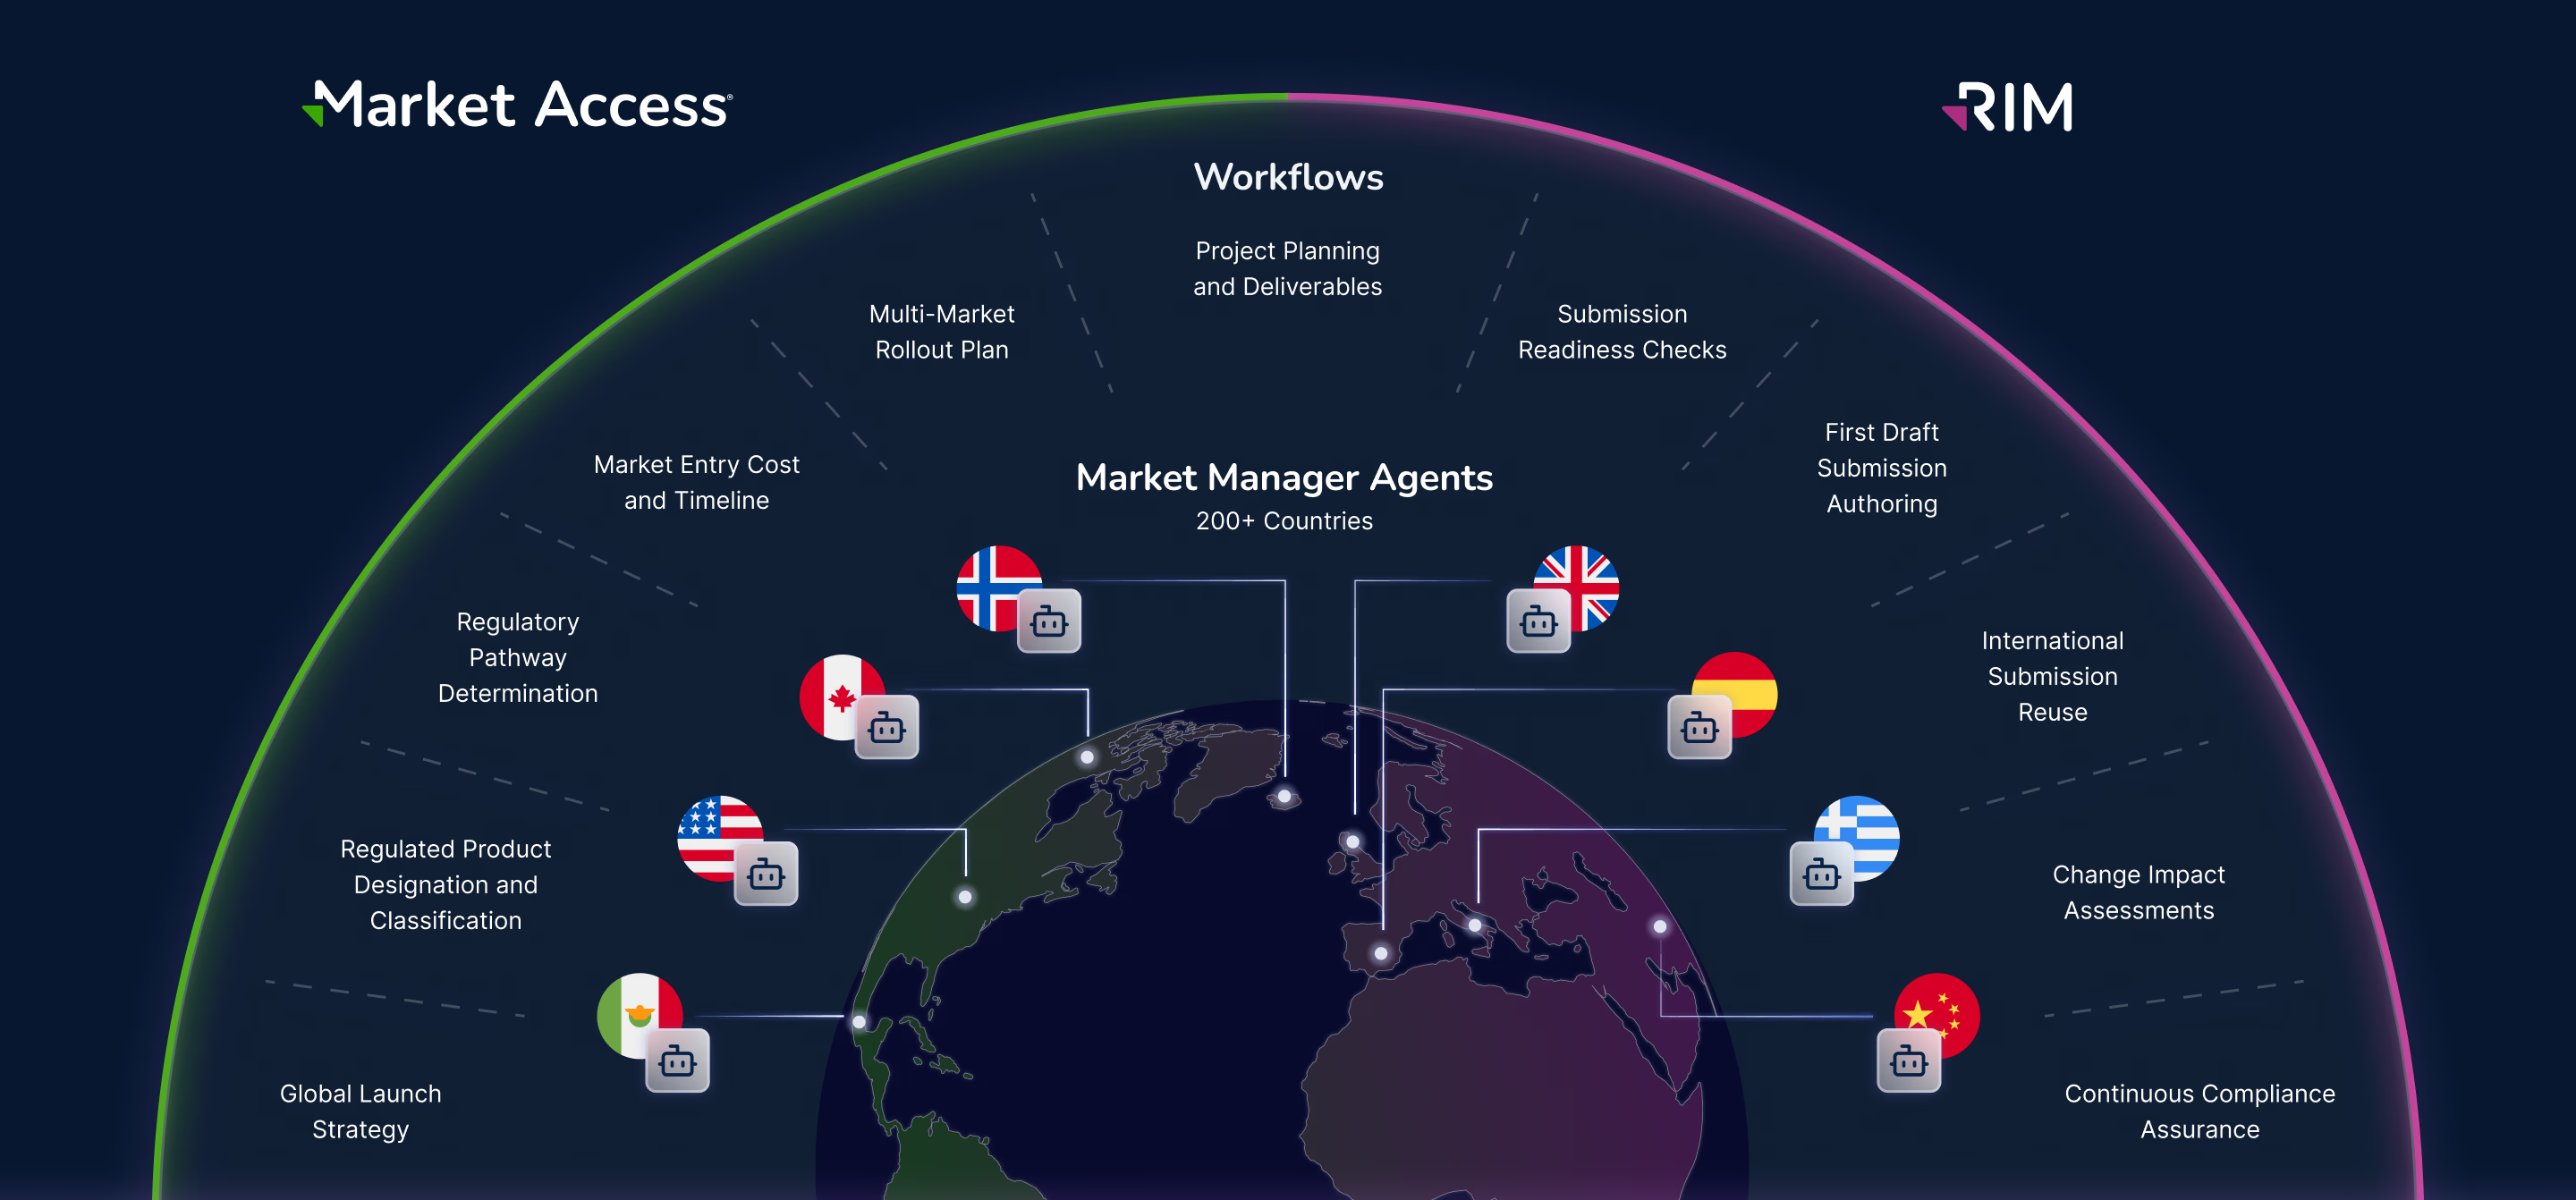
  • AI with human oversight: Uses Rimsys AI and Market Manager Agents to accelerate complex regulatory research while keeping regulatory professionals in control of review and validation.
  • Connected planning and execution: Complements RIM, connecting market-entry decisions with the registrations, submissions, regulatory changes, and ongoing compliance required to execute them.

“Regulatory knowledge has traditionally been highly dependent on individual expertise and market-by-market research,” said James Gianoutsos, Founder and CEO of Rimsys. “Market Access gives companies greater predictability before they make a market investment. Teams can understand whether an opportunity is worth pursuing, what it will take to enter, where regulatory challenges are likely to emerge, and what uncertainty remains. And because every answer is tied to its regulatory source, experts can verify the recommendation rather than simply trust the AI.”

Market Access is powered by Rimsys AI, which works from a curated regulatory corpus spanning more than 200 markets, not the open web. Its Market Manager Agents are AI agents with market-specific regulatory knowledge, including pathways, requirements, and documentation, that support market assessments and global launch planning. The product is also designed to acknowledge areas where reviewer interpretation and expert judgment can affect regulatory outcomes rather than presenting those outcomes as more predictable than they are.

Market Access expands the Rimsys product portfolio from regulatory execution into market planning. Market Access helps regulatory and commercial leaders determine where to compete, while RIM manages the registrations, submissions, regulatory changes, product data, and ongoing compliance required to enter and remain in those markets. Together, they turn regulatory intelligence into coordinated action, helping regulatory teams accomplish more in every hour while giving business leaders greater visibility into opportunities, risk, and the work required to protect and grow revenue.

Beta Availability

Market Access will be available in beta beginning September 1, 2026. During the beta period, Rimsys will continue validating product capabilities and refining workflows ahead of broader commercial availability. 

Organizations interested in learning more about Market Access or participating in the beta program can visit rimsys.io/products/market-access.

About Rimsys

Rimsys is the heart of regulatory operations, connecting AI-native market planning with AI-embedded regulatory execution, giving medtech manufacturers one living system from market-entry decision to execution. 

Rimsys Market Access is the planning layer, answering not just whether a market can be entered, but whether it should be, returning classification, pathway, timeline, and cost before a team commits budget or resources to finding out. Rimsys RIM is the execution layer, managing registrations, submissions, regulatory changes, and compliance so teams can act on that plan with confidence.

Trusted by 6 of the top 12 global MedTech manufacturers and certified to ISO 27001 and SOC 2, with ISO 42001 certification for AI management systems underway, Rimsys keeps regulatory intelligence, product data, approvals, and change management continuously connected so teams can enter new markets with speed and assurance. Learn more at rimsys.io.

Media Contact
letschat@rimsys.io

I agree to the privacy policy including to Rimsys using my contact details to contact me for marketing purposes.
Thank you! Your submission has been received!
Oops! Something went wrong while submitting the form.
Hand holding smartphone showing email app with 12 unread messages notification.