
2023 Regulatory performance report
Today at Rimsys, we unveiled the 2023 MedTech Regulatory Performance Report, a new set of insights into the state of medtech regulatory affairs. Compiled based on interviews with 200 regulatory professionals and executives, the study provides a detailed look into how regulatory teams are staffed, their processes, the tools they use, and ultimately how they perform.
Why did we create this study? There were two driving factors behind the research. The first was a common theme that we heard from a number of our customers: Regulatory leaders don’t have clear data and benchmarks. They don’t necessarily know how long a new market submission should take, and how to plan for or assess the work of their teams. While other studies look at the medtech industry broadly or the state of the regulatory profession, this study tries to build a comprehensive resource for regulatory (and company) leaders.
The second factor was really for ourselves and the team at Rimsys. As a company building solutions specifically for medtech regulatory affairs, we wanted more insight into where companies were successful, where they struggled, and where we can add value.
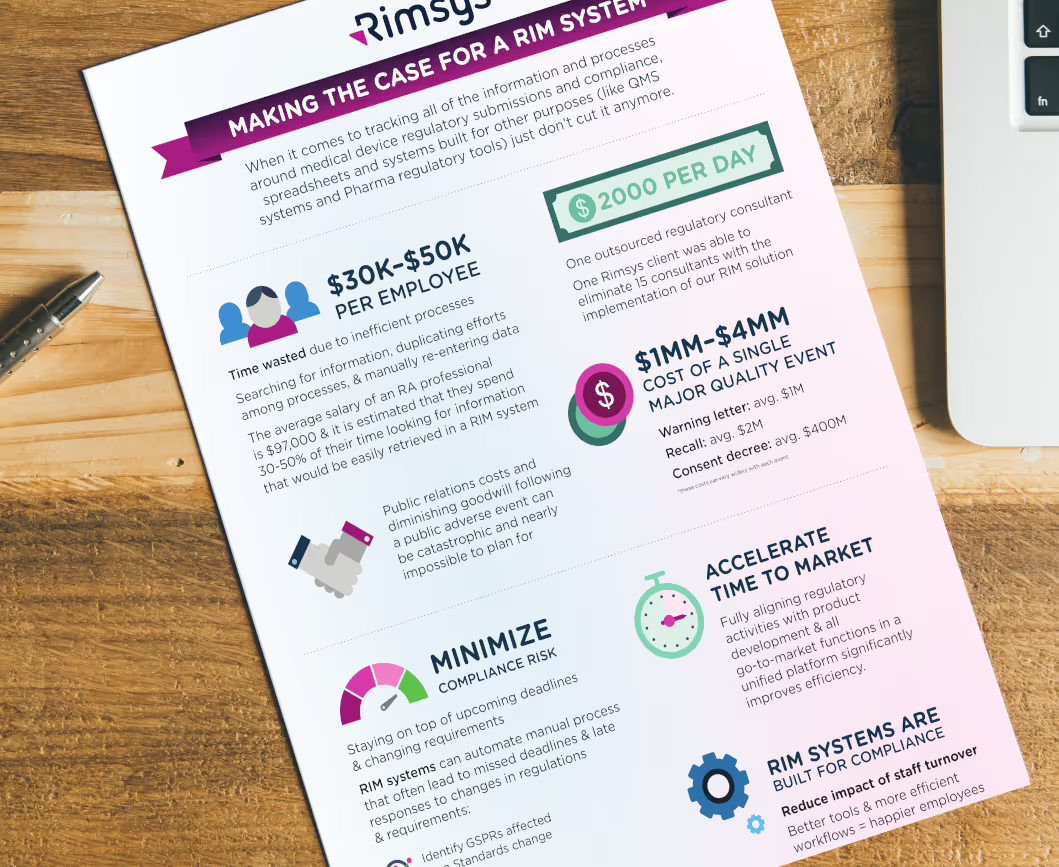
What did we find? Regulatory teams perform a lot of hero work and rate themselves highly for their accomplishments. At the same time there is a lot of opportunity for process improvements, and companies that invest in digital transformation for regulatory affairs see better performance.
Regulatory professionals are superheroes
Regulatory teams are generally pretty small. Most companies have less than 10 full-time regulatory professionals. These small teams complete an enormous amount of work. Last year on average, RA teams completed 50 license renewals, 50 license updates, and 10 new market submissions. This is impressive output.
Digging a bit under the covers, we found that this output relied heavily on the support of external consultants. 90% of companies use consultants to keep pace with their regulatory workload. Front-line employees also struggle with burnout. They were much more likely to report feeling under-resourced than regulatory leaders.
But process problems persist
A lot of regulatory work remains extremely manual. 70% of regulatory teams spend half their time or more on repetitive administrative tasks. All of this manual work increases the frequency of errors and required rework. 61% of companies reported a major non-compliance incident in the past 2 years.
Manual work also makes it difficult to complete regulatory projects in a timely fashion. Teams completed a lot of projects, but each took a long time. Over half of all companies spend 4 months or more on license renewals, license updates, and new market submissions.
Moving regulatory affairs forward
As regulatory requirements become more complex, there’s a natural question about how teams will work moving forward. MDR & IVDR in Europe have significantly increased the regulatory workload required to bring and keep products on the market. Will organizations be able to keep pace with the same resources, tools, and processes?
No, and the performance report shows that medtech companies are investing to improve their regulatory capabilities. The majority of companies are planning to increase the sizes of their RA teams in 2023, and 40% expect to increase their investments in regulatory software. Companies are increasingly adopting specialized software to better support regulatory processes.
Dig into the survey results
The full survey results provide insights into more aspects of regulatory performance. They show that companies need to take a deeper look into their processes and how regulatory resources are allocated. There are two ways to learn more:
- Visit the survey page to see the full results (the survey whitepaper can be downloaded at no cost)
- Watch the recording of our webinar with PA Consulting. We discuss the survey results in more detail and share our regulatory predictions for 2023

Post-market surveillance for medical devices in the European Union
This article is an excerpt from Post-market surveillance for medical device in the European Union.
Table of Contents
- What is post-market surveillance?
- What classes of medical devices require post-market surveillance?
- Components of a successful post-market surveillance plan
- PMS data requirements
- Post-market surveillance system goals
- Required post-market surveillance reporting
- Embracing post-market surveillance as an integral part of your quality program
- Getting started with post-market surveillance
Post-market surveillance (PMS) is designed to monitor the performance of a marketed medical device by collecting and analyzing field use data. Article 10 of the EU MDR and IVDR requires all device manufacturers to have a post-market surveillance system in place. The main elements of the PMS are laid out in Article 83, and additional details for lower-risk and higher-risk devices are covered in articles 84 and85, respectively.
In general, a PMS system consists of both proactive activities and reactive, or vigilance, activities. While post-market surveillance and vigilance are sometimes used interchangeably, vigilance consists of separate activities that feed post-market surveillance programs.
Post-market surveillance systems are used to collect and analyze data not only about the manufacturer’s device but also about related competitors’ devices that are on the market. Data collected through PMS procedures is then used to identify trends that may lead to, among other things, quality improvements, updates to user training and instructions for use, and identification of manufacturing issues.
Note that “market surveillance” encompasses activities performed by a Competent Authority to verify MDR compliance, and should not be confused with the topic of this ebook,“post-market surveillance," which is performed by the manufacturer.
All medical devices marketed in the EU require some level of post-market surveillance, and all medical device manufacturers must implement a post-market surveillance system (PMS). The requirements of the PMS, however, vary and should be “proportionate to the risk class and appropriate for the type of device” (MDR Chapter VII). In particular, the type and frequency of reporting vary based on a device’s risk class.
A post-market surveillance plan (PMS) is an integral part of a manufacturer’s quality management system and provides a system for compiling and analyzing data that is relevant to product quality, performance, and safety throughout the entire lifetime of a device. The PMS should also provide methods for determining the need for and implementing any preventative and corrective actions. A PMS system should include and define:
Surveillance data sources
With the increased focus on proactive risk identification in the MDR, it is important to design post-market surveillance systems that actively acquire knowledge and detect potential risks. It is not sufficient to rely solely on spontaneous reporting by healthcare providers, patients, and other stakeholders.
In addition to information coming from Clinical Evaluation Reports and complaint and adverse event reporting, typical sources of surveillance data include:
• Social media networks: Because many of your stakeholders may be communicating on social media networks, it is important to employ social listening techniques and/or tools to identify issues and concerning trends as they develop.
• Industry and academic literature: Any studies, academic papers, and other literature that addresses similar devices or the specific use cases for which your device is designed should be evaluated. In particular, risk factors and adverse events identified with similar devices should be closely examined. It is also important to identify newer technologies that may affect the benefit-risk ratio and establish a new definition of “state of the art” for the device type.
• EUDAMED: While the European Database on Medical Devices (EUDAMED) is not yet fully functional, it is intended to provide a living picture of the lifecycle of all medical devices marketed in the EU. Manufacturers should take special care to consider information for similar devices made available through the EUDAMED system in the future.
• Registries: Patient, disease, and device registries can provide information that informs the clinical evaluation process which provides input into the post-market surveillance system.
Data analysis methodology
A well-defined data analysis methodology will accurately identify trends and lead to defendable decisions in the application of post-market experience. Once the necessary information has been identified and collected, and potentially cleaned of incomplete or otherwise unusable data, the data needs to be analyzed.
The goal is to identify meaningful trends, correlations, variations, and patterns that can lead to improvements in the safety and efficacy of the device. There are many data analysis tools available that can assist with:
• Regression analysis that will identify correlations between data (e.g. the device location/geography correlates to battery life).
• Data visualization that can be useful in spotting trends in the data.
• Predictive analytics, which can be particularly useful with large data sets, to identify future trends based on historical data.
• Data mining, which is also normally used with large datasets, to organize data and identify data groups for further analysis.
Benefit-risk indicators and thresholds
The MDR requires that medical device manufacturers not only demonstrate the clinical benefit of their device but also quantify the benefit-risk ratio. The benefit of a device must be shown to clearly outweigh the risk for it to gain market approval. Article 2 (24) of the MDR defines the benefit-risk determination as “the analysis of all assessments of benefit and risk of possible relevance for the use of the device for the intended purpose when used in accordance with the intended purpose given by the manufacturer.”
A PMS system should clearly define benefit-risk calculations and the data used to support them. Post-market surveillance activities are critical in order to re-evaluate and maintain the benefit-risk calculations and determinations of a device throughout its life. Information that is gained through a PMS system can lead to:
• Identification of new risk factors.
• Adjustments to risk frequency and/or severity values based on actual use data.
• Adjustments to established risk calculations based on new “state of the art” technologies becoming available.
• Adjustments to established benefit calculations based on actual use data.
While complaint handling and other feedback tracking are more often described as part of post-market vigilance systems, they play a role in the more proactive post-market surveillance processes as well. A PMS system should define ...
To continue reading this ebook, download the full version.
The Future of MedTech Compliance: How AI Is Transforming Regulatory Affairs
MedTech regulatory affairs teams are facing a turning point. Regulations are expanding in number and complexity, resources are limited, and manual processes cannot keep up. At the same time, artificial intelligence (AI) has become a serious topic of discussion in regulatory circles. Leaders are beginning to ask: How can AI help us manage change, reduce risk, and accelerate compliance efforts?
The answer is clear: AI is no longer just a buzzword. When combined with effective regulatory information management (RIM), it can be a powerful enabler of efficiency, accuracy, and strategic decision-making.
Why AI is Trending in Regulatory Affairs
The Surge of Regulatory Data
Regulatory teams must now track requirements from multiple global markets. Each regulator frequently updates its regulations, guidances, templates, and recognized standards, which creates large volumes of data to organize and analyze. AI can scan and classify this information, highlight changes, and prepare it for structured use within RIM systems.
Doing More with Limited Resources
Most teams are expected to deliver more without additional staff. High turnover makes continuity difficult, and according to the 2024 RAPS Global Workforce Report, the number of professionals “open to work” has grown in North America and Europe. AI offers relief by taking on repetitive tasks such as document formatting or data entry, allowing experts to focus on higher-value work.
Global Complexity and Diverging Standards
No two markets are exactly alike. AI can help by flagging differences, surfacing potential risks, and recommending reusable content drawn from a company’s submission history. Faster, more accurate submissions directly improve time-to-market and compliance outcomes.
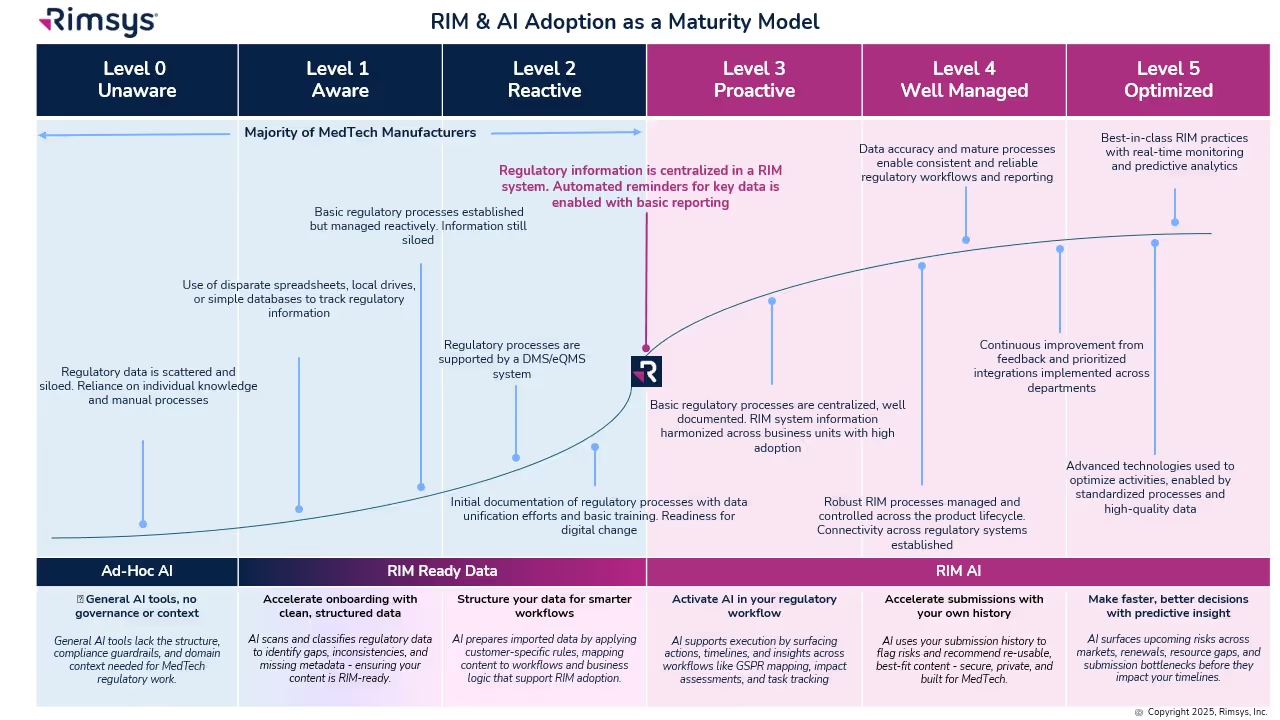
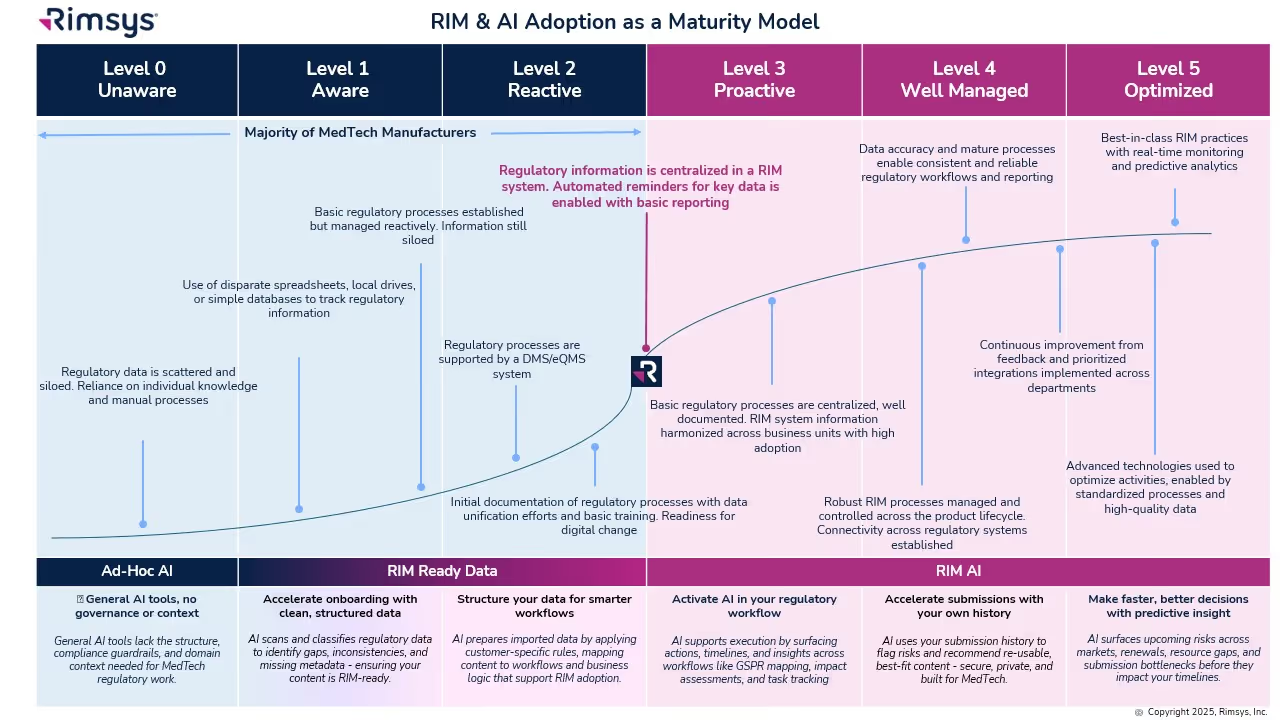
The RIM & AI Adoption Maturity Model
Not every organization is ready to fully embrace AI. Success depends on RIM maturity: how structured and centralized your regulatory processes and data are. The RIM & AI Adoption Maturity Model provides a roadmap from basic to optimized states.
- Levels 0–2: Early Stage
- Data is siloed and processes are ad hoc. AI provides value in isolated ways, such as cleansing records or scanning for regulatory changes.
- Level 3: Proactive
- A RIM system centralizes information. AI begins to surface reminders, deadlines, and global impact assessments.
- Level 4: Well Managed
- Processes are standardized across the lifecycle. AI generates insights, monitors KPIs, and supports reuse of regulatory content.
- Level 5: Optimized
- AI is fully embedded, delivering predictive analytics, continuous monitoring, and smarter decision-making.
Practical Applications of AI Today
Today, regulatory teams see the greatest opportunities in:
- Regulatory submissions: Automatically detecting changes in templates and suggesting updates.
- Document classification: Using natural language processing to tag and organize regulatory documents.
- Regulatory intelligence: Monitoring health authority updates and highlighting what matters most.
- Impact assessments: Linking changes (e.g., regulations/standards/design) directly to the affected products and registrations and evaluate the potential impact.
- Content reuse: Recommending approved content to accelerate submissions.
How to Start Your AI Journey in Regulatory Affairs
Adopting AI is not about jumping to the most advanced capabilities overnight. Instead, consider these steps:
- Assess your RIM maturity. Where does your organization sit on the 0–5 scale? What foundational gaps (data centralization, process standardization) need to be addressed first?
- Identify quick wins. Focus on repetitive, rules-based tasks where AI can add value without major disruption.
- Implement governance. Establish policies for safe, compliant AI use, particularly around data privacy and model training.
- Pilot in phases. Start small, validate results, and expand AI use as confidence and maturity grow.
- Keep people at the center. AI should enhance the expertise of regulatory professionals, not replace it.
Building a Smarter Future for MedTech Compliance
AI is becoming a trending topic in regulatory affairs not just because it’s new, but because it directly addresses the challenges teams face: rising complexity, limited resources, and scattered data.
For organizations that take this approach, the benefits are clear: lower compliance risk, faster execution, and stronger competitive positioning. AI does not replace regulatory professionals. Instead, it enables them to spend less time on manual tasks and more time on strategic contributions that improve patient access to life-changing technologies.
In other words, AI isn’t about futuristic transformation. It’s about helping regulatory teams step off the “data treadmill” and reclaim their time for what matters most: bringing safe, life-changing medical technologies to patients faster.

Rimsys Becomes the Trusted Regulatory Partner for 6 of the Top 12 Global MedTech Manufacturers
“Adoption by half of the top global MedTech manufacturers is a powerful validation that we’re not just a solution, we’re setting the new gold standard for regulatory excellence.”
Pittsburgh, PA - August 7, 2025 - Rimsys, the leading Regulatory Information Management (RIM) software purpose-built for the MedTech industry, today announced a significant milestone: 6 of the world’s top 12 medical device manufacturers now rely on Rimsys to manage and streamline their global regulatory operations.
This milestone further solidifies Rimsys’ position as the trusted partner to the world’s most innovative and quality-focused MedTech companies.
Click here for the full list of the top 12 global MedTech companies.
“Today’s regulatory environment demands more than spreadsheets. Leading manufacturers recognize that regulatory operations are mission-critical, revenue-generating departments and need systems to match that level of importance,” said James Gianoutsos, Founder and CEO of Rimsys.
Rimsys’ unified, enterprise-grade RIM platform centralizes and automates critical regulatory processes—including market registrations, Unique Device Identification (UDI), essential principles/GSPR, and submissions management—reducing compliance risk and accelerating market access. Specifically tailored to the needs of medical device and diagnostics companies, Rimsys enables seamless collaboration across RA, QA, and commercial teams while delivering the audit-ready transparency global regulators demand.
“As more organizations embrace regulatory digital transformation, Rimsys is proud to lead the industry forward,” added Gianoutsos. “Adoption by half of the top global MedTech manufacturers is a powerful validation that we’re not just a solution, we’re setting the new gold standard for regulatory excellence.”
To learn more about the Rimsys, please visit www.rimsys.io.
About Rimsys
Rimsys is the leading provider of Regulatory Information Management (RIM) software purpose-built for MedTech manufacturers. The comprehensive platform digitizes and automates regulatory activities, helping MedTech regulatory affairs teams to efficiently achieve regulatory compliance and get products to market faster. Rimsys is designed around MedTech workflows and supports a full breadth of regulatory functions including registrations, submissions, UDI, EUDAMED compliance, essential principles, and standards management in a unified platform. Rimsys is trusted by half of the world’s top 12 MedTech companies to power their global regulatory operations. For more information, visit www.rimsys.io.

Rimsys Announces Bulk UDI Submission and Rimsys Connect™ to Empower MedTech Regulatory Teams
New solutions deliver enterprise-grade data access and streamlined EUDAMED compliance, driving smarter, faster decisions across the business
Pittsburgh - April 29th, 2025 - Rimsys, the global leader in Regulatory Information Management (RIM) software for the MedTech industry, today announced two major enhancements to its platform: expanded Unique Device Identification (UDI) capabilities to support EUDAMED machine-to-machine (M2M) bulk transmission and Rimsys Connect™, a new enterprise Change Data Capture (CDC) solution that provides near real-time synchronization of Rimsys data with customers’ Business Intelligence (BI) solutions.
Together, these capabilities are designed to help MedTech organizations streamline compliance, reduce manual effort, and unlock the full strategic value of their regulatory data.
New UDI Capabilities Support EUDAMED Readiness
The UDI enhancements extend Rimsys’ industry-leading Universal UDI® framework, enabling MedTech teams to manage complex, global UDI programs in one unified RIM system. Key new capabilities include:
- Approving multiple records simultaneously via a simple, scalable workflow
- EU data governance support with all required attributes for EUDAMED transmission
- Bulk submission of records to both the GUDID and EUDAMED databases
These features allow teams to eliminate time-consuming, record-by-record processing, helping them meet the mandatory January 2026 EUDAMED compliance deadline with confidence.
“We’ve partnered closely with our customers to develop a UDI offering that meets increasing regulatory complexity and is easily scalable as new regulations come online,” said Adam Price, Director of Regulatory and Technical Programs at Rimsys. "We’re not only giving customers the ability to meet EUDAMED compliance but enabling them to manage their global UDI program in a single-sourced RIM solution for complete visibility.”
Introducing Rimsys Connect™: Enterprise Data Access, Redefined
Rimsys Connect™ offers enterprise customers a powerful new way to leverage regulatory data across the business. Built on a scalable, event-driven architecture, it provides secure, structured, near real-time streaming of Rimsys data into any modern data warehouse solution—such as Snowflake, Amazon S3, and Salesforce Bulk API 2.0.
“Rimsys Connect™ is not just a connector—it’s a strategic enabler,” said James Gianoutsos, Founder and CEO of Rimsys. “We’re giving regulatory affairs teams the ability to deliver insights that influence launches, accelerate tender responses, and align compliance with business impact. With Connect, RA teams become true strategic partners.”
By providing full access to customer data—registrations, UDI, projects, tasks, and custom attributes — Rimsys Connect™ supports a wide variety of enterprise use cases with customers’ own business intelligence solutions:
- Tracking on-time submission and decision KPIs
- Aligning registration timelines with product launch dates
- Conducting ROI analysis for renewals and market prioritization
- Accelerating tender readiness by combining RIM and PLM data
- Supporting post-market surveillance dashboards
While the initial release will focus on data access, Rimsys plans to expand Connect with curated BI templates and best practices to further accelerate enterprise customer time-to-value.
Solving the Data Fragmentation Problem for MedTech
Many regulatory affairs teams remain constrained by outdated tools, fragmented data sources, and increasing demands to deliver strategic insights to executive and commercial stakeholders. Rimsys Connect™ addresses these challenges by eliminating manual reporting workflows and enabling teams to analyze their regulatory data alongside financial, marketing, and quality systems.
“With Rimsys Connect™, regulatory teams can visualize and analyze their data in real time, assess launch readiness, and deliver more value to their organizations. This is how RA becomes a catalyst for better decisions—not just compliance,” said Gianoutsos.
Both Rimsys’ expanded UDI capabilities and Rimsys Connect™ will be available this summer. Those interested in learning more about these solutions and how they will enable greater automation, efficiency, and compliance can visit our booth at RAPS Euro Convergence May 13-15 in Brussels, Belgium, or sign up for Rimsys’ product update webinar on Thursday, May 22nd at 10 AM ET.
Read the press release here.
About Rimsys
Rimsys is improving global health by accelerating delivery and increasing availability of life-changing medical technologies. Rimsys Regulatory Information Management (RIM) software digitizes and automates regulatory activities, helping MedTech regulatory affairs teams to plan more effectively, execute more quickly, and confidently ensure global regulatory compliance. Rimsys is designed around MedTech workflows and supports a full breadth of regulatory activities including registrations, submissions, UDI, essential principles, and standards management in a unified platform. For more information, visit www.rimsys.io.
Contacts:
marketing@rimsys.io

Rimsys Enters Strategic Alliance Relationship with KPMG
PITTSBURGH – March 11, 2025 -- Rimsys, the global leader of MedTech Regulatory Information Management (RIM) software, today announced that it has entered into a strategic alliance relationship with KPMG to advance digital transformation in the MedTech industry.
“KPMG’s deep experience in advisory and business transformation services and exceptional reputation make them a valuable alliance relationship for us,” said James Gianoutsos, Founder and CEO of Rimsys. “KPMG is on the cutting edge of industry trends and has a wide breadth of experience in helping companies innovate and scale. We are thrilled to work with them to help MedTech teams transform their regulatory management processes and leverage the benefits of automation and digitization as part of their broader transformation strategy.”
Founded for and by MedTech regulatory affairs professionals in 2017, Rimsys was created to bring efficiency to regulatory information management and fill an inhibitive technology gap in an underserved industry. Rimsys has since grown to support the world’s MedTech leaders backed by a staff that understands their complex workflows and a robust, secure technology infrastructure that allows customers to scale Rimsys software to support their changing regulatory needs and requirements.
“There is tremendous innovation happening in the MedTech industry, and we are excited to work with Rimsys to help clients transform how they manage regulatory information for getting new products to market and sustaining their existing product portfolios. It’s critical to approach these programs as a holistic business transformation across people, process, technology, data, and governance & controls,” said Dipan Karumsi, Principal, Consulting Sector Leader for Life Sciences at KPMG.
“Through our strategic alliance with KPMG, we can further expand our reach to large and enterprise MedTech companies and continue our exponential growth,” said James. “Combined with KPMG’s experience helping organizations mature their data collection and transformation processes to reach RIM readiness, we can enable the MedTech industry to innovate faster, strengthen compliance, and most importantly, improve the availability of life-changing medical technologies.”
See the full press release here.
FDA’s Final Rule on LDTs: What manufacturers need to know
In July 2024, the FDA's final rule in 21 CFR Part 809 on laboratory developed tests (LDTs) went into effect, amending its previous regulations to make it clear that IVDs, including those that are manufactured in laboratories, are classified as devices under the Federal Food, Drug, and Cosmetic Act. Our blog post provides an overview of LDTs, FDA’s final rule, the phase out policy schedule, and how LDT manufacturers can prepare themselves for compliance.
What are LDTs?
Simply put, LDTs are IVDs that are designed, manufactured, and utilized within a certified laboratory and are typically used for high-complexity testing.
Historically, FDA has used enforcement discretion only on LDTs, which means that most LDTs haven’t been subjected to specific regulatory requirements. However, the volume of and risks associated with LDTs have grown over the years. Some examples of modern LDTs include glucose tests, genetic tests for cancer and infectious diseases, and newborn screenings for early diagnostics. Without a regulatory framework in place, patients are at greater risk of receiving inaccurate test results, forgoing necessary or undergoing unnecessary treatment, and adhering to misleading or false product claims, possibly endangering patients and leading to higher healthcare costs.
FDA’s Final Rule on LDTs
Following the final rule that was issued, LDTs are now subject to the same regulatory requirements as other IVDs, including premarket reviews, quality system requirements, labeling requirements, adverse event reporting, and device listing and registration. To prevent disruptions in patient care, there is a four-year transition or phaseout period consisting of the following five stages:
- Stage 1 (May 6, 2025): LDT manufacturers will be expected to comply with FDA medical device reporting (MDR) requirements, correction and removal reporting requirements, and quality system (QS) requirements for complaint files.
- Stage 2 (May 6, 2026): LDT manufacturers will be required to comply with IVD registration and listing requirements, labeling requirements, and investigational use requirements.
- Stage 3 (May 6, 2027): LDT manufacturers will need to comply with all other QS requirements not covered in Stage 1.
- Stage 4 (November 6, 2027): Unless a premarket submission is received before the start of this stage, LDT manufacturers of high-risk products will need to comply with premarket review requirements for IVDs that may be classified into class III or that meet the requirements of section 351 of the Public Health Service Act.
- Stage 5 (May 6, 2028): LDT manufacturers of moderate and low –risk products will need to comply with premarket review requirements for IVDs unless a submission is received before the beginning of this stage.
Manufacturers of LDTs that don’t meet the requirements in each stage are deemed non-compliant to the regulations governing IVDs and may be subject to FDA 483 observations or warning letters, financial penalties, and even worse, involuntary removal of products from the market.
Note that some LDTs will be exempt from these requirements. Refer to the FDA’s website for more guidance.
Preparing for Compliance
Despite a four-year phaseout period, it’s crucial for LDT manufacturers to start assembling a compliance plan. Starting as early as May 2025, manufacturers will be required to comply with FDA Medical Device Reporting (MDR) requirements, correction and removal reporting requirements, and quality system requirements for complaint files.
It's good practice to conduct an internal regulatory assessment to ensure you have the resources, processes, and tools in place to successfully meet new requirements for LDT devices. It’s also essential to make sure your team is well-versed in these new requirements and the documentation and timelines involved. Including all relevant stakeholders early on, getting a comprehensive project plan in place, and meeting regularly to ensure all tasks are completed would be helpful during the phaseout period and beyond.
Regarding the LDT changes, the FDA has provided a Q&A sheet that you may find helpful.
If you're looking for guidance on FDA premarket submissions, see our Beginner’s Guides to the FDA 510(k), De Novo, and PMA processes.
How Regulatory Tools Can Help
FDA’s final rule on LDTs will add complexity to the regulatory information management of laboratory diagnostic tests. There are digital solutions that can help manufacturers stay current on updated regulations and manage the additional information and documentation needed because of these updates.
A regulatory intelligence database like Rimsys Intel can provide detailed global market entrance requirements, application timelines, fees, risk class specifications, and documentation needed for medical devices and IVDs so that manufacturers can start preparing their premarket strategies.
Regulatory Information Management (RIM) software like Rimsys can help boost efficiency, reduce compliance risk, and increase collaboration by centralizing regulatory information and automating time-consuming, manual processes. As a result, medical device manufacturers gain complete visibility into their submission management and selling status so that they can plan more effectively, avoid costly product delays, and execute faster.

Planning Your Enterprise’s UDI Strategy for EUDAMED and Beyond
Rimsys recently hosted a webinar with RAPS titled “Planning Your Enterprise’s UDI Strategy for EUDAMED and Beyond.” During the session, our experts, Adam Price, Director of Regulatory and Technical Programs, and James Gianoutsos, Founder and CEO, provided best practices to help MedTech companies navigate the complexities of Unique Device Identification (UDI) compliance. The session covered key topics such as comparing US and EU UDI regulations, preparing for the January 2026 mandatory EUDAMED submission date, and using EUDAMED as a foundation for a global UDI strategy that can scale as additional markets adopt UDI requirements.
Here are some of the key topics we discussed to help Medtech teams prepare their strategy:
1. The importance of UDI to regulatory affairs teams
UDI is a globally unique identifier for medical devices that helps improve traceability, manage adverse event reporting, and ensure compliance with regulatory requirements. Regulatory affairs teams play a critical role in both establishing and maintaining UDI data to meet submission timelines and market placement requirements.
Responsibility for UDI management can vary between organizations, with some MedTech companies assigning the responsibility to IT or supply chain teams. When we asked our audience who is responsible for UDI management at their company, 80% of respondents said that it was their regulatory affairs team, emphasizing UDI’s crucial role in market placement.
2. Comparing US and EU UDI Requirements
FDA GUDID requirements have been in place since 2014. As such, some MedTech teams may plan to use their GUDID submission strategy as a baseline for the development of their EUDAMED submission strategy. While the US FDA’s GUDID system has standardized requirements, the EU’s EUDAMED is more complex due to additional data fields, language-specific entries, and the use of the EMDN nomenclature instead of GMDN. EUDAMED also introduces the Basic UDI-DI, which connects related devices within family groupings and links different EUDAMED modules. Companies must adapt their UDI strategies to account for these variations.
“Unlike the standardized US GUDID, the EU’s EUDAMED is more complex, featuring multiple device categories, language-specific data, and the EMDN nomenclature.” - Adam Price
3. The Interconnectivity of EUDAMED
EUDAMED consists of multiple interconnected modules. The Basic UDI serves as the key data element linking these modules together. This interconnected structure ensures that regulatory bodies, manufacturers, and economic operators can efficiently track devices throughout their lifecycle. Properly aligning UDI submissions with these modules is essential for seamless compliance and data consistency.
“The interconnectivity of EUDAMED is really important because the Basic UDI family groupings are the key piece of data that is going to tie the different modules together. For example, your notified body certificates for the regulations will have your Basic UDI numbers indicated on them. And then that's connected to the Basic UDI numbers that you have associated to your single registration number.” - Adam Price
4. Deadlines, Compliance Risks, and Recommended Actions per Device Category
The deadline for mandatory UDI submissions to EUDAMED is January 1, 2026, with a six-month grace period until June 2026 for devices already on the market. Failure to meet these deadlines could result in compliance risks, including regulatory penalties and restricted market access. Different categories of devices are subject to specific rules:
- Regulated Devices: Devices compliant with MDR or IVDR regulations must be linked to Basic UDI groupings and classified accurately under EMDN. Recommended actions include ensuring all data aligns with the new requirements and preparing for immediate submissions of new devices after January 2026.
- Legacy Devices: Devices compliant with older EU directives that remain on the market do not require a Basic UDI but must still have all essential data submitted. Companies should consolidate and verify legacy data, ensuring consistency with prior regulatory approvals.
- Non-Registered Devices: Devices no longer placed on the market but still have post-market surveillance activities must be registered for traceability. Manufacturers should identify these devices, gather historical data, and ensure that post-market surveillance records are aligned with regulatory expectations
.
5. Data Preparation and Submission Process
Data preparation for EUDAMED compliance is going to take some time, and the time to start preparing is now. It's important to recognize some of these high-level steps needed for effective UDI data preparation and submissions so that MedTech teams can plan appropriately:
- Identify: Collect and review all required UDI data, including language-specific fields and Basic UDI groupings.
- Verify: Ensure data accuracy and alignment with regulatory requirements, using EUDAMED’s business rules as a reference.
- Format: Structure data to meet EUDAMED’s formatting and enumeration rules, ensuring compatibility with submission methods.
- Submit: Transmit data using manual entry, XML uploads, or automated machine-to-machine (M2M) transmission, depending on the volume and complexity of your product portfolio.
“Ultimately, manufacturers need to be in a position to have all the data in for all products - not just the new products on the market but for all products by the end of that transitional period. It sounds a little apocalyptic but given the right planning, I think it's very achievable." - Adam Price
6. Getting Data into EUDAMED
Manufacturers have multiple options for submitting UDI data into EUDAMED, depending on their resources and portfolio size. Data can be manually entered through the EUDAMED user interface, uploaded via XML files, or submitted using automated machine-to-machine (M2M) transmission. Organizations managing large product portfolios are encouraged to adopt M2M transmission to ensure efficiency and data accuracy. Testing data submissions in the EUDAMED playground before official submission is highly recommended to identify and resolve any formatting or compliance issues.
“With more products, more data, and more BUDI families, there's more complexity. And as you're looking across your product portfolio, that's something that you need to consider as you plan your submission strategy.” -Adam Price
7. UDI Management Solutions
Organizations can choose from various UDI management solutions depending on their size and needs:
- Spreadsheets: Might be suitable for small companies with limited product portfolios but are time-consuming to manage and error-prone
- Repurposed Internal Systems: PLM or PIM systems can manage UDI data but may require manual processes and in-house expertise.
- Purpose-Built Connectors: Provide transmission capabilities for UDI data but don’t provide data capture and management
- RIM Systems (e.g., Rimsys): Provide a centralized, automated solution that integrates UDI data with product registrations and certificates, supporting seamless M2M transmissions.
“For large, complex product portfolios, RIM systems like Rimsys provide automated UDI management and seamless M2M transmissions.”
“Having an integrated data hub that's the source of truth for your UDI information is key. Then being able to associate your UDI data with all other data needed for UDI transmission in a RIM system, your registrations, certificates, legal entity, manufacturer, manufacturing locations, all the product data, all your BUDI data, having that all centralized in a RIM system is really critical to understanding and organizing your data in a way that can be meaningfully managed moving forward.” - James Gianoutsos
8. EUDAMED as a Foundation for Global UDI Compliance
There is a tremendous opportunity for EUDAMED’s comprehensive data requirements to serve as a foundation for global UDI compliance, as many countries are adopting similar regulations. Australia, Switzerland, the UK, and India are among the markets expected to enforce UDI requirements soon, with varying deadlines and data requirements. By using their EUDAMED data output as the foundation for a global UDI program, MedTech companies can build a UDI strategy that positions them to meet current and future requirements as they evolve.
“The application of UDI is not a one and done approach like it had possibly felt for manufacturers in the past. These requirements are going to need to be continually maintained from the perspective of your product characteristic data as well as new technologies as they're being introduced. So it's maintaining compliance to the current requirements, establishing compliance to the new countries’ UDI programs as those get rolled out, and being in a good position to control and consistently apply your UDI data.” -Adam Price
Conclusion
With the January 2026 deadline approaching, companies must act now to ensure compliance, streamline submissions, and prepare for future UDI requirements worldwide. Establishing a robust UDI strategy today will not only support compliance with EUDAMED but also lay the groundwork for expanding regulatory demands in other global markets. Leveraging centralized systems like RIM platforms can help manage the increasing complexity of global UDI regulations, ensuring consistent and efficient compliance.
“This seems to be a trend globally that UDI has become this single source data point to control and manage and maintain your products on the market. And so EUDAMED is just really the tip of the iceberg. Even though we know how complex it is, this is really just the beginning of this global UDI strategy that you have to be thinking about as you go into EUDAMED preparation.” -James Gianoutsos
Those interested in watching the webinar can access the recording here.
With an integrated and automated approach to UDI, Rimsys is a trusted and proven solution to help MedTech teams simplify EUDAMED compliance and assemble a global UDI program that meets current and future requirements. Request a custom demo rimsys.io/demo.