The beginner's guide to the FDA De Novo classification process
This article is an excerpt from The beginner's guide to the FDA De Novo classification process ebook.
Contents
- Introduction
- Chapter 1: What is an FDA De Novo request?
- Chapter 2: Contents of a De Novo request
- Chapter 3: Submitting a De Novo request
- Appendix A: Acceptance review checklist
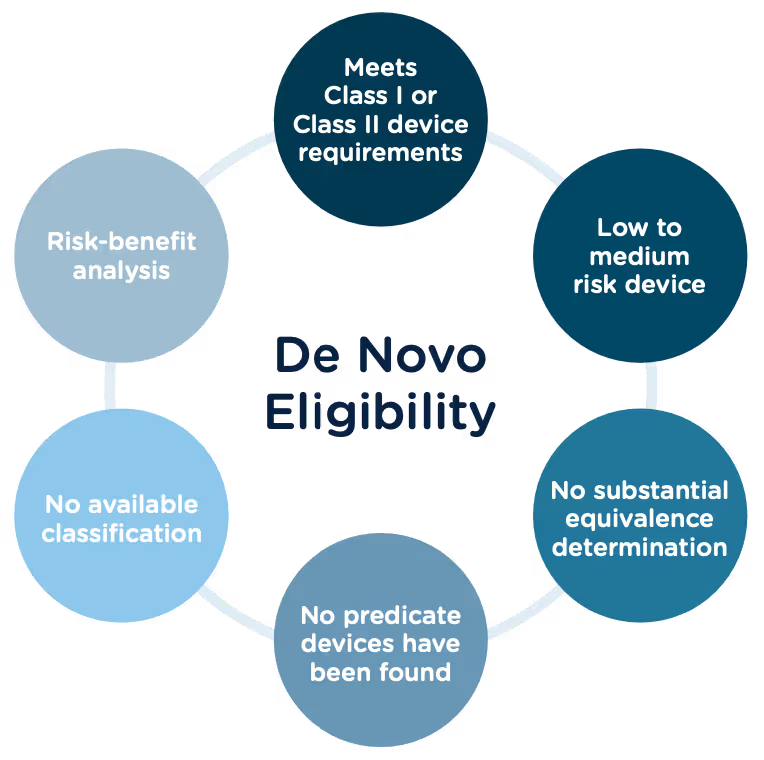
Congratulations, you have successfully developed a new medical device! Now you need to take it to market. Normally in the United States this would mean completing a 510(k) submission. However, the 510(k) relies on “substantial equivalence”—a comparison to a similar device already on the market (also called a predicate device) to assess the risk profile of the new device. What if your device is totally new, and there isn’t a similar device to compare it to? Enter the FDA De Novo process. The De Novo process provides a pathway to market for novel devices with a low to medium risk profile.
What does De Novo mean?
According to the Merriman-Webster dictionary, de novo is a Latin word meaning “as if for the first time; or anew.” Perfectly fitting that the FDA uses this term “De Novo” to describe market approval requests for new medical devices or technology where there is no comparable predicate device on the market.
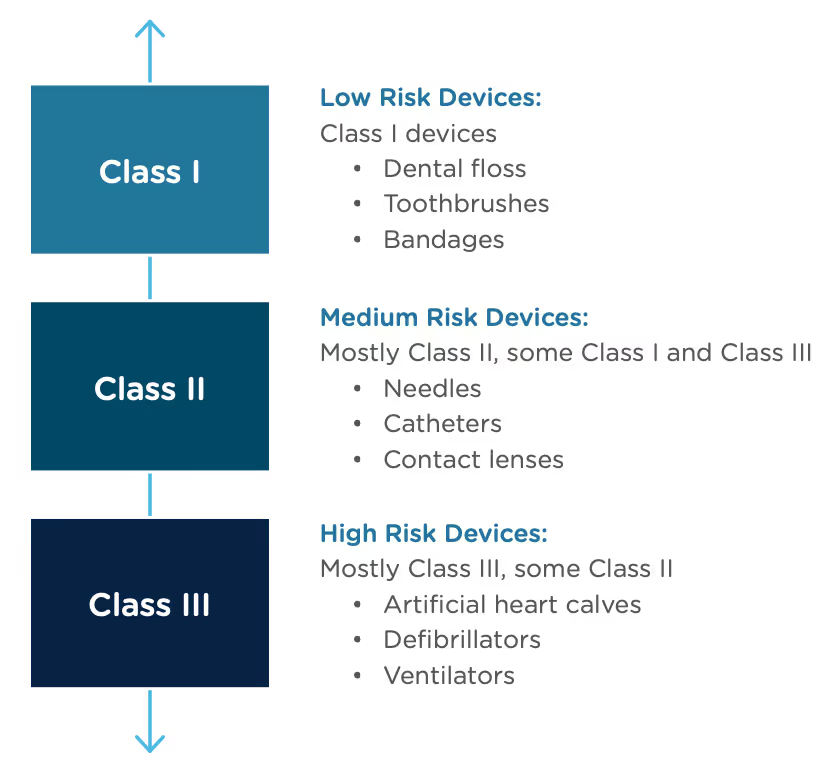
The Food and Drug Administration Modernization Act of 1996 provided the FDA with the authority to create the De Novo Classification Process. It's a process that uses a risk-based strategy for a new, novel kind of medical device, in vitro diagnostic, or medical software solution whose type has previously not been identified and/or classified. It’s a process by which a novel medical device can be classified as a Class I or Class II device, instead of being automatically classified as Class III, which may not be appropriate. Before the implementation of the De Novo process in 1997, all the “not substantially equivalent” (NSE) products were required to be initially classified as a Class III device. But for a lot of devices, this risk class didn’t really make sense. The De Novo process provides a pathway for more accurate classifications of novel, lower-risk devices.
October, 2021, the FDA released a final guidance document "De Novo Classification Process (Evaluation of Automatic Class III Designation)" to provide guidance to the requester (also known as the manufacturer) and the FDA on the process for the submission and review of a De Novo Classification Request under section 513(f)(2) of the Federal Food, Drug, and Cosmetic Act (the FD&C Act). This process provides a pathway to an initial Class I or Class II risk classification for medical devices for which general controls or general and special controls, provide a reasonable assurance of safety and effectiveness, but for which there is no legally marketed predicate device. This guidance document replaced the "New Section 513(f)(2) – Evaluation of Automatic Class III Designation, Guidance for Industry and CDRH Staff" document, dated February 19, 1998.
Consistent with the final rule, the FDA updated the guidance documents below to provide recommendations for submitting De Novo requests, as well as criteria and procedures for accepting, withdrawing, reviewing, and making decisions on De Novo requests, effective January 3, 2022.
- User Fees and Refunds for De Novo Classification Requests
- FDA and Industry Actions on De Novo Classification Requests: Effect on FDA Review clock and Goals
- Acceptance Review for De Novo Classification Requests
The 510(k) and the De Novo processes are similar in that they are both pathways to market for medical devices with low to moderate risk, which is Class I and Class II. The biggest difference between the two is that the 510(k) heavily relies on the concept of "substantial equivalence" to an existing medical device. You must prove this to get the clearance of your 510(k) submission. In the De Novo process, there isn’t a product currently on the market that is “substantially equivalent” to yours, so it’s like starting with a clean slate. For more on the 510(k) process, see our Beginner’s Guide to the 510(k) ebook.
A result of the De Novo process to be aware of is that a successful submission will lead to a new predicate device type that someone else can reference to bring their product to market through the 510(k) process. You’ve done all the work, so now it’s available for anyone to use to provide "substantial equivalence".
De Novo history/timeline
Preparing a De Novo request
1. Do your research! Be sure to complete all the necessary research prior to your submission. You want to be sure that your device is not substantially equivalent to an existing device. Resources to review include:
- The Center for Devices and Radiological Health (CDRH)
- U.S. FDA Device Classification Database
- Device Classification Under Section 513(f)(2)(De Novo)
2. A De Novo request can be submitted with or without a preceding 510(k). There are two options for when you can submit a De Novo request:
Option A: After receiving a not substantially equivalent (NSE) determination (that is, no predicate, new intended use, or different technological characteristics that raise different questions of safety and effectiveness) in response to a 510(k) submission.
Option B: If you’ve determined, after extensive research, that there is no legally marketed device on which to base a determination of substantial equivalence.
3. Be sure all fees are paid to the FDA in advance of submitting a De Novo request. The FDA’s fiscal year begins in October and runs through the following September. Fees have increased each year since they were introduced, but the FDA’s percentage of reviews completed within the 150-day window has increased as well.
A business that is qualified and certified as a “small business” is eligible for a substantial reduction in most of the FDA user fees, including De Novo. The CDRH is responsible for the Small Business Program that determines whether a business is qualified.
Medical Device User Fee Amendments (MDUFA) guidance documents can provide more detailed information about all FDA user fees.
4. The initial request process serves only to determine if the De Novo request is administratively acceptable based upon the Acceptance Checklist. The initial acceptance is followed by substantive review which will determine the final risk classification of your device.
5. A Pre-Submission (Pre-Sub) is a formal written request for feedback from the FDA that is provided in formal written form, and then followed by a meeting. Although a Pre-Sub is not required prior to a De Novo request, it can be extremely helpful to receive early feedback, especially for devices that have not previously been reviewed under a 510(k). If you think you would like to submit a pre-sub first, there are suggested guidelines for submission you should consider:
- Describe your rationale for a Class I or Class II classification for your device.
- Provide the search results of FDA public databases and other resources used to determine that no legally marketed device and no classification for the same device type exists.
- Provide a list of regulations and/or product codes that may be relevant.
- Provide a rationale for why the subject device does not fit within and/or is different from any identified classification regulations, based on available information.
- Identify each health risk associated with the device and the reason for each risk.
- Briefly describe any ongoing and/or planned protocols/studies that need to be completed in order to collect the necessary data to establish the device’s risk profile.
- Provide information regarding the safety and effectiveness of the device. Cite the types of valid scientific evidence you anticipate providing in your De Novo request, including types of data/studies relating to the device’s safety and effectiveness.
- Briefly describe any ongoing and/or planned protocols/studies that need to be completed to collect the necessary safety and effectiveness data.
- Provide protocols for non-clinical and clinical studies (if applicable), including how they will address the risks you anticipate and targeted performance levels that will demonstrate that general controls or general and special controls are sufficient to provide reasonable assurance of safety and effectiveness.
- Share any proposed mitigation measure(s)/control(s) for each risk, based on the best available information at the time of the submission. Highlight which mitigations are general controls and which are special controls and provide details on each.
- Include any other risks that may be applicable, in addition to those identified in the Pre-Sub, given the indications for use for the device.
- If applicable, provide any controls that should be considered to provide a reasonable assurance of safety and effectiveness for the device.
- Provide any non-clinical study protocols that are sufficient to allow the collection of data from which conclusions about device safety and/or effectiveness can be drawn. These protocols should address whether the identified level of concern is the appropriate level of concern for the device software, and if any additional biocompatibility and/or sterility testing is required.
- If clinical data is needed, provide information to show that the proposed study design and selected control groups are appropriate?
6. The FDA will attempt to review the De Novo request submission within 15 calendar days of receipt of the request to make a determination that the submission is declined or accepted for review. If they are unable to complete the review within the 15 days, your submission will automatically move to “accepted for review” status. https://www.fda.gov/regulatory-information/search-fda-guidance-documents/de-novo-classification-process-evaluation-automatic-class-iii-designation
7. There are times when the FDA will refund your application fee. They have created a guidance document “User Fees and Refunds for De Novo Classification Requests” for the purpose of identifying:
- the types of De Novo requests subject to user fees
- exceptions to user fees
- the actions that may result in refunds of user fees that have been paid
When is a De Novo request subject to a user fee?
When will the FDA refund a De Novo user fee?
What fee must be paid for a new device submission following a De Novo “decline” determination?
To continue reading this eBook including a detailed walk-through of all the Traditional 510(k) components, submission requirements and timelines, and an overview of the other 510(k) forms including the Abbreviated 510(k) and the Special 510(k), please register to download the full version.
Arena Solutions and Rimsys announce partnership to offer an end-to-end quality and product-centric regulatory solution
Foster City, Calif., January 12, 2021 – Arena Solutions, a leader providing cloud-based product development solutions for high tech, consumer electronics, and medical device industries, today announced a new partnership with Rimsys Inc., a world-leading provider of a holistic Regulatory Information Management (RIM) software platform designed specifically for medical technology (medtech) companies.
The Arena and Rimsys partnership offers a secure cloud-based, product-centric regulatory solution for the medtech industry. Rimsys seamlessly integrates with Arena’s QMS and PLM solutions by pulling product and documentation information directly into Rimsys to create, manage, and maintain marketing applications such as 510(k), Summary Technical Documentation (STED), and Table of Contents (ToCs).
The Arena product development platform connects product and quality processes allowing dispersed teams throughout the product design and manufacturing process to work together. Rimsys integrates with Arena’s platform by syncing product information so companies can better manage global registrations and selling status at the SKU level. Rimsys has the capability to pull in QMS records and documentation to create and compile regulatory applications from approved documentation. The integration automatically monitors for documentation changes and alerts users when updates occur with additional reporting based on document location ensuring a single and accurate source of truth.
"The seamless and deep integration between Rimsys and Arena solutions reduces the day-to-day regulatory management," said James Gianoutsos, Founder and President of Rimsys. "The administrative burden of compiling marketing applications and the maintenance of product data is completely eliminated, allowing for increased compliance, efficiency, and visibility throughout the organization."
"Our partnership with Rimsys makes it easier for MedTech companies to address regulatory affairs, product registration, and standards management more effectively," said George Lewis, VP of Business Development and Strategy for Arena Solutions. "This new integration streamlines regulatory compliance processes by accelerating the notification of updates to critical quality records and documents."
About Arena Solutions
Arena Solutions helps innovative electronic high tech and medical device companies create products that change the world. Arena unifies product lifecycle (PLM) and quality management (QMS) processes, allowing every participant throughout the product realization process from design to manufacturing to work together. With Arena, teams accelerate product development and delivery to increase profits. For more information, visit ArenaSolutions.com.
About Rimsys
Rimsys is a world-leading provider of Regulatory Information Management (RIM) software for medical technology companies. Built by and for regulatory affairs professionals, Rimsys digitizes, automates, and creates regulatory order to ensure products adhere to changing global regulations. It is the only holistic RIM software for medical devices, in-vitro diagnostics, and medical device software that makes it easy to manage global UDI requirements and navigate the pillars of regulatory affairs, including product registration, standards management, essential principles/GSPR, and regulatory intelligence. rimsys.io

Rimsys releases new automated unique device identification (UDI) module for its holistic medtech RIM platform
Rimsys, a world-leading Regulatory Information Management (RIM) software platform for medical technology companies, announced the release of an innovative and automated solution to address the global Unique Device Identification (UDI) requirements. This enhancement to the Rimsys ecosystem is immediately available to new and existing customers, providing a compliant method for tracking and traceability of UDI data that is fully compatible with global health authority databases.
UDI is a global harmonization effort to provide more consistent data that benefits overall product traceability throughout the medical device supply chain. It aims to adequately identify medical devices from manufacturing through distribution to patient use. Because global markets have different UDI requirements, product data can become a maintenance nightmare, even for the most mature medical technology companies. Data management is further complicated as UDI data is not static and must be updated based on changing regulatory requirements, such as market dates and registration status.
"Rimsys is the first automated UDI solution to fully integrate into a product’s registration lifecycle, ensuring that changing regulatory information is properly captured, registered, and maintained with the product and within government UDI databases like the GUDID (USA) and EUDAMED (EU)," said James Gianoutsos, Founder & President at Rimsys, "The current solutions landscape revolves around creating in-house, custom, and expensive projects with custom-built software or existing ERP software that were not designed to keep up with the fast-paced and ever-changing regulatory landscape."
In order to stay compliant and competitive, medical technology companies need to adapt quickly and flawlessly to the regulatory environment. As UDI requirements are rolled out across the globe, Rimsys’ UDI module seamlessly integrates with product registration data that is already actively managed within Rimsys, resulting in less maintenance, reduced costs, better compliance, and ultimately decreased time-to-market.
"We have already seen how our system has helped world-leading medical technology companies, such as Johnson & Johnson, Omron, and Terumo, digitize, automate, and navigate the global regulatory landscape,” said Brad Ryba, Chief Technology Officer at Rimsys, "As these requirements and demands continue to evolve, we are committed to ensuring that regulatory professionals can leverage the data integrations within Rimsys to more proactively manage their regulatory information."
Interested customers can schedule a preview of the new UDI module.

Rimsys named 2020 top technology company finalist by Tech 50 awards
Rimsys has been named a Finalist in Tech 50’s 2020 awards program hosted by the Pittsburgh Technology Council. The awards program, which was held November 12, 2020 at a virtual gala event, announced the winners and finalists. Rimsys was nominated as Innovator of the Year in the MedTech category, along with Sentact, Philips, and TeleTracking Technologies. In addition to the Company’s nomination, Rimsys’ Founder and CEO, James Gianoutsos, was nominated for CEO of the Year.
The awards program honors technology innovation in the region. Rimsys Inc., provider of the only holistic Regulatory Information Management (RIM) software for medtech is headquartered in Pittsburgh, Pennsylvania and was proud to be among the 2020 nominees. Rimsys was named a finalist this year, as Sentact, a provider of healthcare technology solutions for the patient care journey, walked away as the winner. Barbara VanKirk, CEO of IQ Inc., won this year’s CEO of the Year award.
"Rimsys is honored to be a finalist in both categories. Thank you to the Pittsburgh Technology Council for highlighting all of these innovative companies, and congratulations to this year’s winners," said Gianoutsos, "Rimsys is already used and trusted by leading companies, including Johnson & Johnson and Terumo, and we look forward to the near future where Rimsys’ automation of the global regulatory landscape is commonplace for medical technology companies and regulatory affairs professionals."
James Gianoutsos founded Rimsys in 2017, recognizing that while regulatory management platforms existed for pharmaceutical companies, medical technology companies face an entirely different set of challenges and requirements. Equipped with the expertise and drive to give regulatory affairs professionals a better way, he developed Rimsys: a RIM software built 100% for medtech companies.
Rimsys consolidates all the major functions of regulatory affairs on a 100% secure, cloud-based software, making product registration, standards management, essential principles/GSPR, and regulatory intelligence easy. Its newest integrated module manages global UDI requirements.
"It’s been an unparalleled year with unthinkable challenges," said Audrey Russo, President & CEO of the Pittsburgh Technology Council, "The COVID-19 Pandemic has tested everyone’s mettle in ways never imagined. This year’s Tech 50 is a testament to the fortitude, creativity, and innovation that exists across our industry, membership, and in every Tech 50 winner and finalist."
For a full list of finalists and winners, visit the Pittsburgh Technology Council’s website.
Rimsys secures $1.5 million to accelerate growth of its leading medtech regulatory information management platform
Rimsys Inc., provider of the leading Regulatory Information Management (“RIM”) software designed specifically for medical technology (“medtech”) companies, announced today it closed a $1.5 million investment round, led by Allos Ventures. The financing round will support Rimsys’ penetration of the medtech RIM market through planned expansions of its product offering, sales, and marketing execution. Rimsys serves an expanding list of enterprise customers, including industry leaders such as Johnson & Johnson and Terumo.
"Complex regulatory challenges create barriers that delay product time-to-market, stall revenue growth and increase exposure to compliance risks for even the most experienced medtech companies," said James Gianoutsos, founder & president of Rimsys. "Regulatory teams are in need of digital solutions that provide more efficient and compliant ways to stay on top of a constantly changing regulatory environment"
Rimsys’ solution seamlessly integrates with medtech manufacturers’ existing quality management systems, product lifecycle management systems, and sales and distribution software systems. Rimsys’ robust digital platform, with its intuitive user interface and global intelligence, enables its customers to meet market entrance requirements and grow internationally. John McIlwraith, managing director at Allos Ventures will join Rimsys’ board of directors.
"The maze of global regulations covering medical technology is growing more complex and frequently results in dead ends and delays that can greatly impact commercialization of these products," said McIlwraith. "This financing round will enable the company to further accelerate its momentum with large enterprise customers."
For more information on Rimsys, please visit www.rimsys.io.
Rimsys announces new brand identity for world-leading regulatory information management (RIM) software
Rimsys, a world-leading Regulatory Information Management (RIM) software platform for medical technology companies, announced a new brand identity and tagline. Rimsys, the only holistic RIM software on the market designed specifically for medical technology companies, with functionality for the pillars of regulatory affairs, rebranded to accurately reflect its novel and advanced offering through a distinct, modern brand identity and messaging platform.
The new visual identity, including logo, color palette, graphic elements, and iconography mirror the trusted, relatable, and modern personality of Rimsys. The colors, which feature a vibrant purple and dark blue, were strategically chosen to differentiate Rimsys from other RIM solutions on the market. The visuals are accompanied by a new mission statement, “to digitize, automate, and create regulatory order for the medical technology industry,” and messaging platform that conveys its value propositions:
- Used and trusted by the world’s leading medtech companies
- Built by and for regulatory affairs professionals in medtech
- Easy to start and simple to scale as your company grows
"We’re pleased to announce this rebrand to our customers and partners, with complete confidence that the new identity and messaging accurately reflect our enterprise software solution," said James Gianoutsos, Founder & President at Rimsys.
Rimsys consolidates all the major functions of regulatory affairs on a 100% secure, cloud-based software, making product registration, standards management, essential principles/GSPR, and regulatory intelligence easy. These functions, plus soon-to-be-released features, were formally messaged in the rebrand, including superior server security, system compliance, and simple integrations with many leading enterprise QMS and ERP/PLM software systems.
"With successful implementations in some of the world’s leading medical technology companies including Johnson & Johnson and Terumo, we already know Rimsys has the power to modernize medtech companies of all sizes and scales," said Gianoutsos. "And now, with a brand that mirrors our capabilities, we’re even better poised to reach out and help more."
Components of the rebrand can be seen on rimsys.io, and the full rebrand will be revealed on a new website that is currently under development.

Announcing the release of Rimsys 3.0: Rimsys Insight
Rimsys Regulatory Management Software, the leading and only Regulatory Information Management (RIM) system platform designed specifically for the medtech industry, has just released Rimsys 3.0, which includes its much anticipated regulatory intelligence module, Rimsys Insight.
Rimsys 3.0 now offers curated regulatory news along with changes to laws, regulations, and guidances that are timely and linked to customers’ specific products, markets and the other customizable filters available to each company. Additionally, the integration of market entrance requirements for the top 50+ global markets, further strengthen the already powerful Rimsys Registrations workflow which manages medical device and IVD registrations at the SKU level.
"Information is meaningless unless you can apply context to it as it relates to your company and products. As regulatory professionals ourselves, we know the specific pain that is experienced by our customers. The user experience should be seamless, fully integrated, and intuitive enough that, regardless of how long that person has been in the industry"
Rimsys was built for and by regulatory affairs professionals, enabling teams to digitize and automate otherwise disjointed, paper-based and manual processes. Rimsys has spent the last year working with customers from some of the world’s leading medical device manufacturers to develop the requirements that seamlessly integrate their business processes with its existing ecosystem of cloud-based regulatory affairs software solutions. This includes global product registrations, submissions management, standards management, essential principles and more.
"There’s simply no other holistic and vertically integrated solution quite like Rimsys out in the marketplace. Our initial features were focused on bringing MedTech companies together to collaborate and connect with each other internally. Built on this solid foundation, our latest release truly empowers and connects them to the entire global regulatory ecosystem"
Rimsys is a holistic approach to modernizing, digitizing and automating regulatory affairs activities for the MedTech industry by providing a single source of truth. Rimsys is now aligned and positioned to continue its growth and mission to "digitize, automate and createregulatory order for the medical technology industry™."