Announcing AI-Native Market Access Product for Regulated Products

Learn More

Blogs

MedTech

FDA PMA submission process: a beginner's guide

By

Wendy Levine

April 27, 2022

4 min read

This article is an excerpt from The beginner's guide to the FDA PMA submission process ebook.

Table of Contents

Introduction

If your organization is planning to market a new medical device in the United States, you first need to determine which regulatory class the device falls under. The vast majority of medical devices regulated by the FDA are either Class I or Class II medical devices, requiring a 510(k) premarket notification or a simple registration if exempt from 510(k) requirements. However, if your device sustains or supports life, is implanted, or presents a “potential unreasonable risk of illness or injury,” your device is likely a Class III device which will require Premarket Approval (PMA) from the FDA before it can be marketed in the United States. Novel devices, for which there are no existing substantially equivalent devices, are automatically classified as Class III as well. Novel devices with a lower risk profile, however, may qualify for the De Novo process instead of the PMA. Just 10% of devices regulated by the FDA are Class III devices.

This ebook provides an overview of the PMA process and its requirements, but it is not designed to be the only resource used in compiling a PMA submission. The FDA provides significant documentation on this process, starting with the regulation governing premarket approval that is located in Title 21 Code of Federal Regulations (CFR) Part 814.

Chapter 1: PMA Basics

FDA: Background and device oversight 

Before we explain what a PMA is, let’s first talk generally about the Food and Drug Administration (FDA) and device oversight. The FDA is the U.S. governmental agency responsible for overseeing medical devices, drugs, food, and tobacco products. When it comes to medical devices, the FDA’s mission is to “protect the public health by ensuring the safety, efficacy, and security of...medical devices.” At the same time, the FDA also has an interest in “advancing public health by helping to speed innovations.” In other words, the FDA’s goal is to make sure devices are safe and effective for public use, while also ensuring that devices have a quick and efficient path to market.

In order to achieve this balance of safety and efficiency, the FDA has three different levels of oversight depending on the risk level of the device: (1) exempt from premarket notification, (2) Premarket Notification, also known as 510(k), and (3) Premarket Approval (PMA). 
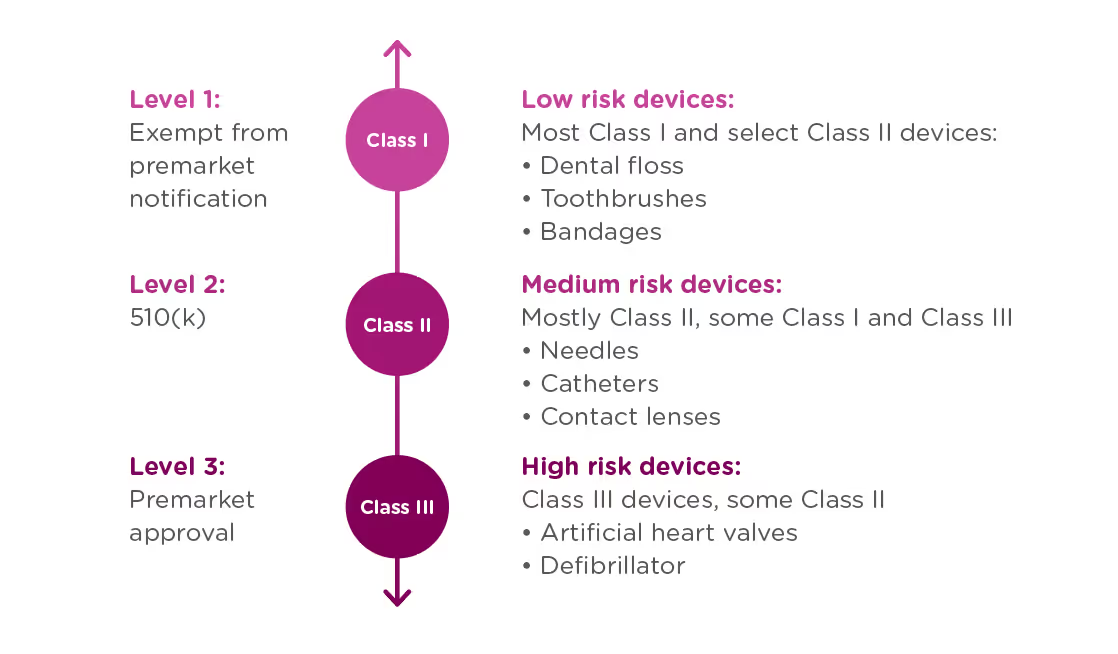
PMA submissions - medical device classes

When is a PMA required?

The PMA process is the most stringent regulatory process for medical device approval under the FDA and applies to almost all Class III devices. To determine whether your device requires a PMA, you must first Classify your device by searching the Product Classification Database. The database will provide you with similar devices; their name, classification, and link to the Code of Federal Regulations (CFR) if applicable.

  • If a substantial equivalent is found in the Product Classification Database with a submission type of 510(k), you should submit a 510(k), not a PMA.
  • If the product classification database identifies your device as Class III and/or requiring a PMA - you should submit a PMA.
  • If your device involves a new concept and does not have a classification regulation in the CFR, the database will list only the device type name and product code. In this case, the three-letter product code can be used to search the PMA database and the 510(k). 
  • If  your device cannot be found in the product classification database because it is a new type of device and should be classified as a Class III device because of the level of risk it presents*.

Class III devices support or sustain human life, are of substantial importance in preventing impairment of human health, or present a potential and unreasonable risk of illness or injury.

Note that if your device is a new concept without a substantial equivalent, but does not present the level of risk of a class III device, it may be eligible for the De Novo process as a class I or class II device.

PMA vs 510(k)

Not only are PMA and 510(k) processes applicable to different types of devices, they have different purposes.

510(k): A 510(k) is intended to demonstrate that the device for which approval is being sought is as safe and effective as a currently marketed device that does not require a PMA.

PMA: A PMA is intended to prove that a new device is safe and effective for the end user. A PMA is much more detailed and in-depth than a 510(k). Device manufacturers are typically required to present human clinical trial data, in addition to laboratory testing data.

The difference in complexity between a PMA and 510(k) also affects the time needed to process the submissions. The FDA typically accepts or rejects a 510(k) submission within 30-90 days, at which point the device is posted to the FDA’s 510(k) database. A PMA submission can take up to 180 days to be processed, at which point the FDA can approve or deny the application. The FDA may also issue an “approvable” or “not approvable” letter, which the applicant can choose to respond to, thereby adding time to the submission process. 

PMA application methods

There are a number of types of PMA application methods. While most devices which require a PMA will follow the traditional process, be sure to verify that you are using the correct application process to maximize your chances for success and avoid unnecessary delays:

Traditional PMA

The most common method for attaining FDA clearance for Class III devices, the traditional PMA is the appropriate option for most devices that have completed clinical testing. 

Modular PMA

The modular PMA is the appropriate application method for devices that have not yet completed clinical testing. Applicants complete individual “modules,” with final confirmation granted once all sections are completed. For additional information on specific requirements of a modular PMA, read the FDA’s Premarket Approval Application Modular Review.

Product Development Protocol

Use the Product Development Protocol (PDP) with medical devices that are based on well-established technology. The PDP process for gaining market approval merges the clinical evaluation and development of information, and involves an agreement between the manufacturer and the FDA. The process provides the advantage of early predictability for the manufacturer and allows early interaction that can identifyFDA concerns as soon as possible in the development process. Because the PDP identifies the agreed upon design and development details, a completed PDP is considered to have an approved PMA. For additional information, read more about the FDA’s PMA Application Methods.

Humanitarian Device Exemption

A Humanitarian Use Device (HUD) is specifically defined as a device intended to benefit patients that are affected by a disease or condition that affects less than 8,000 individuals in the U.S. per year. TheHumanitarian Device Exemption (HDE) approval process is designed to encourage clinical activity around rare conditions, and does have certain restrictions, including:

  • After receiving HDE approval, a HUD is eligible to be sold for profit only if the device is intended to address a disease or condition that occurs primarily in pediatric patients, or occurs in pediatric patients in small numbers.
  • If an HDE is approved to be sold for profit, the FDA will determine an annual distribution number(ADN). Any devices sold beyond the ADN limit are required to be sold for no profit.

For more information see the FDA’s explanation of the Humanitarian Device Exemption.

CBER Submissions

There are two centers within the FDA responsible for evaluating medical devices. While the majority of devices will go through the Center for Devices and Radiological Health (CDRH), some will be managed by The Center for Biologics Evaluation and Research (CBER). CBER regulates medical devices related to blood and cellular products, including blood collection and processing procedures as well as cellular therapies. This ebook focuses on submissions made through the CDRH, but you can view CBER Regulatory Submissions – Electronic and Paper for more information on the CBER process.

Chapter 2: FDA Interactions

To continue reading this eBook, including a walk through of the different types of required and optional FDA meetings and communications, a detailed list of the contents of a traditional PMA submission, and an overview of quality management system requirements, please register to download the full version.

Similar posts

Company

Product Updates

Rimsys Announces AI-Native Market Access for Regulated Products

By

Bethaney Lentz

September 1, 2026

4 min read

New software gives regulatory and commercial leaders a source-linked, actionable view of market viability, requirements, timing, and risk before committing resources

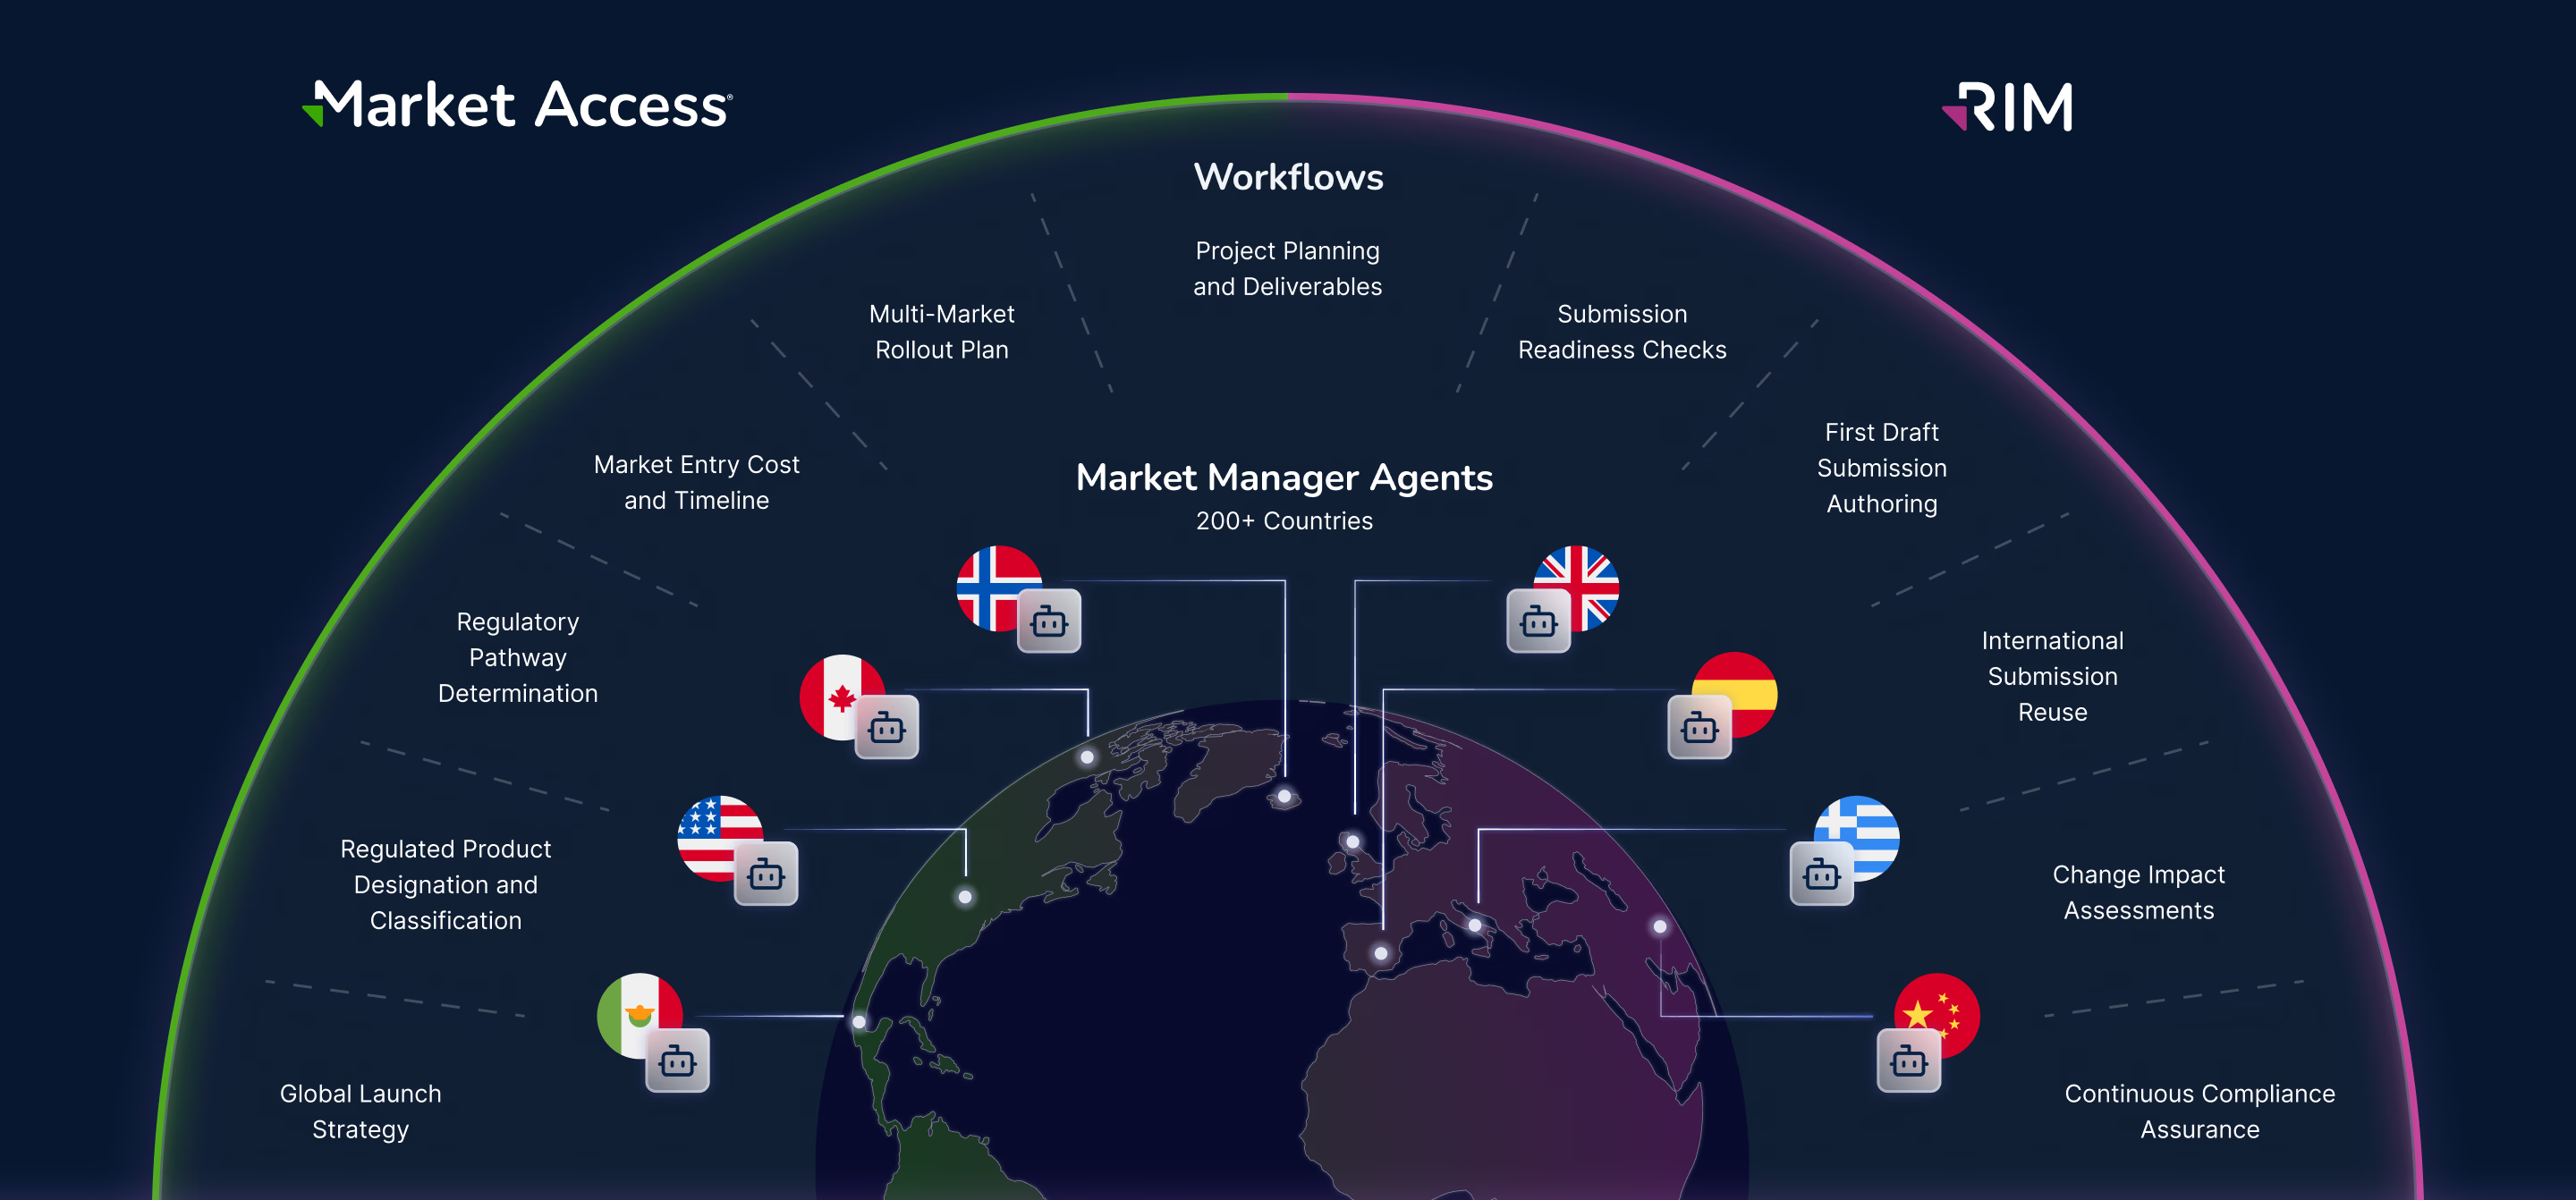
PITTSBURGH, PA, September 1, 2026 - Rimsys, the heart of medtech regulatory operations, today announced Market Access, a new AI-native product designed to help companies determine whether a market is worth entering and what it will take to get there. Market Access will be available in beta beginning September 1, 2026, with general availability to be announced separately.

Market Access replaces what can be a 6-to-9-month research and assessment process with a same-day, actionable read on market viability. It gives regulatory and commercial leaders the classification, pathway, timeline, cost considerations, and potential regulatory gaps they need to make a more defensible investment decision before committing budget and resources.

Predictability is at the core of Market Access. Rather than simply identifying a regulatory pathway, it helps teams understand what to expect, which requirements or gaps are most likely to slow down a submission, and where uncertainty remains. Every answer traces back to the specific regulation, guidance, or law behind it, allowing regulatory professionals to verify the underlying source rather than simply trust an AI-generated response.

Key features and benefits include:

  • Same-day market assessment: Provides an actionable read on whether a market is worth entering, replacing research and assessment that can take 6 to 9 months.
  • Cross-designation classification: Evaluates products across medical device, IVD, cosmetic, consumer, and OTC designations, so the same product receives the appropriate regulatory read regardless of how a given market classifies it.
  • Clear market-entry requirements: Provides classification, regulatory pathway, timeline, cost considerations, and key requirements before teams commit resources.
  • Earlier visibility into regulatory gaps: Surfaces the specific requirements and gaps most likely to slow down a submission and helps teams understand their potential severity.
  • Greater predictability: Provides visibility into anticipated regulatory response windows, risks, and areas of uncertainty so teams can better plan for what lies ahead.
  • Source-linked intelligence: Traces answers to the underlying regulations, guidance, and laws so regulatory professionals can verify the basis for an assessment.
  • AI with human oversight: Uses Rimsys AI and Market Manager Agents to accelerate complex regulatory research while keeping regulatory professionals in control of review and validation.
  • Connected planning and execution: Complements RIM, connecting market-entry decisions with the registrations, submissions, regulatory changes, and ongoing compliance required to execute them.

“Regulatory knowledge has traditionally been highly dependent on individual expertise and market-by-market research,” said James Gianoutsos, Founder and CEO of Rimsys. “Market Access gives companies greater predictability before they make a market investment. Teams can understand whether an opportunity is worth pursuing, what it will take to enter, where regulatory challenges are likely to emerge, and what uncertainty remains. And because every answer is tied to its regulatory source, experts can verify the recommendation rather than simply trust the AI.”

Market Access is powered by Rimsys AI, which works from a curated regulatory corpus spanning more than 200 markets, not the open web. Its Market Manager Agents are AI agents with market-specific regulatory knowledge, including pathways, requirements, and documentation, that support market assessments and global launch planning. The product is also designed to acknowledge areas where reviewer interpretation and expert judgment can affect regulatory outcomes rather than presenting those outcomes as more predictable than they are.

Market Access expands the Rimsys product portfolio from regulatory execution into market planning. Market Access helps regulatory and commercial leaders determine where to compete, while RIM manages the registrations, submissions, regulatory changes, product data, and ongoing compliance required to enter and remain in those markets. Together, they turn regulatory intelligence into coordinated action, helping regulatory teams accomplish more in every hour while giving business leaders greater visibility into opportunities, risk, and the work required to protect and grow revenue.

Beta Availability

Market Access will be available in beta beginning September 1, 2026. During the beta period, Rimsys will continue validating product capabilities and refining workflows ahead of broader commercial availability. 

Organizations interested in learning more about Market Access or participating in the beta program can visit rimsys.io/products/market-access.

About Rimsys

Rimsys is the heart of regulatory operations, connecting AI-native market planning with AI-embedded regulatory execution, giving medtech manufacturers one living system from market-entry decision to execution. 

Rimsys Market Access is the planning layer, answering not just whether a market can be entered, but whether it should be, returning classification, pathway, timeline, and cost before a team commits budget or resources to finding out. Rimsys RIM is the execution layer, managing registrations, submissions, regulatory changes, and compliance so teams can act on that plan with confidence.

Trusted by 6 of the top 12 global MedTech manufacturers and certified to ISO 27001 and SOC 2, with ISO 42001 certification for AI management systems underway, Rimsys keeps regulatory intelligence, product data, approvals, and change management continuously connected so teams can enter new markets with speed and assurance. Learn more at rimsys.io.

Media Contact
letschat@rimsys.io

Registration deadlines don't have to be a risk. See how centralized data, automated alerts, and standardized workflows keep global registrations on track

MedTech

RIM

Product Updates

How to Prevent Registration Deadline Misses in 2026

By

Bethaney Lentz

August 4, 2026

4 min read

A missed registration deadline can halt sales, trigger regulatory penalties, and damage your organization's credibility with health authorities. For MedTech regulatory affairs teams managing hundreds or thousands of product registrations across dozens of markets, preventing deadline misses requires more than calendar reminders. It demands structured data, clear workflows, and real-time visibility into expiration timelines.

This guide explains why registration compliance deadlines are missed and how your team can prevent delays through stronger data control, workflow visibility, and process standardization.

Key Takeaways: How to Prevent Registration Deadline Misses in 2026

  • Registration deadline misses typically stem from fragmented data, unclear ownership, and siloed communication between teams.
  • Proactive renewal management requires real-time visibility into expiration dates across your entire global product portfolio.
  • Standardized workflows with assigned accountability eliminate the confusion that leads to overlooked deadlines.
  • Rimsys centralizes registration data and automates renewal alerts, reducing the risk of missed deadlines by up to 90%.
  • Investing in a purpose-built RIM platform protects revenue continuity and maintains uninterrupted market access.

Why Do MedTech Teams Miss Registration Compliance Deadlines?

Registration deadline misses  occur because the operating model cannot scale with growing portfolio complexity. Understanding root causes is the first step toward prevention.

Fragmented Data Across Systems and Regions

Many organizations track registrations using disconnected regional trackers, shared drives, and individual team member records. When product data lives in multiple locations, no single source of truth exists. This fragmentation makes it difficult to identify upcoming expirations before they become urgent.

A registration that expires in Brazil may be tracked in one system while the same product's EU approval is monitored elsewhere. Without connection between these records, teams lack portfolio-wide visibility into renewal timelines.

Unclear Ownership and Accountability

When responsibilities are distributed across regional leads, in-country partners, and distributors without clear accountability structures, deadlines fall through the cracks. Team members may assume someone else is handling a renewal, leading to inaction until the deadline passes.

Organizations with high turnover face additional risk. When key personnel leave, institutional knowledge about pending renewals often leaves with them.

Reactive Tracking Instead of Proactive Management

Calendar-based reminders and email notifications are inherently reactive. By the time a reminder appears, teams may not have enough runway to gather required documentation, coordinate with authorities, or address unexpected complications.

Effective deadline management requires proactive monitoring that surfaces upcoming expirations months in advance, not days before they occur.

What Happens When Registration Deadlines Are Missed?

The consequences of missed registration deadlines extend beyond administrative inconvenience. They create tangible business disruptions that affect revenue, reputation, and regulatory standing.

Loss of Market Access and Revenue Disruption

An expired registration means your product cannot legally be sold in that market. Regulatory teams must halt shipments, distribution partners cannot fulfill orders, and revenue stops flowing. For high-volume products or critical markets, even brief interruptions to the supply chain can translate to significant financial impact.

Re-registering a lapsed product often takes longer than a standard renewal. Your team may face additional scrutiny, updated documentation requirements, or queue delays that extend the time to market restoration.

Regulatory Penalties and Increased Scrutiny

Health authorities track compliance history. Missed deadlines signal operational weaknesses that can trigger increased inspection frequency, additional documentation requests, or enhanced scrutiny on future submissions. Building trust with regulators takes years; eroding it takes one avoidable miss.

Damage to Distributor and Partner Relationships

In-country partners and distributors depend on your organization to maintain valid registrations. When market access lapses, partners bear the burden of explaining delays to healthcare providers and end customers. Repeated misses can damage long-term business relationships and competitive positioning.

How to Identify Products at Risk of Deadline Misses

Prevention starts with identifying which registrations are most vulnerable. A systematic risk assessment helps teams prioritize attention and allocate resources effectively.

Create a Consolidated Registration Inventory

Bring all registration data into a single view. Include product identifiers, registration numbers, approval dates, expiration dates, responsible parties, and market-specific requirements. This inventory becomes your foundation for risk assessment.

Rimsys Registration Software centralizes this data automatically, linking registrations to products, submissions, and regulatory intelligence in one structured system.

Segment by Expiration Timeline and Complexity

Not all renewals carry equal risk. Categorize registrations by time remaining until expiration and by the complexity of renewal requirements. Products expiring within the next 90 days with extensive documentation needs should receive immediate attention.

Consider country-specific factors. Some markets require local testing, updated clinical data, or notified body involvement. These requirements extend lead times and increase the risk of delays.

Assess Documentation Readiness

For each at-risk registration, evaluate whether required documentation is current and accessible. Outdated technical files, missing test reports, or incomplete labeling information create bottlenecks that delay renewal submissions.

Documentation gaps discovered weeks before a deadline often cannot be addressed in time. Early assessment ensures enough runway to resolve issues.

Building a Proactive Renewal Management System

Moving from reactive tracking to proactive management requires intentional process design. The following components form the foundation of an effective renewal management system.

Establish Clear Ownership for Every Registration

Assign a single accountable owner for each registration. This person is responsible for monitoring expiration timelines, coordinating renewal activities, and escalating issues that require additional support.

Ownership should be documented within your registration management system, not stored in separate organizational charts or email threads. When someone leaves the organization, ownership transfer becomes straightforward.

Define Standard Lead Times by Market and Product Type

Different markets have different renewal timelines. EU MDR renewals involve notified body coordination. FDA establishment registrations follow annual cycles. ANVISA submissions may require specific local documentation.

Document the standard lead time needed for each market and product type. Use these lead times to trigger renewal workflows well before deadlines approach.

Implement Automated Expiration Monitoring

Replace manual calendar tracking with automated monitoring that surfaces upcoming expirations across your entire portfolio. Alerts should trigger at multiple intervals, such as 180 days, 90 days, 60 days, and 30 days before expiration.

Automated monitoring ensures that no registration is overlooked, regardless of portfolio size or team bandwidth. Rimsys Global Regulatory Compliance capabilities include lifecycle tracking and automated alerts that reduce missed renewal risk.

Standardizing Workflows to Prevent Oversight

Consistent workflows eliminate the variability that leads to deadlines. When every renewal follows the same structured process, teams can identify and address deviations before they cause problems.

Create Renewal Workflow Templates

Define the steps required for each type of renewal. Include documentation gathering, internal review, submission preparation, authority submission, and post-submission tracking. Assign responsible parties and expected durations for each step.

Templates ensure that new team members can execute renewals correctly without relying on undocumented institutional knowledge.

Build Escalation Paths for Delayed Activities

Not every renewal proceeds according to plan. Documentation may be delayed. Authority responses may take longer than expected. Partners may fail to submit required local information.

Define clear escalation paths that activate when activities fall behind schedule. Escalation should be automatic and visible, not dependent on individual team members raising concerns manually.

Conduct Regular Pipeline Reviews

Schedule recurring reviews of your renewal pipeline. During these reviews, examine registrations approaching expiration, identify any blocked activities, and confirm that responsible parties are on track.

Pipeline reviews surface potential issues early, when corrective action is still possible. They also reinforce accountability by making renewal status visible to leadership.

How Data Control Reduces Deadline Risk

Strong data control is the foundation of reliable deadline management. When registration data is accurate, current, and accessible, teams make better decisions and avoid preventable misses.

Maintain a Single Source of Truth

Eliminate competing data sources by establishing one authoritative system for registration information. All team members, partners, and stakeholders should access the same data, ensuring consistency and reducing confusion.

A single source of truth means that when someone asks about a product's registration status in a specific market, the answer is immediate and reliable. No cross-referencing multiple trackers. No waiting for colleagues to respond to email inquiries.

Connect Registration Data to Product and Submission Records

Registrations do not exist in isolation. They connect to specific products, rely on previous submissions, and may be affected by regulatory changes or product modifications. Systems that link these data types enable more accurate impact analysis.

When a product design change occurs, connected data helps teams identify which registrations may need updated. When regulations evolve, linked intelligence shows which markets and products are affected.

Ensure Version Control and Audit Readiness

Regulatory authorities expect organizations to demonstrate controlled processes. Maintain version history for all registration-related documentation. Track who made changes, when changes occurred, and what was modified.

Audit readiness built into daily operations means less scrambling when inspections occur. It also shows the historical context needed to understand how registration status evolved over time.

Leveraging Technology for Visibility and Automation

Technology plays a critical role in scaling deadline management across large portfolios. Purpose-built regulatory information management platforms offer capabilities that generic tools cannot match.

Dashboard Visibility Across Global Markets

Executive and operational dashboards display at-a-glance visibility into registration status across your entire portfolio. Filter by market, product line, expiration timeline, or responsible party, to focus on relevant subsets.

Dashboard visibility enables leadership to ask and answer questions quickly:

  • How many registrations will expire in the next quarter?
  • Which markets have the highest concentration of upcoming renewals?
  • Are any renewals currently blocked?

Automated Alerts and Notifications

Configure alerts that notify responsible parties when action is required. Notifications should be specific, actionable, and timely. Rather than generic reminders, alerts should identify the specific registration, required action, and deadline.

Effective notification systems reduce the cognitive burden on team members. Instead of tracking deadlines mentally, they receive prompts when attention is needed.

Reporting for Continuous Improvement

Track metrics that indicate process health. Monitor the number of registrations renewed on time, average lead time utilization, and frequency of expedited renewals. Use these metrics to identify process weaknesses and drive improvement.

Organizations that measure renewal performance can demonstrate compliance maturity to regulators, partners, and investors. Metrics also help justify investments in process improvement.

How Rimsys Helps Prevent Registration Deadline Misses

Rimsys is the first and only holistic Regulatory Information Management software purpose-built for MedTech. It centralizes registrations, submissions, regulatory intelligence, and UDI data in one connected platform, giving teams the visibility and automation needed to prevent deadline misses.

Centralized Global Registration Tracking

Rimsys maintains a structured, product-centric data model that connects registrations to products, markets, and submissions. Teams gain real-time visibility into where every product can be sold, which registrations are pending, and which are approaching expiration.

Six of the world's top 12 MedTech manufacturers trust Rimsys to manage global regulatory operations, achieving up to 90% reduction in reporting effort and elimination of manual tracking risk.

Automated Lifecycle Alerts

Rimsys monitors registration lifecycles and automatically alerts teams to upcoming expirations, renewals, and information requests. Automated alerts ensure that no registration is overlooked, regardless of the portfolio size.

With Rimsys AI, teams can accelerate regulatory work through AI-assisted workflows that reduce repetitive tasks while keeping human judgment in control.

Connected Regulatory Intelligence

Registration deadlines do not exist in isolation from regulatory change. New requirements can affect renewal timelines, documentation needs, or market access conditions.

Rimsys connects registration data to regulatory intelligence and impact assessment workflows, helping teams understand how changes affect their portfolio before deadlines become urgent.

Creating a Culture of Deadline Accountability

Technology and process alone do not prevent deadline misses. Organizations must also cultivate a culture where deadline accountability is valued and reinforced.

Make Renewal Performance Visible

Share renewal metrics with teams and leadership regularly. Celebrate on-time renewals and analyze near-misses to identify improvement opportunities. Visibility creates accountability without requiring punitive measures.

Invest in Team Training and Development

Ensure that team members understand the importance of deadline management and have the skills to execute renewal workflows effectively. Training should cover both procedural requirements and the business impact of missed deadlines.

Support Cross-Functional Collaboration

Registrations often require input from quality, engineering, labeling, and commercial teams. Foster collaboration across functions to ensure that dependencies are identified early and addressed proactively.

When regulatory affairs teams operate in silos, they lack the information needed to anticipate complications. Cross-functional visibility enables earlier intervention.

In Conclusion: How to Prevent Registration Deadline Misses

Preventing registration deadline misses requires intentional effort across three dimensions: data control, workflow visibility, and process standardization. Organizations that invest in these areas protect revenue continuity, maintain regulatory standing, and preserve partner relationships.

The complexity of global regulatory operations is not slowing down. As portfolios expand and requirements evolve, teams need infrastructure that scales. Rimsys transforms regulatory operations from administrative tracking into strategic enablement, bringing speed, visibility, and confidence to global expansion.

Ready to eliminate deadline risk from your registration management? Speak with the Rimsys team to see how leading MedTech manufacturers manage global registrations with precision and control.

References:

How Smith & Nephew Repositioned Regulatory as a Strategic Commercial Partner

MedTech

RIM

How Smith & Nephew Repositioned Regulatory as a Strategic Commercial Partner

By

Caroline La

May 28, 2026

4 min read

Smith & Nephew is a global medical device manufacturerwith a broad portfolio spanning orthopedics, sports medicine, and woundmanagement, sold and registered across markets worldwide. Before Rimsys,regulatory data was scattered across spreadsheets, shared drives, anddisconnected systems.

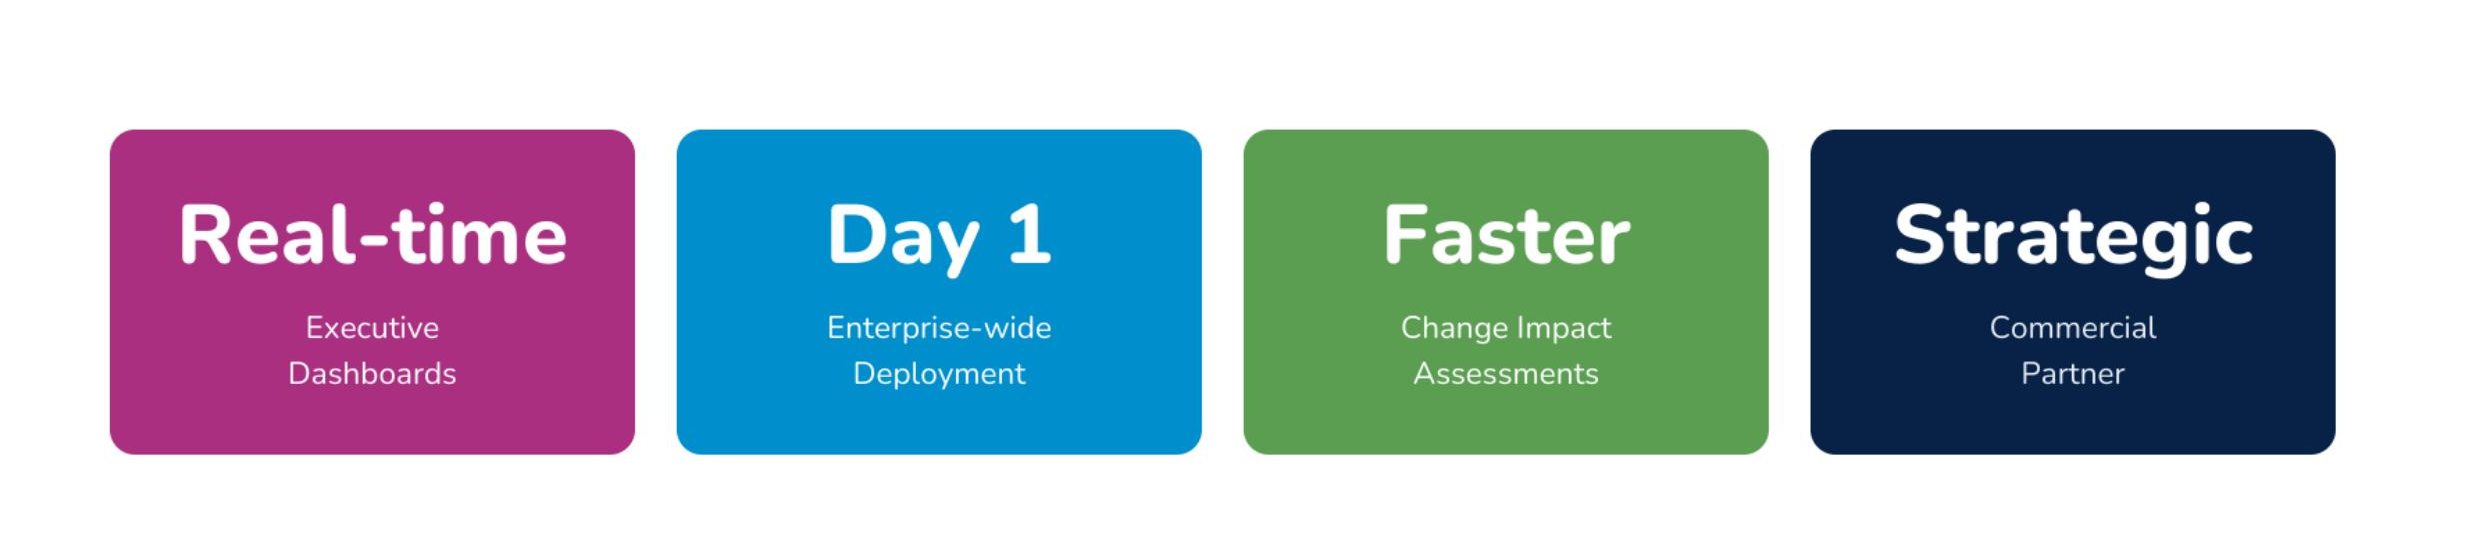
When Smith & Nephew selected Rimsys, they deployed enterprise-wide from day one. Executive reporting moved from manual fire drills to real-time dashboards. Change impact assessments became faster and more consistent. The regulatory team made the shift from reactive compliance function to strategic partner to the business.

The Challenge

Regulatory data at Smith & Nephew lived in multiplespreadsheets, shared drives, SharePoint sites, emails, and disconnectedsystems. Without a centralized record, the team could not reliably trackregistration timelines, measure on-time submissions, assess change impacts, orunderstand the downstream impact of product changes across markets. Preparingexecutive reporting meant manually assembling data from multiple sources, aprocess that consumed time and introduced risk each time.

The Solution

Smith & Nephew selected Rimsys for its configurable, notcustomized, platform: an intuitive user interface, centralized submissionmanagement, robust metrics, change assessment capabilities, and UDI supportwith machine-to-machine transmission. Rimsys’ interconnected modulearchitecture linked products, registrations, projects, change assessments, andUDI in a centralized location.

Rather than piloting in one business unit, Smith &Nephew deployed Rimsys across the entire regulatory organization from day one.The decision was deliberate: a partial deployment would have preserved thefragmentation. Enterprise-wide adoption established consistent metrics,standardized processes, and a single source of truth from the start.

The Results

Executive and board reporting, previously built from manualdata pulls, now flows directly from Rimsys in real time. What had been adisruptive, recurring effort is now a routine view. Leadership has thevisibility to make faster, more confident decisions, and the regulatory team isno longer pulled into reporting fire drills.

Change management has also been transformed. Direct linkagebetween products, registrations, and projects means impact assessments arefaster and less dependent on individual knowledge. UDI operations havesimilarly improved: machine-to-machine transmission has reduced manual uploadsand centralized DI record visibility supports global UDI requirements.

The most significant shift is strategic. With centralizedregulatory intelligence and real-time data, Smith & Nephew’s regulatoryteam now actively supports commercial planning: informing budget cycles,guiding renewal and launch sequencing, and advising on regulatory pathways toaccelerate market entry. Regulatory is no longer a downstream compliancefunction. It is a business partner.

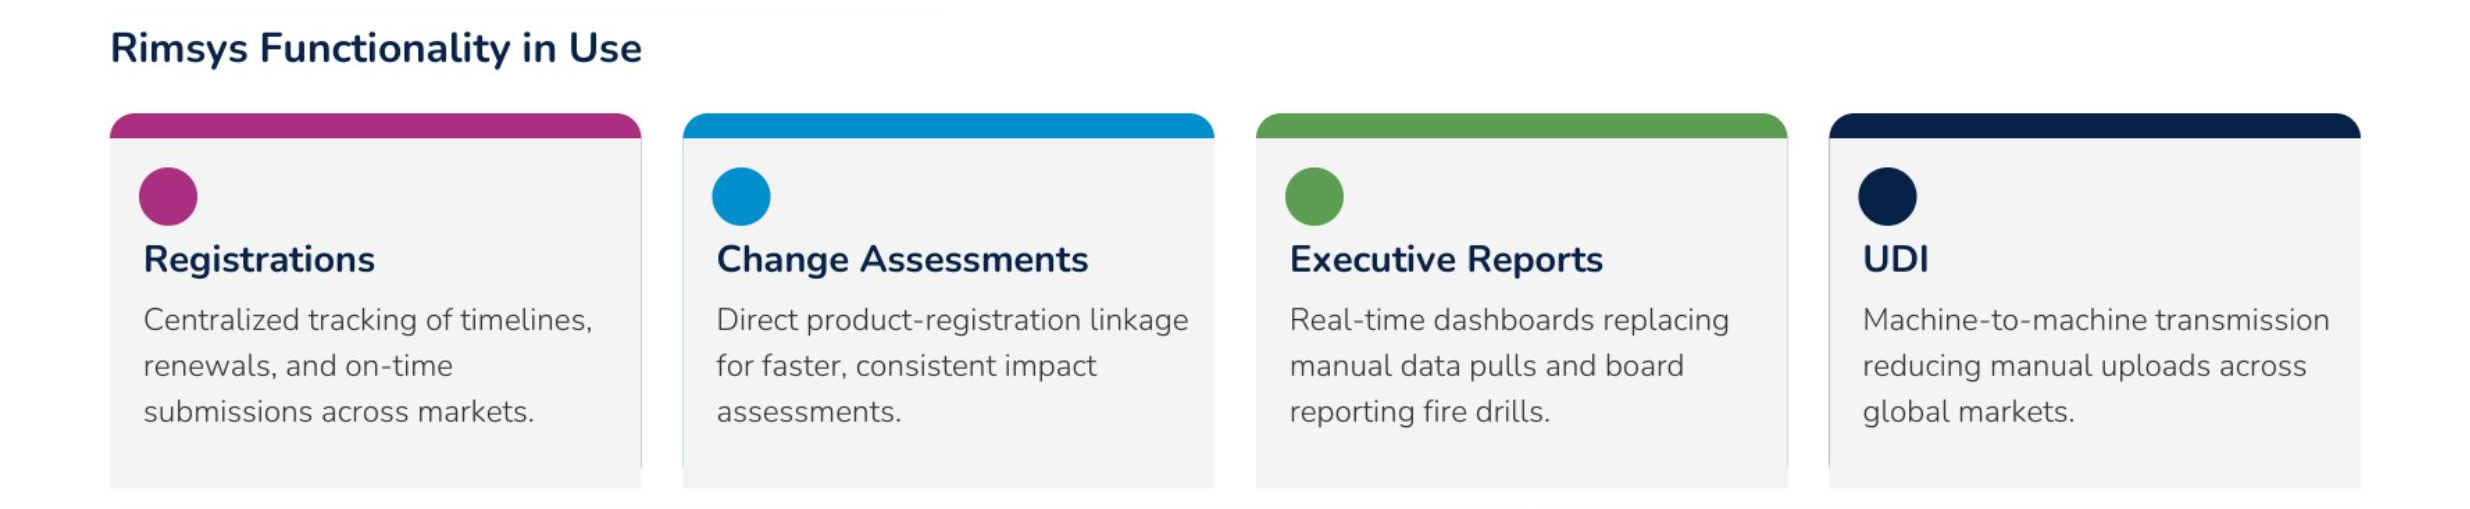
Smith & Nephew now runs four modules across its RIM operation:

  • Registrations— Centralized license tracking across 250 countries and 30+ business units
  • Change Assessments— Direct product-registration linkage for faster, consistent impact assessments
  • Executive Reports— Real-time dashboards replacing manual data pulls and board reporting fire drills
  • UDI— Machine-to-machine transmission reducing manual uploads across global markets

Take this to your team

If you’re evaluating how to modernize RIM operations at scale, the Smith & Nephew case study is a practical reference to share internally. It covers the full implementation story, module breakdown, and results data in a format built for stakeholder conversations.

Download the Case Study

I agree to the privacy policy including to Rimsys using my contact details to contact me for marketing purposes.
Thank you! Your submission has been received!
Oops! Something went wrong while submitting the form.
Hand holding smartphone showing email app with 12 unread messages notification.