


Regulatory Strategy as a Competitive Advantage
This article is an excerpt from the Regulatory strategy as a competitive advantage ebook.
Table of Contents
- The regulatory revenue opportunity
- Regulatory responsibilities
- Limitations of the "cost-center" approach to regulatory affairs
- Regulatory as a revenue function
- Competitive advantage #1: Faster time to market
- Competitive advantage #2: Cost avoidance
- Competitive advantage #3: Out-pacing competitors
- Why invest in regulatory/revenue alignment?
- Getting started - 3 steps to move towards a revenue-aligned RA team
It is well known that medical technology (medtech) companies are highly regulated, given the potential risks their products present. Understanding and complying with the complex regulations in each country is, therefore, a necessary part of marketing and selling medical devices. To realize any revenue from a medical device, it must not only demonstrate compliance with all applicable regulations, it must also receive and maintain market clearance from each country in which it is to be sold. No market clearance means no revenue. Given the key role regulatory compliance plays in revenue attainment, regulatory teams, tools, and processes present a significant opportunity for differentiation for organizations willing to invest in them.
For the majority of medtech companies, however, regulatory departments have traditionally been treated as operational cost centers, with departmental improvements focused on cost reduction and efficiency improvements. Limited investment in people and tools, and limited interest in digital transformation, have left regulatory teams across the medtech industry underfunded and under-resourced.
This has led to great resourcefulness within the RA community, where most members can point to heroes within their team who worked long hours to meet a submission deadline, headed off a disaster by uncovering a pending expiration, created ad-hoc systems to organize information and streamline communication between the RA and QA teams for smoother audits, or have otherwise gone above and beyond their typical responsibilities.
Regulatory teams, however, have the potential to be a revenue-driving competitive weapon for companies that are willing to look at them a little differently and invest in regulatory performance above regulatory cost-effectiveness. Well-supported regulatory teams can provide a true competitive advantage by providing the resources and direction to:
- Capture market share by being first to market with novel devices.
- Avoid lost revenue by effectively tracking and planning for registration renewals/updates.
- Out-pace competitors and grow market share by adapting to regulatory changes more quickly and taking advantage of competitors’ non-compliance or inability to enter a new market.
We believe we are entering a new era for regulatory affairs within the medtech industry. One in which RA teams have a seat at the table when go-to-market, competitive positioning, and strategic decisions are being made.
In the medtech industry, regulatory affairs (RA) teams have a broad range of responsibilities across the product lifecycle:
Premarket regulatory strategy
Obtaining market clearance for a new medical device is the primary activity typically attributed to RA teams. It is not unusual for a regulatory team to be given market entrance projects with little warning, but ideally, the RA team would be brought in as early as possible to contribute to go-to-market discussions.
Premarket regulatory strategy, at a minimum, involves:
- Determining the most appropriate pathway to market approval. For example, a 510(k) or PMA submission in the U.S.
- Working with quality, product, and other teams to gather information needed for market submission.
- Establishing communication with applicable regulatory bodies and third-party approved auditors.
- Compiling and submitting required documentation for market approval. This includes managing follow-up activities, questions, and requests for additional information throughout the approval process.
Forward-thinking organizations often look to bring in RA teams even earlier in the process. As regulatory experts, RA professionals can provide unique insight into product development plans. In consultation with R&D teams, can help to refine product strategies, and steer development in areas that will reduce regulatory hurdles when new products are ready to be commercialized.
Maintaining regulatory compliance for existing products
While the primary focus of regulatory teams is often considered to be new market submissions, the majority of their time is actually spent on maintaining compliance for products that are already in-market. Even in situations where market registrations do not expire, constant vigilance is required to ensure that devices remain compliant with current regulations. These efforts take a considerable time for a typical RA team because information is often spread across disparate systems, where it can be difficult to find and confirm.
Maintaining regulatory compliance for approved devices includes:
- Staying on top of changing standards and making changes as required to existing technical files and other documentation.
- Submitting appropriate documentation updates when there is a change made that could potentially affect the efficacy or safety of the product, such as a material switch or facility change.
- Understanding pending regulatory changes and proactively addressing any that have an impact on devices currently in-market.
- Tracking registration expirations and preparing for timely re-submissions to ensure there is no lapse in market clearance.
Post-market activity
Post-market surveillance and vigilance activities are required by most countries and should involve the cooperation of the quality and regulatory teams. Ensuring that changing post-market reporting requirements are understood and complied with is an important regulatory responsibility.
Regulatory teams typically play a role in:
- Post-market surveillance of adverse events, complaints, and any issues associated with a device in the field.
- Assembling and submitting any required periodic safety reports to country/regional health authorities.
- Post-market vigilance and reporting of serious events to the appropriate regulatory agencies.
- Any required communication with regulatory authorities regarding adverse events or concerning trends in product quality.
Ask any RA professional, and they are likely to tell you that they work long hours and are often scrambling to meet looming deadlines...
To continue reading this ebook, download the full version.

The Future of MedTech Compliance: How AI Is Transforming Regulatory Affairs
MedTech regulatory affairs teams are facing a turning point. Regulations are expanding in number and complexity, resources are limited, and manual processes cannot keep up. At the same time, artificial intelligence (AI) has become a serious topic of discussion in regulatory circles. Leaders are beginning to ask: How can AI help us manage change, reduce risk, and accelerate compliance efforts?
The answer is clear: AI is no longer just a buzzword. When combined with effective regulatory information management (RIM), it can be a powerful enabler of efficiency, accuracy, and strategic decision-making.
Why AI is Trending in Regulatory Affairs
The Surge of Regulatory Data
Regulatory teams must now track requirements from multiple global markets. Each regulator frequently updates its regulations, guidances, templates, and recognized standards, which creates large volumes of data to organize and analyze. AI can scan and classify this information, highlight changes, and prepare it for structured use within RIM systems.
Doing More with Limited Resources
Most teams are expected to deliver more without additional staff. High turnover makes continuity difficult, and according to the 2024 RAPS Global Workforce Report, the number of professionals “open to work” has grown in North America and Europe. AI offers relief by taking on repetitive tasks such as document formatting or data entry, allowing experts to focus on higher-value work.
Global Complexity and Diverging Standards
No two markets are exactly alike. AI can help by flagging differences, surfacing potential risks, and recommending reusable content drawn from a company’s submission history. Faster, more accurate submissions directly improve time-to-market and compliance outcomes.
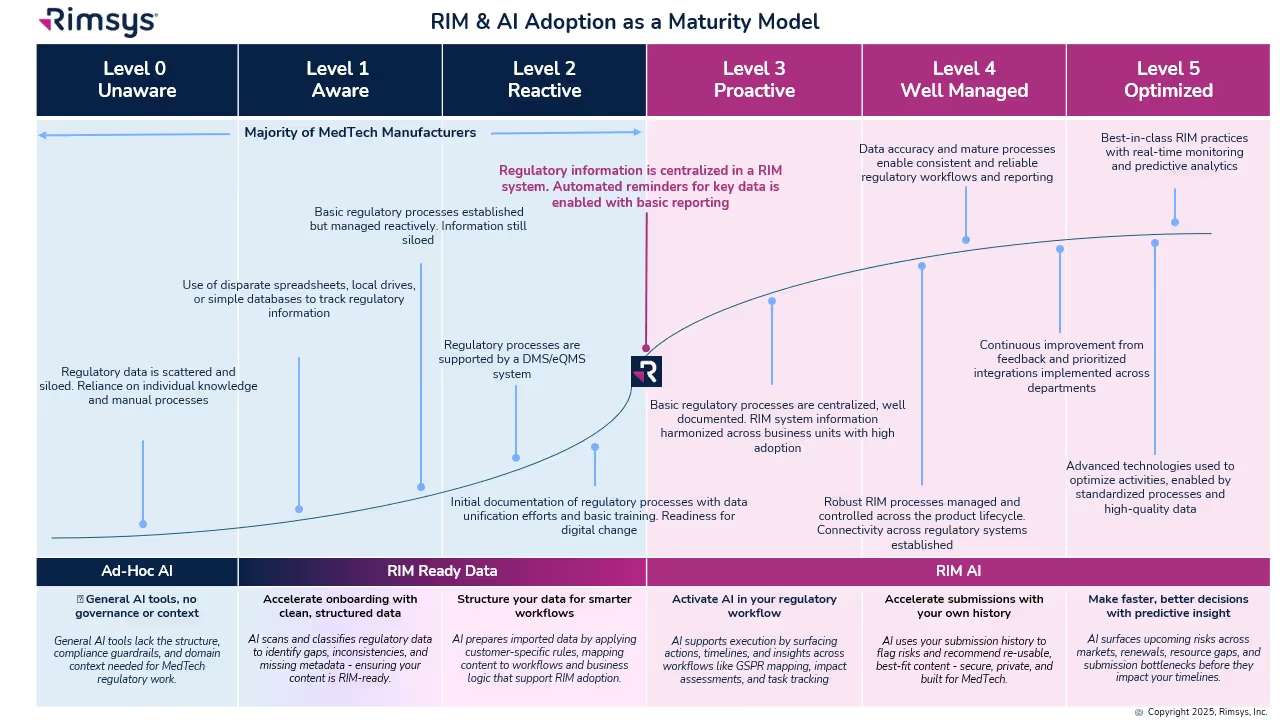
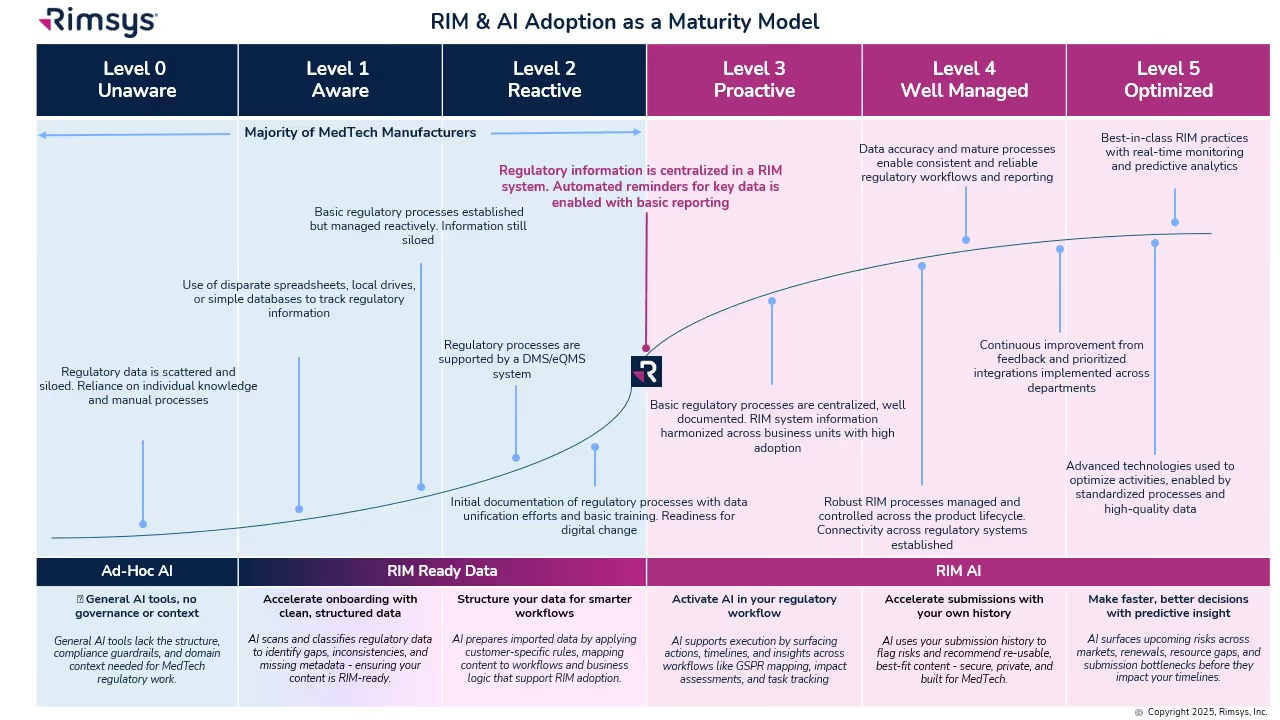
The RIM & AI Adoption Maturity Model
Not every organization is ready to fully embrace AI. Success depends on RIM maturity: how structured and centralized your regulatory processes and data are. The RIM & AI Adoption Maturity Model provides a roadmap from basic to optimized states.
- Levels 0–2: Early Stage
- Data is siloed and processes are ad hoc. AI provides value in isolated ways, such as cleansing records or scanning for regulatory changes.
- Level 3: Proactive
- A RIM system centralizes information. AI begins to surface reminders, deadlines, and global impact assessments.
- Level 4: Well Managed
- Processes are standardized across the lifecycle. AI generates insights, monitors KPIs, and supports reuse of regulatory content.
- Level 5: Optimized
- AI is fully embedded, delivering predictive analytics, continuous monitoring, and smarter decision-making.
Practical Applications of AI Today
Today, regulatory teams see the greatest opportunities in:
- Regulatory submissions: Automatically detecting changes in templates and suggesting updates.
- Document classification: Using natural language processing to tag and organize regulatory documents.
- Regulatory intelligence: Monitoring health authority updates and highlighting what matters most.
- Impact assessments: Linking changes (e.g., regulations/standards/design) directly to the affected products and registrations and evaluate the potential impact.
- Content reuse: Recommending approved content to accelerate submissions.
How to Start Your AI Journey in Regulatory Affairs
Adopting AI is not about jumping to the most advanced capabilities overnight. Instead, consider these steps:
- Assess your RIM maturity. Where does your organization sit on the 0–5 scale? What foundational gaps (data centralization, process standardization) need to be addressed first?
- Identify quick wins. Focus on repetitive, rules-based tasks where AI can add value without major disruption.
- Implement governance. Establish policies for safe, compliant AI use, particularly around data privacy and model training.
- Pilot in phases. Start small, validate results, and expand AI use as confidence and maturity grow.
- Keep people at the center. AI should enhance the expertise of regulatory professionals, not replace it.
Building a Smarter Future for MedTech Compliance
AI is becoming a trending topic in regulatory affairs not just because it’s new, but because it directly addresses the challenges teams face: rising complexity, limited resources, and scattered data.
For organizations that take this approach, the benefits are clear: lower compliance risk, faster execution, and stronger competitive positioning. AI does not replace regulatory professionals. Instead, it enables them to spend less time on manual tasks and more time on strategic contributions that improve patient access to life-changing technologies.
In other words, AI isn’t about futuristic transformation. It’s about helping regulatory teams step off the “data treadmill” and reclaim their time for what matters most: bringing safe, life-changing medical technologies to patients faster.
Rimsys Becomes the Trusted Regulatory Partner for 6 of the Top 12 Global MedTech Manufacturers
“Adoption by half of the top global MedTech manufacturers is a powerful validation that we’re not just a solution, we’re setting the new gold standard for regulatory excellence.”
Pittsburgh, PA - August 7, 2025 - Rimsys, the leading Regulatory Information Management (RIM) software purpose-built for the MedTech industry, today announced a significant milestone: 6 of the world’s top 12 medical device manufacturers now rely on Rimsys to manage and streamline their global regulatory operations.
This milestone further solidifies Rimsys’ position as the trusted partner to the world’s most innovative and quality-focused MedTech companies.
Click here for the full list of the top 12 global MedTech companies.
“Today’s regulatory environment demands more than spreadsheets. Leading manufacturers recognize that regulatory operations are mission-critical, revenue-generating departments and need systems to match that level of importance,” said James Gianoutsos, Founder and CEO of Rimsys.
Rimsys’ unified, enterprise-grade RIM platform centralizes and automates critical regulatory processes—including market registrations, Unique Device Identification (UDI), essential principles/GSPR, and submissions management—reducing compliance risk and accelerating market access. Specifically tailored to the needs of medical device and diagnostics companies, Rimsys enables seamless collaboration across RA, QA, and commercial teams while delivering the audit-ready transparency global regulators demand.
“As more organizations embrace regulatory digital transformation, Rimsys is proud to lead the industry forward,” added Gianoutsos. “Adoption by half of the top global MedTech manufacturers is a powerful validation that we’re not just a solution, we’re setting the new gold standard for regulatory excellence.”
To learn more about the Rimsys, please visit www.rimsys.io.
About Rimsys
Rimsys is the leading provider of Regulatory Information Management (RIM) software purpose-built for MedTech manufacturers. The comprehensive platform digitizes and automates regulatory activities, helping MedTech regulatory affairs teams to efficiently achieve regulatory compliance and get products to market faster. Rimsys is designed around MedTech workflows and supports a full breadth of regulatory functions including registrations, submissions, UDI, EUDAMED compliance, essential principles, and standards management in a unified platform. Rimsys is trusted by half of the world’s top 12 MedTech companies to power their global regulatory operations. For more information, visit www.rimsys.io.
Rimsys Announces Bulk UDI Submission and Rimsys Connect™ to Empower MedTech Regulatory Teams
New solutions deliver enterprise-grade data access and streamlined EUDAMED compliance, driving smarter, faster decisions across the business
Pittsburgh - April 29th, 2025 - Rimsys, the global leader in Regulatory Information Management (RIM) software for the MedTech industry, today announced two major enhancements to its platform: expanded Unique Device Identification (UDI) capabilities to support EUDAMED machine-to-machine (M2M) bulk transmission and Rimsys Connect™, a new enterprise Change Data Capture (CDC) solution that provides near real-time synchronization of Rimsys data with customers’ Business Intelligence (BI) solutions.
Together, these capabilities are designed to help MedTech organizations streamline compliance, reduce manual effort, and unlock the full strategic value of their regulatory data.
New UDI Capabilities Support EUDAMED Readiness
The UDI enhancements extend Rimsys’ industry-leading Universal UDI® framework, enabling MedTech teams to manage complex, global UDI programs in one unified RIM system. Key new capabilities include:
- Approving multiple records simultaneously via a simple, scalable workflow
- EU data governance support with all required attributes for EUDAMED transmission
- Bulk submission of records to both the GUDID and EUDAMED databases
These features allow teams to eliminate time-consuming, record-by-record processing, helping them meet the mandatory January 2026 EUDAMED compliance deadline with confidence.
“We’ve partnered closely with our customers to develop a UDI offering that meets increasing regulatory complexity and is easily scalable as new regulations come online,” said Adam Price, Director of Regulatory and Technical Programs at Rimsys. "We’re not only giving customers the ability to meet EUDAMED compliance but enabling them to manage their global UDI program in a single-sourced RIM solution for complete visibility.”
Introducing Rimsys Connect™: Enterprise Data Access, Redefined
Rimsys Connect™ offers enterprise customers a powerful new way to leverage regulatory data across the business. Built on a scalable, event-driven architecture, it provides secure, structured, near real-time streaming of Rimsys data into any modern data warehouse solution—such as Snowflake, Amazon S3, and Salesforce Bulk API 2.0.
“Rimsys Connect™ is not just a connector—it’s a strategic enabler,” said James Gianoutsos, Founder and CEO of Rimsys. “We’re giving regulatory affairs teams the ability to deliver insights that influence launches, accelerate tender responses, and align compliance with business impact. With Connect, RA teams become true strategic partners.”
By providing full access to customer data—registrations, UDI, projects, tasks, and custom attributes — Rimsys Connect™ supports a wide variety of enterprise use cases with customers’ own business intelligence solutions:
- Tracking on-time submission and decision KPIs
- Aligning registration timelines with product launch dates
- Conducting ROI analysis for renewals and market prioritization
- Accelerating tender readiness by combining RIM and PLM data
- Supporting post-market surveillance dashboards
While the initial release will focus on data access, Rimsys plans to expand Connect with curated BI templates and best practices to further accelerate enterprise customer time-to-value.
Solving the Data Fragmentation Problem for MedTech
Many regulatory affairs teams remain constrained by outdated tools, fragmented data sources, and increasing demands to deliver strategic insights to executive and commercial stakeholders. Rimsys Connect™ addresses these challenges by eliminating manual reporting workflows and enabling teams to analyze their regulatory data alongside financial, marketing, and quality systems.
“With Rimsys Connect™, regulatory teams can visualize and analyze their data in real time, assess launch readiness, and deliver more value to their organizations. This is how RA becomes a catalyst for better decisions—not just compliance,” said Gianoutsos.
Both Rimsys’ expanded UDI capabilities and Rimsys Connect™ will be available this summer. Those interested in learning more about these solutions and how they will enable greater automation, efficiency, and compliance can visit our booth at RAPS Euro Convergence May 13-15 in Brussels, Belgium, or sign up for Rimsys’ product update webinar on Thursday, May 22nd at 10 AM ET.
Read the press release here.
About Rimsys
Rimsys is improving global health by accelerating delivery and increasing availability of life-changing medical technologies. Rimsys Regulatory Information Management (RIM) software digitizes and automates regulatory activities, helping MedTech regulatory affairs teams to plan more effectively, execute more quickly, and confidently ensure global regulatory compliance. Rimsys is designed around MedTech workflows and supports a full breadth of regulatory activities including registrations, submissions, UDI, essential principles, and standards management in a unified platform. For more information, visit www.rimsys.io.
Contacts:
marketing@rimsys.io
Rimsys Enters Strategic Alliance Relationship with KPMG
PITTSBURGH – March 11, 2025 -- Rimsys, the global leader of MedTech Regulatory Information Management (RIM) software, today announced that it has entered into a strategic alliance relationship with KPMG to advance digital transformation in the MedTech industry.
“KPMG’s deep experience in advisory and business transformation services and exceptional reputation make them a valuable alliance relationship for us,” said James Gianoutsos, Founder and CEO of Rimsys. “KPMG is on the cutting edge of industry trends and has a wide breadth of experience in helping companies innovate and scale. We are thrilled to work with them to help MedTech teams transform their regulatory management processes and leverage the benefits of automation and digitization as part of their broader transformation strategy.”
Founded for and by MedTech regulatory affairs professionals in 2017, Rimsys was created to bring efficiency to regulatory information management and fill an inhibitive technology gap in an underserved industry. Rimsys has since grown to support the world’s MedTech leaders backed by a staff that understands their complex workflows and a robust, secure technology infrastructure that allows customers to scale Rimsys software to support their changing regulatory needs and requirements.
“There is tremendous innovation happening in the MedTech industry, and we are excited to work with Rimsys to help clients transform how they manage regulatory information for getting new products to market and sustaining their existing product portfolios. It’s critical to approach these programs as a holistic business transformation across people, process, technology, data, and governance & controls,” said Dipan Karumsi, Principal, Consulting Sector Leader for Life Sciences at KPMG.
“Through our strategic alliance with KPMG, we can further expand our reach to large and enterprise MedTech companies and continue our exponential growth,” said James. “Combined with KPMG’s experience helping organizations mature their data collection and transformation processes to reach RIM readiness, we can enable the MedTech industry to innovate faster, strengthen compliance, and most importantly, improve the availability of life-changing medical technologies.”
See the full press release here.
FDA’s Final Rule on LDTs: What manufacturers need to know
In July 2024, the FDA's final rule in 21 CFR Part 809 on laboratory developed tests (LDTs) went into effect, amending its previous regulations to make it clear that IVDs, including those that are manufactured in laboratories, are classified as devices under the Federal Food, Drug, and Cosmetic Act. Our blog post provides an overview of LDTs, FDA’s final rule, the phase out policy schedule, and how LDT manufacturers can prepare themselves for compliance.
What are LDTs?
Simply put, LDTs are IVDs that are designed, manufactured, and utilized within a certified laboratory and are typically used for high-complexity testing.
Historically, FDA has used enforcement discretion only on LDTs, which means that most LDTs haven’t been subjected to specific regulatory requirements. However, the volume of and risks associated with LDTs have grown over the years. Some examples of modern LDTs include glucose tests, genetic tests for cancer and infectious diseases, and newborn screenings for early diagnostics. Without a regulatory framework in place, patients are at greater risk of receiving inaccurate test results, forgoing necessary or undergoing unnecessary treatment, and adhering to misleading or false product claims, possibly endangering patients and leading to higher healthcare costs.
FDA’s Final Rule on LDTs
Following the final rule that was issued, LDTs are now subject to the same regulatory requirements as other IVDs, including premarket reviews, quality system requirements, labeling requirements, adverse event reporting, and device listing and registration. To prevent disruptions in patient care, there is a four-year transition or phaseout period consisting of the following five stages:
- Stage 1 (May 6, 2025): LDT manufacturers will be expected to comply with FDA medical device reporting (MDR) requirements, correction and removal reporting requirements, and quality system (QS) requirements for complaint files.
- Stage 2 (May 6, 2026): LDT manufacturers will be required to comply with IVD registration and listing requirements, labeling requirements, and investigational use requirements.
- Stage 3 (May 6, 2027): LDT manufacturers will need to comply with all other QS requirements not covered in Stage 1.
- Stage 4 (November 6, 2027): Unless a premarket submission is received before the start of this stage, LDT manufacturers of high-risk products will need to comply with premarket review requirements for IVDs that may be classified into class III or that meet the requirements of section 351 of the Public Health Service Act.
- Stage 5 (May 6, 2028): LDT manufacturers of moderate and low –risk products will need to comply with premarket review requirements for IVDs unless a submission is received before the beginning of this stage.
Manufacturers of LDTs that don’t meet the requirements in each stage are deemed non-compliant to the regulations governing IVDs and may be subject to FDA 483 observations or warning letters, financial penalties, and even worse, involuntary removal of products from the market.
Note that some LDTs will be exempt from these requirements. Refer to the FDA’s website for more guidance.
Preparing for Compliance
Despite a four-year phaseout period, it’s crucial for LDT manufacturers to start assembling a compliance plan. Starting as early as May 2025, manufacturers will be required to comply with FDA Medical Device Reporting (MDR) requirements, correction and removal reporting requirements, and quality system requirements for complaint files.
It's good practice to conduct an internal regulatory assessment to ensure you have the resources, processes, and tools in place to successfully meet new requirements for LDT devices. It’s also essential to make sure your team is well-versed in these new requirements and the documentation and timelines involved. Including all relevant stakeholders early on, getting a comprehensive project plan in place, and meeting regularly to ensure all tasks are completed would be helpful during the phaseout period and beyond.
Regarding the LDT changes, the FDA has provided a Q&A sheet that you may find helpful.
If you're looking for guidance on FDA premarket submissions, see our Beginner’s Guides to the FDA 510(k), De Novo, and PMA processes.
How Regulatory Tools Can Help
FDA’s final rule on LDTs will add complexity to the regulatory information management of laboratory diagnostic tests. There are digital solutions that can help manufacturers stay current on updated regulations and manage the additional information and documentation needed because of these updates.
A regulatory intelligence database like Rimsys Intel can provide detailed global market entrance requirements, application timelines, fees, risk class specifications, and documentation needed for medical devices and IVDs so that manufacturers can start preparing their premarket strategies.
Regulatory Information Management (RIM) software like Rimsys can help boost efficiency, reduce compliance risk, and increase collaboration by centralizing regulatory information and automating time-consuming, manual processes. As a result, medical device manufacturers gain complete visibility into their submission management and selling status so that they can plan more effectively, avoid costly product delays, and execute faster.

Planning Your Enterprise’s UDI Strategy for EUDAMED and Beyond
Rimsys recently hosted a webinar with RAPS titled “Planning Your Enterprise’s UDI Strategy for EUDAMED and Beyond.” During the session, our experts, Adam Price, Director of Regulatory and Technical Programs, and James Gianoutsos, Founder and CEO, provided best practices to help MedTech companies navigate the complexities of Unique Device Identification (UDI) compliance. The session covered key topics such as comparing US and EU UDI regulations, preparing for the January 2026 mandatory EUDAMED submission date, and using EUDAMED as a foundation for a global UDI strategy that can scale as additional markets adopt UDI requirements.
Here are some of the key topics we discussed to help Medtech teams prepare their strategy:
1. The importance of UDI to regulatory affairs teams
UDI is a globally unique identifier for medical devices that helps improve traceability, manage adverse event reporting, and ensure compliance with regulatory requirements. Regulatory affairs teams play a critical role in both establishing and maintaining UDI data to meet submission timelines and market placement requirements.
Responsibility for UDI management can vary between organizations, with some MedTech companies assigning the responsibility to IT or supply chain teams. When we asked our audience who is responsible for UDI management at their company, 80% of respondents said that it was their regulatory affairs team, emphasizing UDI’s crucial role in market placement.
2. Comparing US and EU UDI Requirements
FDA GUDID requirements have been in place since 2014. As such, some MedTech teams may plan to use their GUDID submission strategy as a baseline for the development of their EUDAMED submission strategy. While the US FDA’s GUDID system has standardized requirements, the EU’s EUDAMED is more complex due to additional data fields, language-specific entries, and the use of the EMDN nomenclature instead of GMDN. EUDAMED also introduces the Basic UDI-DI, which connects related devices within family groupings and links different EUDAMED modules. Companies must adapt their UDI strategies to account for these variations.
“Unlike the standardized US GUDID, the EU’s EUDAMED is more complex, featuring multiple device categories, language-specific data, and the EMDN nomenclature.” - Adam Price
3. The Interconnectivity of EUDAMED
EUDAMED consists of multiple interconnected modules. The Basic UDI serves as the key data element linking these modules together. This interconnected structure ensures that regulatory bodies, manufacturers, and economic operators can efficiently track devices throughout their lifecycle. Properly aligning UDI submissions with these modules is essential for seamless compliance and data consistency.
“The interconnectivity of EUDAMED is really important because the Basic UDI family groupings are the key piece of data that is going to tie the different modules together. For example, your notified body certificates for the regulations will have your Basic UDI numbers indicated on them. And then that's connected to the Basic UDI numbers that you have associated to your single registration number.” - Adam Price
4. Deadlines, Compliance Risks, and Recommended Actions per Device Category
The deadline for mandatory UDI submissions to EUDAMED is January 1, 2026, with a six-month grace period until June 2026 for devices already on the market. Failure to meet these deadlines could result in compliance risks, including regulatory penalties and restricted market access. Different categories of devices are subject to specific rules:
- Regulated Devices: Devices compliant with MDR or IVDR regulations must be linked to Basic UDI groupings and classified accurately under EMDN. Recommended actions include ensuring all data aligns with the new requirements and preparing for immediate submissions of new devices after January 2026.
- Legacy Devices: Devices compliant with older EU directives that remain on the market do not require a Basic UDI but must still have all essential data submitted. Companies should consolidate and verify legacy data, ensuring consistency with prior regulatory approvals.
- Non-Registered Devices: Devices no longer placed on the market but still have post-market surveillance activities must be registered for traceability. Manufacturers should identify these devices, gather historical data, and ensure that post-market surveillance records are aligned with regulatory expectations
.
5. Data Preparation and Submission Process
Data preparation for EUDAMED compliance is going to take some time, and the time to start preparing is now. It's important to recognize some of these high-level steps needed for effective UDI data preparation and submissions so that MedTech teams can plan appropriately:
- Identify: Collect and review all required UDI data, including language-specific fields and Basic UDI groupings.
- Verify: Ensure data accuracy and alignment with regulatory requirements, using EUDAMED’s business rules as a reference.
- Format: Structure data to meet EUDAMED’s formatting and enumeration rules, ensuring compatibility with submission methods.
- Submit: Transmit data using manual entry, XML uploads, or automated machine-to-machine (M2M) transmission, depending on the volume and complexity of your product portfolio.
“Ultimately, manufacturers need to be in a position to have all the data in for all products - not just the new products on the market but for all products by the end of that transitional period. It sounds a little apocalyptic but given the right planning, I think it's very achievable." - Adam Price
6. Getting Data into EUDAMED
Manufacturers have multiple options for submitting UDI data into EUDAMED, depending on their resources and portfolio size. Data can be manually entered through the EUDAMED user interface, uploaded via XML files, or submitted using automated machine-to-machine (M2M) transmission. Organizations managing large product portfolios are encouraged to adopt M2M transmission to ensure efficiency and data accuracy. Testing data submissions in the EUDAMED playground before official submission is highly recommended to identify and resolve any formatting or compliance issues.
“With more products, more data, and more BUDI families, there's more complexity. And as you're looking across your product portfolio, that's something that you need to consider as you plan your submission strategy.” -Adam Price
7. UDI Management Solutions
Organizations can choose from various UDI management solutions depending on their size and needs:
- Spreadsheets: Might be suitable for small companies with limited product portfolios but are time-consuming to manage and error-prone
- Repurposed Internal Systems: PLM or PIM systems can manage UDI data but may require manual processes and in-house expertise.
- Purpose-Built Connectors: Provide transmission capabilities for UDI data but don’t provide data capture and management
- RIM Systems (e.g., Rimsys): Provide a centralized, automated solution that integrates UDI data with product registrations and certificates, supporting seamless M2M transmissions.
“For large, complex product portfolios, RIM systems like Rimsys provide automated UDI management and seamless M2M transmissions.”
“Having an integrated data hub that's the source of truth for your UDI information is key. Then being able to associate your UDI data with all other data needed for UDI transmission in a RIM system, your registrations, certificates, legal entity, manufacturer, manufacturing locations, all the product data, all your BUDI data, having that all centralized in a RIM system is really critical to understanding and organizing your data in a way that can be meaningfully managed moving forward.” - James Gianoutsos
8. EUDAMED as a Foundation for Global UDI Compliance
There is a tremendous opportunity for EUDAMED’s comprehensive data requirements to serve as a foundation for global UDI compliance, as many countries are adopting similar regulations. Australia, Switzerland, the UK, and India are among the markets expected to enforce UDI requirements soon, with varying deadlines and data requirements. By using their EUDAMED data output as the foundation for a global UDI program, MedTech companies can build a UDI strategy that positions them to meet current and future requirements as they evolve.
“The application of UDI is not a one and done approach like it had possibly felt for manufacturers in the past. These requirements are going to need to be continually maintained from the perspective of your product characteristic data as well as new technologies as they're being introduced. So it's maintaining compliance to the current requirements, establishing compliance to the new countries’ UDI programs as those get rolled out, and being in a good position to control and consistently apply your UDI data.” -Adam Price
Conclusion
With the January 2026 deadline approaching, companies must act now to ensure compliance, streamline submissions, and prepare for future UDI requirements worldwide. Establishing a robust UDI strategy today will not only support compliance with EUDAMED but also lay the groundwork for expanding regulatory demands in other global markets. Leveraging centralized systems like RIM platforms can help manage the increasing complexity of global UDI regulations, ensuring consistent and efficient compliance.
“This seems to be a trend globally that UDI has become this single source data point to control and manage and maintain your products on the market. And so EUDAMED is just really the tip of the iceberg. Even though we know how complex it is, this is really just the beginning of this global UDI strategy that you have to be thinking about as you go into EUDAMED preparation.” -James Gianoutsos
Those interested in watching the webinar can access the recording here.
With an integrated and automated approach to UDI, Rimsys is a trusted and proven solution to help MedTech teams simplify EUDAMED compliance and assemble a global UDI program that meets current and future requirements. Request a custom demo rimsys.io/demo.