
2023 Regulatory performance report
Today at Rimsys, we unveiled the 2023 MedTech Regulatory Performance Report, a new set of insights into the state of medtech regulatory affairs. Compiled based on interviews with 200 regulatory professionals and executives, the study provides a detailed look into how regulatory teams are staffed, their processes, the tools they use, and ultimately how they perform.
Why did we create this study? There were two driving factors behind the research. The first was a common theme that we heard from a number of our customers: Regulatory leaders don’t have clear data and benchmarks. They don’t necessarily know how long a new market submission should take, and how to plan for or assess the work of their teams. While other studies look at the medtech industry broadly or the state of the regulatory profession, this study tries to build a comprehensive resource for regulatory (and company) leaders.
The second factor was really for ourselves and the team at Rimsys. As a company building solutions specifically for medtech regulatory affairs, we wanted more insight into where companies were successful, where they struggled, and where we can add value.
What did we find? Regulatory teams perform a lot of hero work and rate themselves highly for their accomplishments. At the same time there is a lot of opportunity for process improvements, and companies that invest in digital transformation for regulatory affairs see better performance.
Regulatory professionals are superheroes
Regulatory teams are generally pretty small. Most companies have less than 10 full-time regulatory professionals. These small teams complete an enormous amount of work. Last year on average, RA teams completed 50 license renewals, 50 license updates, and 10 new market submissions. This is impressive output.
Digging a bit under the covers, we found that this output relied heavily on the support of external consultants. 90% of companies use consultants to keep pace with their regulatory workload. Front-line employees also struggle with burnout. They were much more likely to report feeling under-resourced than regulatory leaders.
But process problems persist
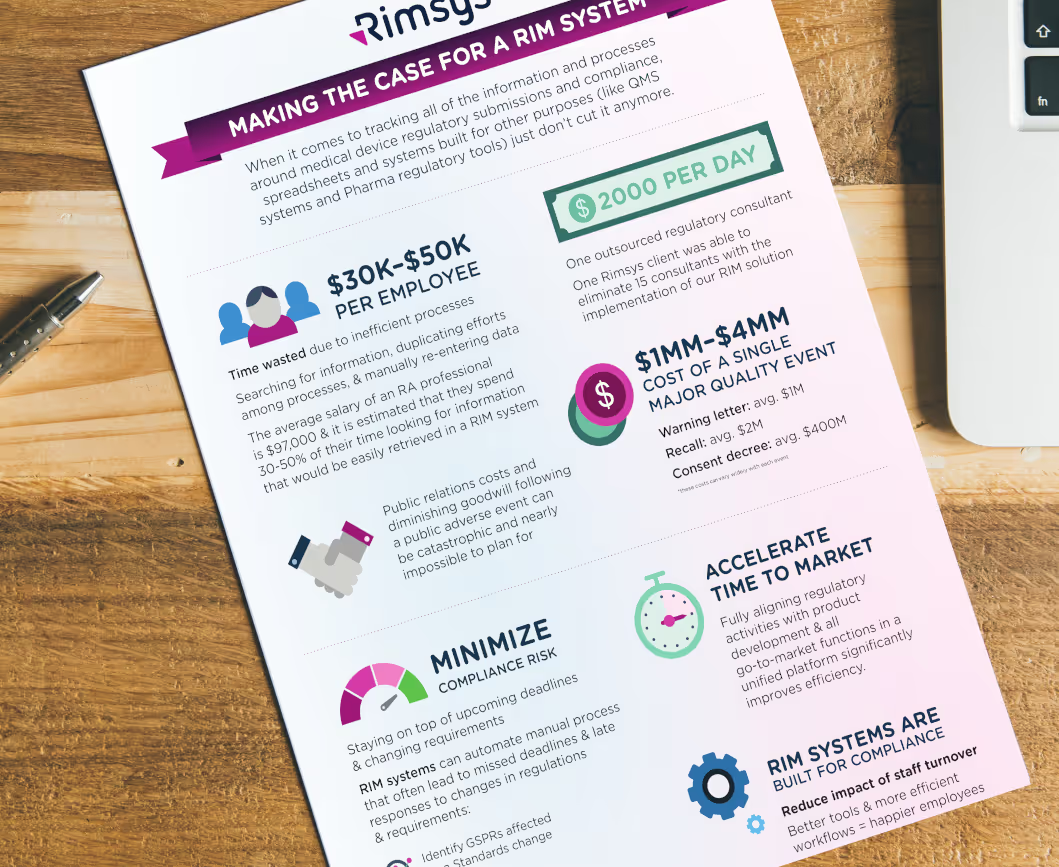
A lot of regulatory work remains extremely manual. 70% of regulatory teams spend half their time or more on repetitive administrative tasks. All of this manual work increases the frequency of errors and required rework. 61% of companies reported a major non-compliance incident in the past 2 years.
Manual work also makes it difficult to complete regulatory projects in a timely fashion. Teams completed a lot of projects, but each took a long time. Over half of all companies spend 4 months or more on license renewals, license updates, and new market submissions.
Moving regulatory affairs forward
As regulatory requirements become more complex, there’s a natural question about how teams will work moving forward. MDR & IVDR in Europe have significantly increased the regulatory workload required to bring and keep products on the market. Will organizations be able to keep pace with the same resources, tools, and processes?
No, and the performance report shows that medtech companies are investing to improve their regulatory capabilities. The majority of companies are planning to increase the sizes of their RA teams in 2023, and 40% expect to increase their investments in regulatory software. Companies are increasingly adopting specialized software to better support regulatory processes.
Dig into the survey results
The full survey results provide insights into more aspects of regulatory performance. They show that companies need to take a deeper look into their processes and how regulatory resources are allocated. There are two ways to learn more:
- Visit the survey page to see the full results (the survey whitepaper can be downloaded at no cost)
- Watch the recording of our webinar with PA Consulting. We discuss the survey results in more detail and share our regulatory predictions for 2023

Post-market surveillance for medical devices in the European Union
This article is an excerpt from Post-market surveillance for medical device in the European Union.
Table of Contents
- What is post-market surveillance?
- What classes of medical devices require post-market surveillance?
- Components of a successful post-market surveillance plan
- PMS data requirements
- Post-market surveillance system goals
- Required post-market surveillance reporting
- Embracing post-market surveillance as an integral part of your quality program
- Getting started with post-market surveillance
Post-market surveillance (PMS) is designed to monitor the performance of a marketed medical device by collecting and analyzing field use data. Article 10 of the EU MDR and IVDR requires all device manufacturers to have a post-market surveillance system in place. The main elements of the PMS are laid out in Article 83, and additional details for lower-risk and higher-risk devices are covered in articles 84 and85, respectively.
In general, a PMS system consists of both proactive activities and reactive, or vigilance, activities. While post-market surveillance and vigilance are sometimes used interchangeably, vigilance consists of separate activities that feed post-market surveillance programs.
Post-market surveillance systems are used to collect and analyze data not only about the manufacturer’s device but also about related competitors’ devices that are on the market. Data collected through PMS procedures is then used to identify trends that may lead to, among other things, quality improvements, updates to user training and instructions for use, and identification of manufacturing issues.
Note that “market surveillance” encompasses activities performed by a Competent Authority to verify MDR compliance, and should not be confused with the topic of this ebook,“post-market surveillance," which is performed by the manufacturer.
All medical devices marketed in the EU require some level of post-market surveillance, and all medical device manufacturers must implement a post-market surveillance system (PMS). The requirements of the PMS, however, vary and should be “proportionate to the risk class and appropriate for the type of device” (MDR Chapter VII). In particular, the type and frequency of reporting vary based on a device’s risk class.
A post-market surveillance plan (PMS) is an integral part of a manufacturer’s quality management system and provides a system for compiling and analyzing data that is relevant to product quality, performance, and safety throughout the entire lifetime of a device. The PMS should also provide methods for determining the need for and implementing any preventative and corrective actions. A PMS system should include and define:
Surveillance data sources
With the increased focus on proactive risk identification in the MDR, it is important to design post-market surveillance systems that actively acquire knowledge and detect potential risks. It is not sufficient to rely solely on spontaneous reporting by healthcare providers, patients, and other stakeholders.
In addition to information coming from Clinical Evaluation Reports and complaint and adverse event reporting, typical sources of surveillance data include:
• Social media networks: Because many of your stakeholders may be communicating on social media networks, it is important to employ social listening techniques and/or tools to identify issues and concerning trends as they develop.
• Industry and academic literature: Any studies, academic papers, and other literature that addresses similar devices or the specific use cases for which your device is designed should be evaluated. In particular, risk factors and adverse events identified with similar devices should be closely examined. It is also important to identify newer technologies that may affect the benefit-risk ratio and establish a new definition of “state of the art” for the device type.
• EUDAMED: While the European Database on Medical Devices (EUDAMED) is not yet fully functional, it is intended to provide a living picture of the lifecycle of all medical devices marketed in the EU. Manufacturers should take special care to consider information for similar devices made available through the EUDAMED system in the future.
• Registries: Patient, disease, and device registries can provide information that informs the clinical evaluation process which provides input into the post-market surveillance system.
Data analysis methodology
A well-defined data analysis methodology will accurately identify trends and lead to defendable decisions in the application of post-market experience. Once the necessary information has been identified and collected, and potentially cleaned of incomplete or otherwise unusable data, the data needs to be analyzed.
The goal is to identify meaningful trends, correlations, variations, and patterns that can lead to improvements in the safety and efficacy of the device. There are many data analysis tools available that can assist with:
• Regression analysis that will identify correlations between data (e.g. the device location/geography correlates to battery life).
• Data visualization that can be useful in spotting trends in the data.
• Predictive analytics, which can be particularly useful with large data sets, to identify future trends based on historical data.
• Data mining, which is also normally used with large datasets, to organize data and identify data groups for further analysis.
Benefit-risk indicators and thresholds
The MDR requires that medical device manufacturers not only demonstrate the clinical benefit of their device but also quantify the benefit-risk ratio. The benefit of a device must be shown to clearly outweigh the risk for it to gain market approval. Article 2 (24) of the MDR defines the benefit-risk determination as “the analysis of all assessments of benefit and risk of possible relevance for the use of the device for the intended purpose when used in accordance with the intended purpose given by the manufacturer.”
A PMS system should clearly define benefit-risk calculations and the data used to support them. Post-market surveillance activities are critical in order to re-evaluate and maintain the benefit-risk calculations and determinations of a device throughout its life. Information that is gained through a PMS system can lead to:
• Identification of new risk factors.
• Adjustments to risk frequency and/or severity values based on actual use data.
• Adjustments to established risk calculations based on new “state of the art” technologies becoming available.
• Adjustments to established benefit calculations based on actual use data.
While complaint handling and other feedback tracking are more often described as part of post-market vigilance systems, they play a role in the more proactive post-market surveillance processes as well. A PMS system should define ...
To continue reading this ebook, download the full version.

How to Prevent Registration Deadline Misses in 2026
A missed registration deadline can halt sales, trigger regulatory penalties, and damage your organization's credibility with health authorities. For MedTech regulatory affairs teams managing hundreds or thousands of product registrations across dozens of markets, preventing deadline misses requires more than calendar reminders. It demands structured data, clear workflows, and real-time visibility into expiration timelines.
This guide explains why registration compliance deadlines are missed and how your team can prevent delays through stronger data control, workflow visibility, and process standardization.
Key Takeaways: How to Prevent Registration Deadline Misses in 2026
- Registration deadline misses typically stem from fragmented data, unclear ownership, and siloed communication between teams.
- Proactive renewal management requires real-time visibility into expiration dates across your entire global product portfolio.
- Standardized workflows with assigned accountability eliminate the confusion that leads to overlooked deadlines.
- Rimsys centralizes registration data and automates renewal alerts, reducing the risk of missed deadlines by up to 90%.
- Investing in a purpose-built RIM platform protects revenue continuity and maintains uninterrupted market access.
Why Do MedTech Teams Miss Registration Compliance Deadlines?
Registration deadline misses occur because the operating model cannot scale with growing portfolio complexity. Understanding root causes is the first step toward prevention.
Fragmented Data Across Systems and Regions
Many organizations track registrations using disconnected regional trackers, shared drives, and individual team member records. When product data lives in multiple locations, no single source of truth exists. This fragmentation makes it difficult to identify upcoming expirations before they become urgent.
A registration that expires in Brazil may be tracked in one system while the same product's EU approval is monitored elsewhere. Without connection between these records, teams lack portfolio-wide visibility into renewal timelines.
Unclear Ownership and Accountability
When responsibilities are distributed across regional leads, in-country partners, and distributors without clear accountability structures, deadlines fall through the cracks. Team members may assume someone else is handling a renewal, leading to inaction until the deadline passes.
Organizations with high turnover face additional risk. When key personnel leave, institutional knowledge about pending renewals often leaves with them.
Reactive Tracking Instead of Proactive Management
Calendar-based reminders and email notifications are inherently reactive. By the time a reminder appears, teams may not have enough runway to gather required documentation, coordinate with authorities, or address unexpected complications.
Effective deadline management requires proactive monitoring that surfaces upcoming expirations months in advance, not days before they occur.
What Happens When Registration Deadlines Are Missed?
The consequences of missed registration deadlines extend beyond administrative inconvenience. They create tangible business disruptions that affect revenue, reputation, and regulatory standing.
Loss of Market Access and Revenue Disruption
An expired registration means your product cannot legally be sold in that market. Regulatory teams must halt shipments, distribution partners cannot fulfill orders, and revenue stops flowing. For high-volume products or critical markets, even brief interruptions to the supply chain can translate to significant financial impact.
Re-registering a lapsed product often takes longer than a standard renewal. Your team may face additional scrutiny, updated documentation requirements, or queue delays that extend the time to market restoration.
Regulatory Penalties and Increased Scrutiny
Health authorities track compliance history. Missed deadlines signal operational weaknesses that can trigger increased inspection frequency, additional documentation requests, or enhanced scrutiny on future submissions. Building trust with regulators takes years; eroding it takes one avoidable miss.
Damage to Distributor and Partner Relationships
In-country partners and distributors depend on your organization to maintain valid registrations. When market access lapses, partners bear the burden of explaining delays to healthcare providers and end customers. Repeated misses can damage long-term business relationships and competitive positioning.
How to Identify Products at Risk of Deadline Misses
Prevention starts with identifying which registrations are most vulnerable. A systematic risk assessment helps teams prioritize attention and allocate resources effectively.
Create a Consolidated Registration Inventory
Bring all registration data into a single view. Include product identifiers, registration numbers, approval dates, expiration dates, responsible parties, and market-specific requirements. This inventory becomes your foundation for risk assessment.
Rimsys Registration Software centralizes this data automatically, linking registrations to products, submissions, and regulatory intelligence in one structured system.
Segment by Expiration Timeline and Complexity
Not all renewals carry equal risk. Categorize registrations by time remaining until expiration and by the complexity of renewal requirements. Products expiring within the next 90 days with extensive documentation needs should receive immediate attention.
Consider country-specific factors. Some markets require local testing, updated clinical data, or notified body involvement. These requirements extend lead times and increase the risk of delays.
Assess Documentation Readiness
For each at-risk registration, evaluate whether required documentation is current and accessible. Outdated technical files, missing test reports, or incomplete labeling information create bottlenecks that delay renewal submissions.
Documentation gaps discovered weeks before a deadline often cannot be addressed in time. Early assessment ensures enough runway to resolve issues.
Building a Proactive Renewal Management System
Moving from reactive tracking to proactive management requires intentional process design. The following components form the foundation of an effective renewal management system.
Establish Clear Ownership for Every Registration
Assign a single accountable owner for each registration. This person is responsible for monitoring expiration timelines, coordinating renewal activities, and escalating issues that require additional support.
Ownership should be documented within your registration management system, not stored in separate organizational charts or email threads. When someone leaves the organization, ownership transfer becomes straightforward.
Define Standard Lead Times by Market and Product Type
Different markets have different renewal timelines. EU MDR renewals involve notified body coordination. FDA establishment registrations follow annual cycles. ANVISA submissions may require specific local documentation.
Document the standard lead time needed for each market and product type. Use these lead times to trigger renewal workflows well before deadlines approach.
Implement Automated Expiration Monitoring
Replace manual calendar tracking with automated monitoring that surfaces upcoming expirations across your entire portfolio. Alerts should trigger at multiple intervals, such as 180 days, 90 days, 60 days, and 30 days before expiration.
Automated monitoring ensures that no registration is overlooked, regardless of portfolio size or team bandwidth. Rimsys Global Regulatory Compliance capabilities include lifecycle tracking and automated alerts that reduce missed renewal risk.
Standardizing Workflows to Prevent Oversight
Consistent workflows eliminate the variability that leads to deadlines. When every renewal follows the same structured process, teams can identify and address deviations before they cause problems.
Create Renewal Workflow Templates
Define the steps required for each type of renewal. Include documentation gathering, internal review, submission preparation, authority submission, and post-submission tracking. Assign responsible parties and expected durations for each step.
Templates ensure that new team members can execute renewals correctly without relying on undocumented institutional knowledge.
Build Escalation Paths for Delayed Activities
Not every renewal proceeds according to plan. Documentation may be delayed. Authority responses may take longer than expected. Partners may fail to submit required local information.
Define clear escalation paths that activate when activities fall behind schedule. Escalation should be automatic and visible, not dependent on individual team members raising concerns manually.
Conduct Regular Pipeline Reviews
Schedule recurring reviews of your renewal pipeline. During these reviews, examine registrations approaching expiration, identify any blocked activities, and confirm that responsible parties are on track.
Pipeline reviews surface potential issues early, when corrective action is still possible. They also reinforce accountability by making renewal status visible to leadership.
How Data Control Reduces Deadline Risk
Strong data control is the foundation of reliable deadline management. When registration data is accurate, current, and accessible, teams make better decisions and avoid preventable misses.
Maintain a Single Source of Truth
Eliminate competing data sources by establishing one authoritative system for registration information. All team members, partners, and stakeholders should access the same data, ensuring consistency and reducing confusion.
A single source of truth means that when someone asks about a product's registration status in a specific market, the answer is immediate and reliable. No cross-referencing multiple trackers. No waiting for colleagues to respond to email inquiries.
Connect Registration Data to Product and Submission Records
Registrations do not exist in isolation. They connect to specific products, rely on previous submissions, and may be affected by regulatory changes or product modifications. Systems that link these data types enable more accurate impact analysis.
When a product design change occurs, connected data helps teams identify which registrations may need updated. When regulations evolve, linked intelligence shows which markets and products are affected.
Ensure Version Control and Audit Readiness
Regulatory authorities expect organizations to demonstrate controlled processes. Maintain version history for all registration-related documentation. Track who made changes, when changes occurred, and what was modified.
Audit readiness built into daily operations means less scrambling when inspections occur. It also shows the historical context needed to understand how registration status evolved over time.
Leveraging Technology for Visibility and Automation
Technology plays a critical role in scaling deadline management across large portfolios. Purpose-built regulatory information management platforms offer capabilities that generic tools cannot match.
Dashboard Visibility Across Global Markets
Executive and operational dashboards display at-a-glance visibility into registration status across your entire portfolio. Filter by market, product line, expiration timeline, or responsible party, to focus on relevant subsets.
Dashboard visibility enables leadership to ask and answer questions quickly:
- How many registrations will expire in the next quarter?
- Which markets have the highest concentration of upcoming renewals?
- Are any renewals currently blocked?
Automated Alerts and Notifications
Configure alerts that notify responsible parties when action is required. Notifications should be specific, actionable, and timely. Rather than generic reminders, alerts should identify the specific registration, required action, and deadline.
Effective notification systems reduce the cognitive burden on team members. Instead of tracking deadlines mentally, they receive prompts when attention is needed.
Reporting for Continuous Improvement
Track metrics that indicate process health. Monitor the number of registrations renewed on time, average lead time utilization, and frequency of expedited renewals. Use these metrics to identify process weaknesses and drive improvement.
Organizations that measure renewal performance can demonstrate compliance maturity to regulators, partners, and investors. Metrics also help justify investments in process improvement.
How Rimsys Helps Prevent Registration Deadline Misses
Rimsys is the first and only holistic Regulatory Information Management software purpose-built for MedTech. It centralizes registrations, submissions, regulatory intelligence, and UDI data in one connected platform, giving teams the visibility and automation needed to prevent deadline misses.
Centralized Global Registration Tracking
Rimsys maintains a structured, product-centric data model that connects registrations to products, markets, and submissions. Teams gain real-time visibility into where every product can be sold, which registrations are pending, and which are approaching expiration.
Six of the world's top 12 MedTech manufacturers trust Rimsys to manage global regulatory operations, achieving up to 90% reduction in reporting effort and elimination of manual tracking risk.
Automated Lifecycle Alerts
Rimsys monitors registration lifecycles and automatically alerts teams to upcoming expirations, renewals, and information requests. Automated alerts ensure that no registration is overlooked, regardless of the portfolio size.
With Rimsys AI, teams can accelerate regulatory work through AI-assisted workflows that reduce repetitive tasks while keeping human judgment in control.
Connected Regulatory Intelligence
Registration deadlines do not exist in isolation from regulatory change. New requirements can affect renewal timelines, documentation needs, or market access conditions.
Rimsys connects registration data to regulatory intelligence and impact assessment workflows, helping teams understand how changes affect their portfolio before deadlines become urgent.
Creating a Culture of Deadline Accountability
Technology and process alone do not prevent deadline misses. Organizations must also cultivate a culture where deadline accountability is valued and reinforced.
Make Renewal Performance Visible
Share renewal metrics with teams and leadership regularly. Celebrate on-time renewals and analyze near-misses to identify improvement opportunities. Visibility creates accountability without requiring punitive measures.
Invest in Team Training and Development
Ensure that team members understand the importance of deadline management and have the skills to execute renewal workflows effectively. Training should cover both procedural requirements and the business impact of missed deadlines.
Support Cross-Functional Collaboration
Registrations often require input from quality, engineering, labeling, and commercial teams. Foster collaboration across functions to ensure that dependencies are identified early and addressed proactively.
When regulatory affairs teams operate in silos, they lack the information needed to anticipate complications. Cross-functional visibility enables earlier intervention.
In Conclusion: How to Prevent Registration Deadline Misses
Preventing registration deadline misses requires intentional effort across three dimensions: data control, workflow visibility, and process standardization. Organizations that invest in these areas protect revenue continuity, maintain regulatory standing, and preserve partner relationships.
The complexity of global regulatory operations is not slowing down. As portfolios expand and requirements evolve, teams need infrastructure that scales. Rimsys transforms regulatory operations from administrative tracking into strategic enablement, bringing speed, visibility, and confidence to global expansion.
Ready to eliminate deadline risk from your registration management? Speak with the Rimsys team to see how leading MedTech manufacturers manage global registrations with precision and control.
References:

How Smith & Nephew Repositioned Regulatory as a Strategic Commercial Partner
Smith & Nephew is a global medical device manufacturerwith a broad portfolio spanning orthopedics, sports medicine, and woundmanagement, sold and registered across markets worldwide. Before Rimsys,regulatory data was scattered across spreadsheets, shared drives, anddisconnected systems.
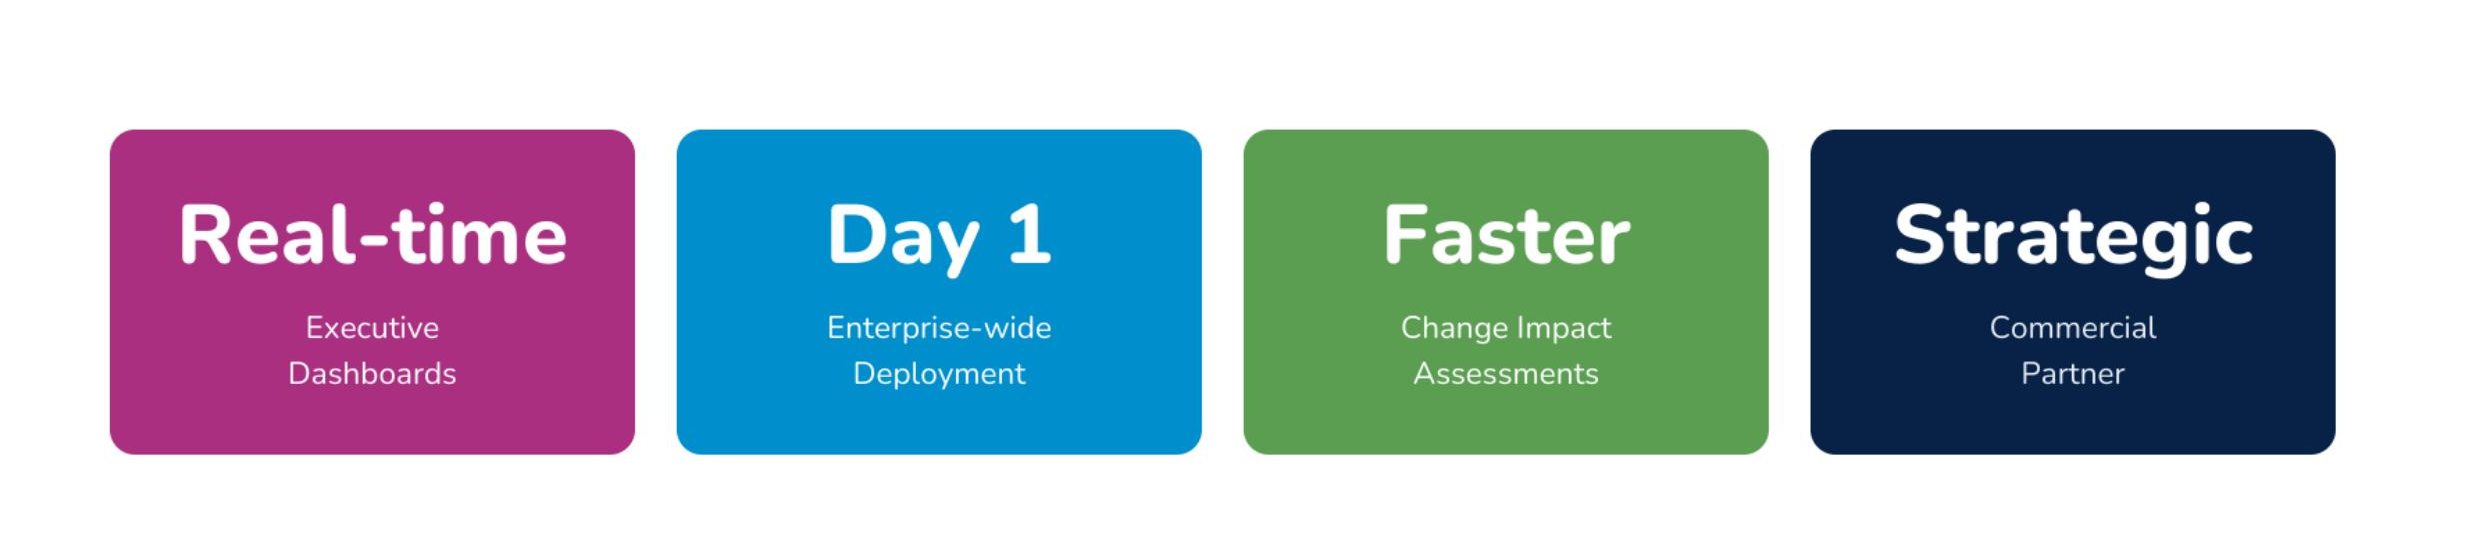
When Smith & Nephew selected Rimsys, they deployed enterprise-wide from day one. Executive reporting moved from manual fire drills to real-time dashboards. Change impact assessments became faster and more consistent. The regulatory team made the shift from reactive compliance function to strategic partner to the business.
The Challenge
Regulatory data at Smith & Nephew lived in multiplespreadsheets, shared drives, SharePoint sites, emails, and disconnectedsystems. Without a centralized record, the team could not reliably trackregistration timelines, measure on-time submissions, assess change impacts, orunderstand the downstream impact of product changes across markets. Preparingexecutive reporting meant manually assembling data from multiple sources, aprocess that consumed time and introduced risk each time.
The Solution
Smith & Nephew selected Rimsys for its configurable, notcustomized, platform: an intuitive user interface, centralized submissionmanagement, robust metrics, change assessment capabilities, and UDI supportwith machine-to-machine transmission. Rimsys’ interconnected modulearchitecture linked products, registrations, projects, change assessments, andUDI in a centralized location.
Rather than piloting in one business unit, Smith &Nephew deployed Rimsys across the entire regulatory organization from day one.The decision was deliberate: a partial deployment would have preserved thefragmentation. Enterprise-wide adoption established consistent metrics,standardized processes, and a single source of truth from the start.
The Results
Executive and board reporting, previously built from manualdata pulls, now flows directly from Rimsys in real time. What had been adisruptive, recurring effort is now a routine view. Leadership has thevisibility to make faster, more confident decisions, and the regulatory team isno longer pulled into reporting fire drills.
Change management has also been transformed. Direct linkagebetween products, registrations, and projects means impact assessments arefaster and less dependent on individual knowledge. UDI operations havesimilarly improved: machine-to-machine transmission has reduced manual uploadsand centralized DI record visibility supports global UDI requirements.
The most significant shift is strategic. With centralizedregulatory intelligence and real-time data, Smith & Nephew’s regulatoryteam now actively supports commercial planning: informing budget cycles,guiding renewal and launch sequencing, and advising on regulatory pathways toaccelerate market entry. Regulatory is no longer a downstream compliancefunction. It is a business partner.
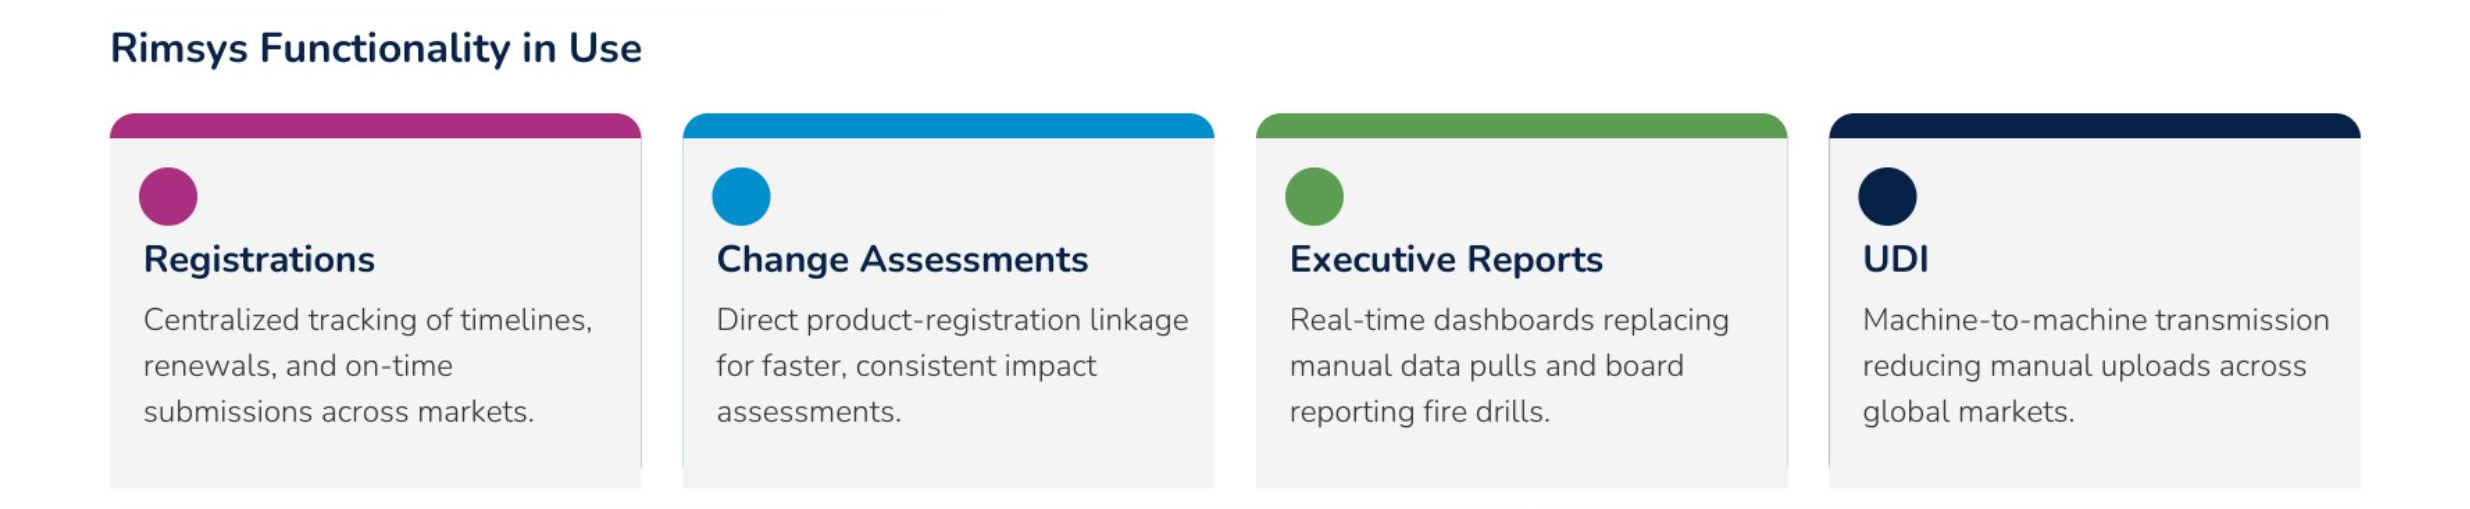
Smith & Nephew now runs four modules across its RIM operation:
- Registrations— Centralized license tracking across 250 countries and 30+ business units
- Change Assessments— Direct product-registration linkage for faster, consistent impact assessments
- Executive Reports— Real-time dashboards replacing manual data pulls and board reporting fire drills
- UDI— Machine-to-machine transmission reducing manual uploads across global markets
Take this to your team
If you’re evaluating how to modernize RIM operations at scale, the Smith & Nephew case study is a practical reference to share internally. It covers the full implementation story, module breakdown, and results data in a format built for stakeholder conversations.

How Philips Scaled Active Product Registrations More Than 20x
Philips Healthcare operates one of the largest regulatory portfolios in global MedTech: products registered across 250 countries, with a footprint that grows with every acquisition. Before Rimsys, that complexity was managed through email and spreadsheets. Submission packages moved through inboxes with no audit trail, no performance data, and no reliable view of where products were authorized to ship.
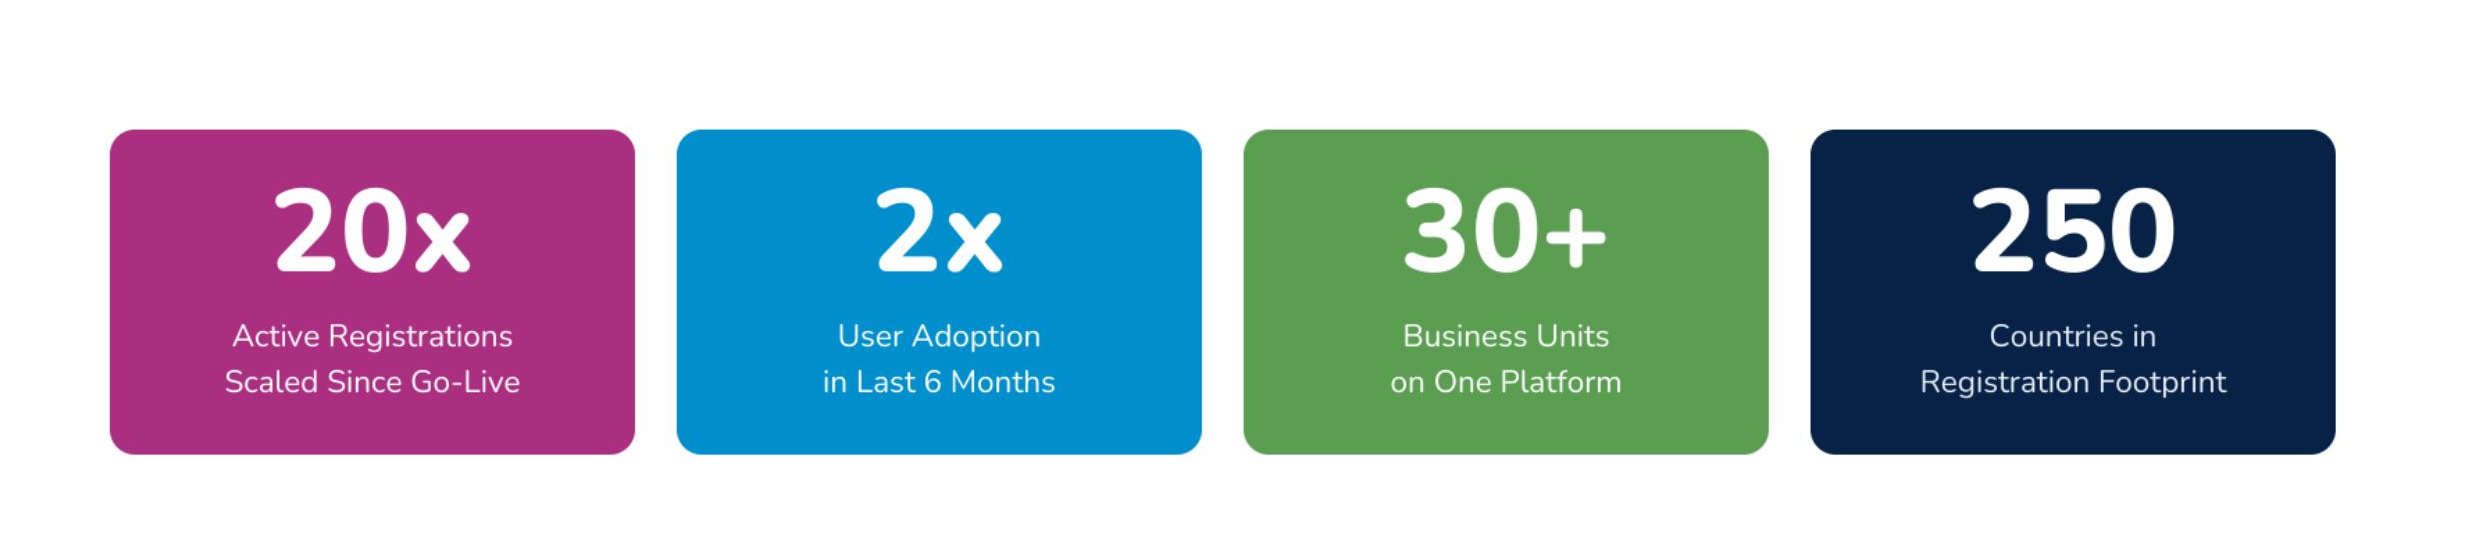
Philips selected Rimsys in 2022 as the enterprise RIM platform to bring regulatory order to that complexity. Since go-live, active product registrations have scaled more than 20x, user adoption has doubled in the last six months, and the regulatory affairs function now operates from a single source of truth spanning the entire enterprise.
The Challenge
Without structured data, Philips could not measure regulatory performance, track license expiration across the portfolio, or identify where submission work was stalling. Every acquisition made it worse: incoming business units arrived with their own workflows and systems, absorbing more fragmentation rather than resolving it.
The Solution
Philips evaluated multiple platforms against requirements built with both market-facing and business regulatory affairs teams. Rimsys won on two dimensions: an interface that made complex product and registration data immediately visible, and more enterprise-ready features than competing platforms at the right price point.
Philips went live with Rimsys Registrations and Submissions modules in July 2022. The team deployed platform experts for train-the-trainer sessions and launched regular drop-in sessions where users could ask questions and surface issues. Standing up a dedicated Regulatory Operations team focused exclusively on rest-of-world registration accelerated adoption further.
When an early business unit pushed back on workflow efficiency, Philips and Rimsys worked through it together. A hands-on process walkthrough identified exactly what needed to change, a resolution plan was shared, and that transparency and collaboration became the foundation for sustained user buy-in across the enterprise.
The Results
Since go-live, Philips has scaled active product registrations more than 20x, with further growth already underway. What started as a single deployment now spans 30+ business units across 250 countries, with Rimsys serving as the single source of truth for regulatory data across the enterprise, including businesses acquired since implementation.
For the first time, Philips can measure its own regulatory performance. KPIs flow directly from the platform, giving leadership real-time visibility into registration health. When anomalies surface, they drive data correction and user training, closing gaps that previously went undetected until they affected revenue.
Now with Rimsys AI-assisted Submissions and Regulatory Intelligence now in use, Philips expects to accelerate further: reducing administrative burden so skilled regulatory professionals can focus on strategy.
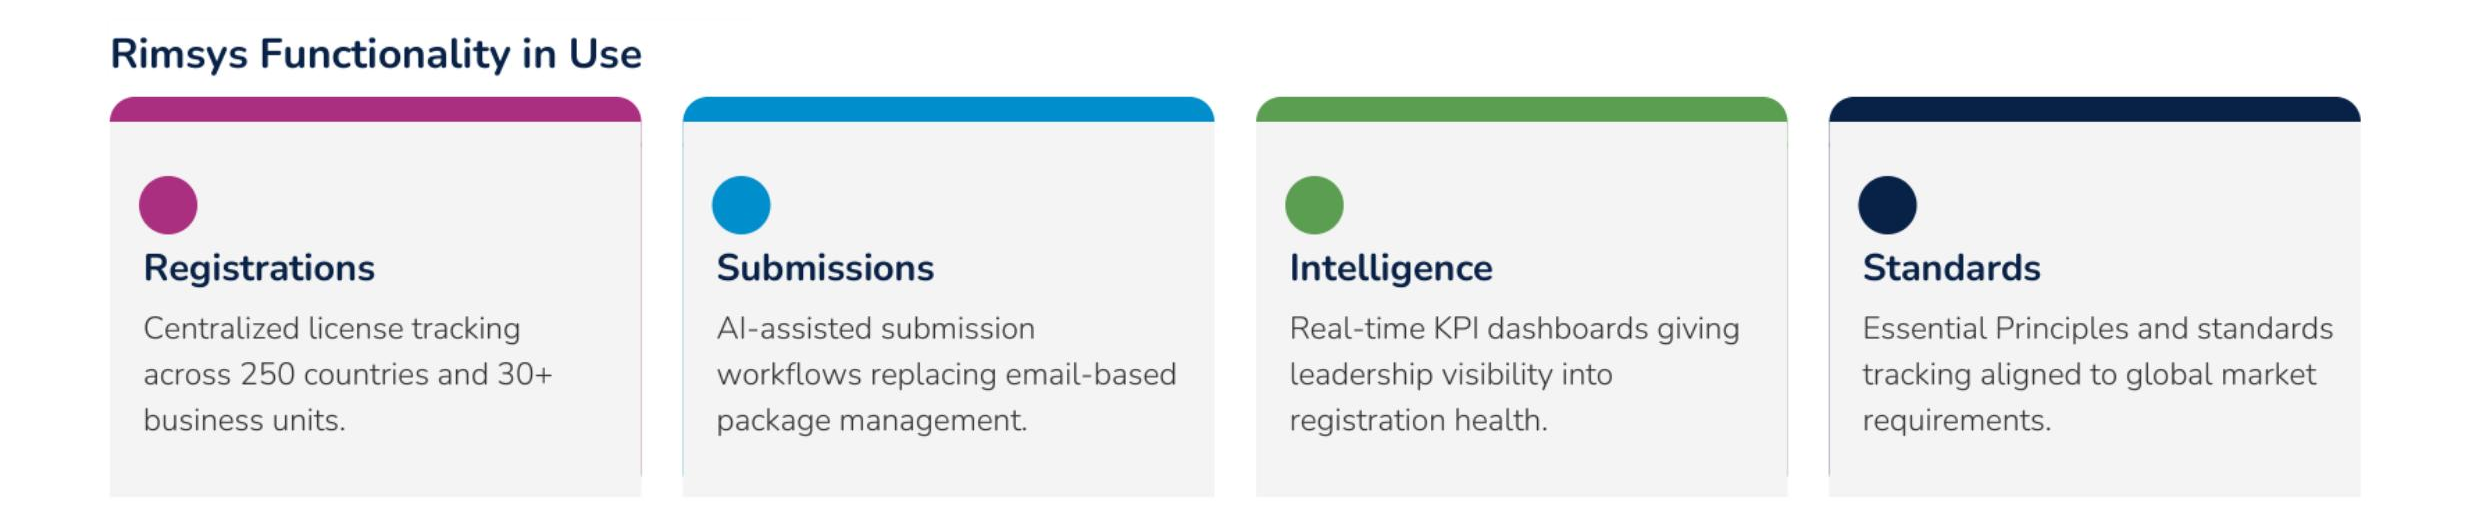
Philips now runs four modules across its RIM operation:
- Registrations— Centralized license tracking across 250 countries and 30+ business units
- Submissions— AI-assisted submission workflows replacing email-based package management
- Intelligence— Real-time KPI dashboards giving leadership visibility into registration health
- Standards— Essential Principles and standards tracking aligned to global market requirements
Take this to your team
If you’re evaluating how to modernize RIM operations at scale, the Philips Healthcare case study is a practical reference to share internally. It covers the full implementation story, module breakdown, and results data in a format built for stakeholder conversations.

What RAPS Euro Convergence 2026 Told Us About the Future of MedTech Regulation
Last week, the MedTech regulatory community gathered in Lisbon for RAPS Euro Convergence 2026: nearly 100 sessions, hundreds of professionals, and one overriding theme: transformation.The European regulatory landscape is shifting faster than it has in two decades, and the pressure is on every RA team to keep pace.
We were there. And here is what we took away.
The Dominant Signal: Change Is Accelerating
For MedTech manufacturers, the immediate reality is demanding. MDR 2.0 is advancing. The EU AI Act is creating new compliance obligations for software-enabled devices. EUDAMED continues to mature. And teams are being asked to absorb all of this while still meeting existing registration and renewal deadlines.
The practical implication is clear: RA functions that rely on manual tracking, disconnected spreadsheets, and tribal knowledge are being outrun by the pace of change. Across the industry, teams are moving from talking about AI to actively experimenting with it, using it to handle the volume and complexity that manual processes simply cannot absorb. The teams emerging as strategic forces are the ones who have connected, real-time regulatory infrastructure and are putting AI to work within it.
AI Is No Longer Optional Thinking
The conversation at Euro Convergence made one thing clear: AI has moved from future-state to present-tense. Regulatory professionals were encouraged to embrace AI while maintainingaccountability for the outcome and challenging the algorithms.
" Our role is to make sure that the AI does the right interpretations appropriate to our products, to our business."
— João Martins, Director of Regulatory Affairs at Abbott at RAPS Euro Convergence 2026 Opening Plenary
That framing resonates deeply with how we have built AI into Rimsys. The goal was never to replace regulatory judgment; it is to amplify it. Rimsys AI is domain-specific, built on the regulatory data structures and logic that reflect real-world requirements, country-specific nuances, and product context. It proposes, analyzes, and alerts. Your team reviews, approves, and decides.
For teams that are ready to accelerate, Rimsys AI accelerates regulatory intelligence monitoring and submission authoring, removing the repetitive, detail-heavy work so skilled professionals can focus on strategy, market expansion, and the higher-order decisions that increasingly complex regulations demand.
"As future regulators, we will need to be scientifically strong, comfortable with complexity, open to innovation, and also be able to work in increasingly complex environments."
— Rui Santos Ivo, President of Portugal's National Authority of Medicines and Health Products (INFARMED) and chair of the EMA management board, RAPS Euro Convergence 2026 Opening Plenary
MDR 2.0: Reform With Guardrails
A panel of experts representing regulators, industry, and notified bodies gave their views on the proposed revision of the EU Medical Device Regulation at the conference. While their sentiments were largely supportive, notified body representatives urged the European Commission to maintain proactive surveillance of devices to protect patients.
The discussion acknowledged the complexity of balancing reform with patient safety. Simplification and innovation go hand in hand, though if it is overly complicated or overly simplified, it becomes difficult to innovate. Structured dialogues in MDR/IVDR will provide transparency and predictability for manufacturers, especially in early product development.
Regulatory Workflows Cannot Be an Afterthought
A recurring observation across sessions was that MDR 2.0, EUDAMED, and the EU AI Act are only as effective as the operational workflows behind them. Structured dialogues, risk-proportionate pathways, and submissions all require teams to move quickly with accurate, up-to-date product data. That is simply not possible when that data lives across email threads, spreadsheets, and disconnected systems.
The workflows that came up most in Lisbon (change control, renewals, new product introductions, and registration management) are exactly the areas where manual processes create the most risk. A missed renewal. A design change that triggers 40 country-level impact assessments with no system to coordinate them. A registration record that no one has updated since the last audit.
Rimsys keeps these workflows connected and proactive. Renewal expiration reminders fire before deadlines become a risk. Change control impact surveys are configurable to your SOPs, so teams can assign tasks and coordinate work across regions without relying on someone to manually track progress. New product introductions move faster because previous submission content can be reused across markets. Target market data, registration history, and approval status are already centralized, so teams are building on existing work rather than starting from scratcheach time.
The result is regulatory operations that reduce time to market by weeks to months, not add to it. Access information in seconds rather than hours. Regulatory release authorization in minutes rather than weeks. More than 90% reduction in regional regulatory reporting time. These are not projections. They are outcomes reported by Rimsys customers operating in exactly the kind of complex, multi-market environments that dominated the conversation in Lisbon.
The Regulatory Professional Is Evolving
Perhaps the most striking thread across sessions was the evolution of the RA function itself. Regulatory work was once seen mainly in terms of compliance procedures and submissions. Today, the profession is much broader than that.
This evolution is exactly the transition Rimsys is designed to support. When regulatory data is centralized, connected, and visible in real time, RA teams stop spending their days chasing down registration status and start contributing to commercial strategy: market expansion decisions, launch sequencing, change control planning, and executive-level risk communication.
The heart of regulatory operations is not a filing cabinet. It is a living, connected system that elevates the entire function.
What It All Points To
RAPS Euro Convergence 2026 made one thing clear: the organizations that will thrive are those who have invested in regulatory infrastructure that can absorb change without breaking. Rimsys is the platform built for exactly this moment: enterprise-grade, intuitive enough for global teams to actually use, and trusted by 6 of the top 12 global MedTech manufacturers worldwide.

Rimsys Launches the Regulatory Execution Engine for MedTech
Spring 2026 embeds submission authoring, AI-powered regulatory monitoring, and configurable impact workflows inside a single RIM platform, the first step toward Rimsys' AI vision for global regulatory operations.
PITTSBURGH, PA, May 5, 2026 – Regulatory Information Management (RIM) software was built to store records. That foundation has served its purpose and reached its limit. Today, Rimsys announces the Spring 2026 release: a platform designed not to hold regulatory data, but to execute on it.
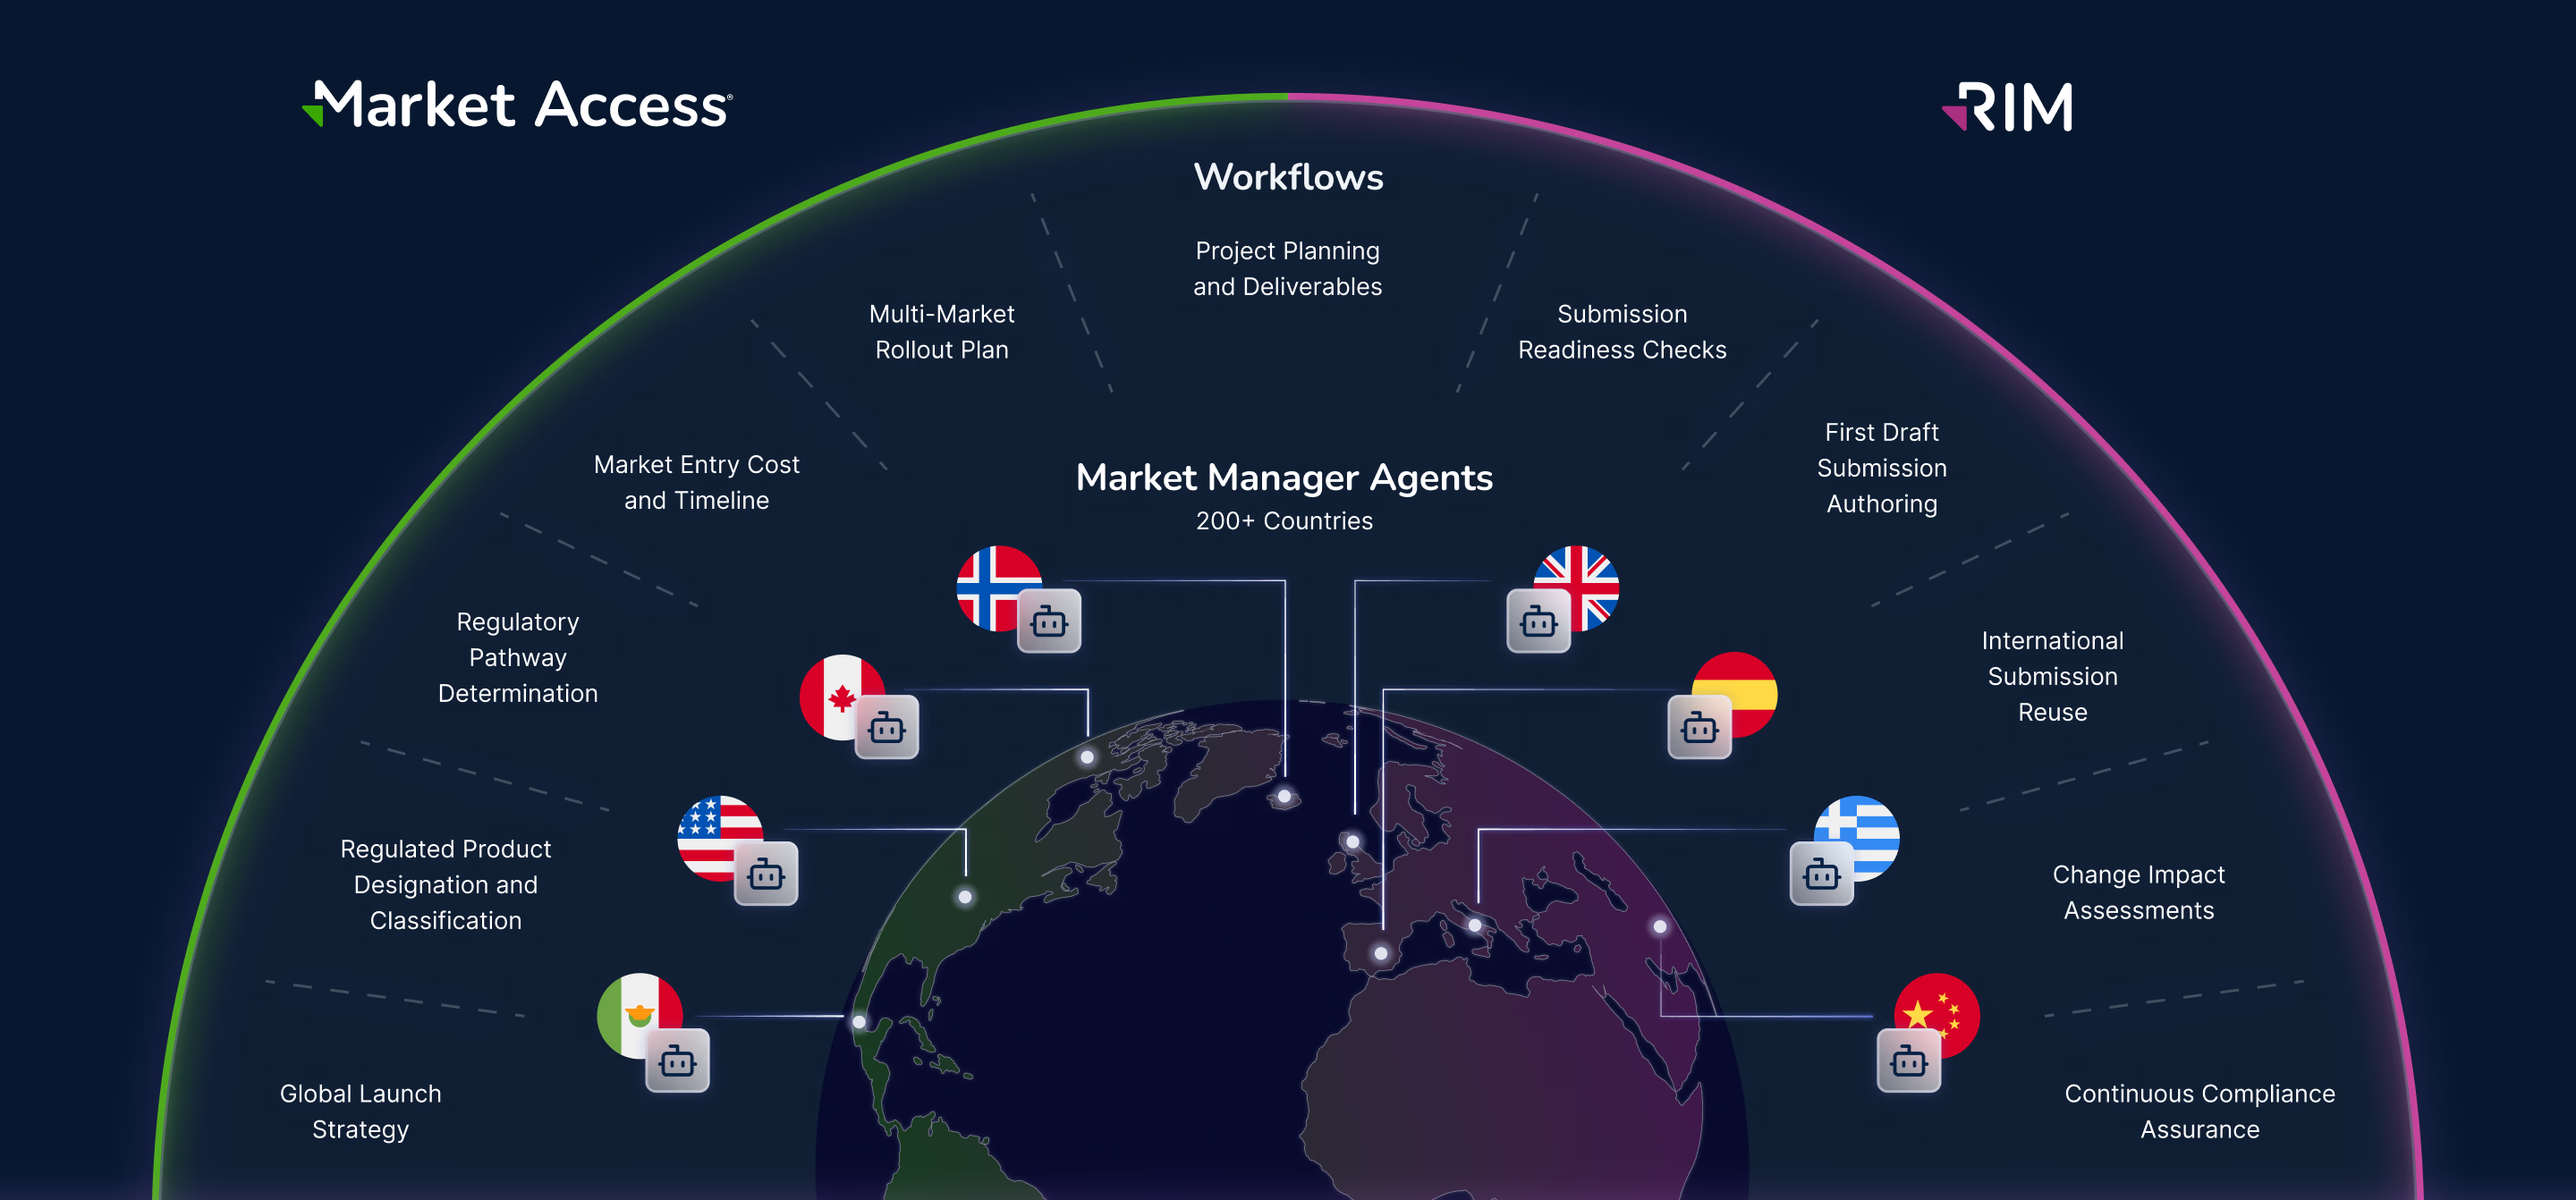
Submission volumes are growing. Markets are multiplying. Regulatory change is accelerating. Spring 2026 gives regulatory teams the tools to keep pace: embedded authoring, reusable submission content, configurable impact workflows, and AI-powered intelligence, all inside a single platform.
"Our vision for Rimsys is a platform that makes regulatory expertise go further, companies move faster, and products reach more markets than any team could accomplish alone. Spring 2026 is another meaningful step toward that vision. We are embedding the tools and intelligence that allow regulatory affairs professionals to operate at a different level, doing more strategic work, entering markets faster, and staying ahead of regulatory change rather than reacting to it. What we are building next makes this release the starting line." – James Gianoutsos, CEO
What Spring 2026 Delivers
A brand new website that provides in-depth information about the Rimsys offering and the benefits to MedTech manufacturers, including details on these new products:
Intelligence: AI-Powered Regulatory Monitoring
Rimsys Intelligence provides access to regulations, guidance documents, safety alerts, and legislation across more than 90 countries. AI triage and prioritization surface the updates most relevant to each customer's specific products and markets, eliminating hours of manual surveillance and putting the right information in front of the right people.
When a change requires action, teams can move directly from regulatory signal to impact assessment without a manual handoff. Intelligence represents Rimsys' first production deployment of context-aware AI operating across a customer's live regulatory data, a foundation that will expand significantly in future releases.
Advanced Submissions: A Unified Submission Execution Workflow
Advanced Submissions consolidates everything required to create, manage, and publish a regulatory submission into a single workflow inside Rimsys, eliminating the disconnected tools, manual reformatting, and version fragmentation that have defined submission work for too long. Three capabilities anchor it:
Rimsys Editor
The Rimsys Editor is the cornerstone of Advanced Submissions and the most significant capability in this release. It brings word-compatible authoring and editing natively inside Rimsys, fully compatible with Microsoft Word®, allowing regulatory teams to create, co-author, review, and publish submission content without leaving the platform for the first time.
The Editor supports real-time co-authoring, tracked changes and redlining, rich content including tables and images, document comparison, and PDF publishing with standardized headers, footers, and company branding applied automatically. AI-assisted authoring is available as a configurable option, enabling teams to summarize, refine, expand, and translate content within their workflow. Rimsys AI is human-in-the-loop by design.
Universal Submissions
Universal Submissions enables teams to build from a single universal template (an IMDRF Technical Document) with content automatically mapped into market-specific templates. One master structure, many markets, without rebuilding from scratch.
Reusable Submissions
Reusable Submissions takes a completed submission from one market and uses it as the starting point for a new one. The system automatically maps content into the target market's template, carrying applicable sections forward reducing the content creation time up to 90% and compressing the time required to enter each additional market.
Configurable Impact Surveys: Governed Change Assessment at Scale
Impact Surveys are now fully configurable. Templates can be defined for specific change event types, tied to countries or registrations, and triggered automatically from Rimsys Intelligence findings replacing ad hoc assessments with repeatable, governed workflows. This integration creates a direct line from change event to regulatory scope, with results tracked in a single audit-ready trail.
A Platform Built for What's Next
Spring 2026 establishes more than a set of new capabilities. It establishes the execution infrastructure, structured data model, and embedded AI foundation on which Rimsys' longer-term vision is being built.
That vision: a world where regulatory experts are amplified by intelligence, not constrained by information. Where the knowledge required to enter a new market, interpret a regulatory change, or scope a submission is instantly available to every member of the team. Where regulatory operations scale not by spreading experts thin, but by giving them tools that multiply their impact.
Spring is the first production step in that direction. Every submission authored inside the platform, every intelligence signal triaged by AI, and every impact assessment connected to structured regulatory data deepens the foundation. Future releases will build on it directly, expanding AI capabilities, automating more of the regulatory workflow, and ultimately enabling teams to do work that today requires external expertise to be done inside Rimsys.
Regulatory Execution as a Business Lever
Spring 2026 is built to move metrics that matter: reduced submission cycle time variance, improved approval predictability, lower marginal effort per market, and increased team capacity without proportional headcount growth. For executive leadership, earlier approvals translate directly into faster market access and accelerated revenue recognition.
Availability
Spring 2026 is now Generally Available. Existing customers on the Organizer product will retain access to their current experience.
To learn more about the Spring 2026 release and how Rimsys can accelerate your regulatory operations, visit rimsys.io or contact your Rimsys representative.
About Rimsys
Rimsys is the heart of regulatory operations for the medical device industry and the platform at the center of an AI-driven transformation in how regulated products reach global markets. A living, connected regulatory platform, Rimsys keeps regulatory intelligence, product data, approvals, and change management continuously connected, enabling organizations to expand into global markets with speed, precision, and confidence. Enterprise-ready yet intuitive to use, Rimsys is trusted by 6 of the top 12 global MedTech manufacturers to accelerate time to market and scale regulatory operations worldwide. To learn more, visit rimsys.io.
Media Contact
rimsys.io
The Real Cost of “We’ll Build It Ourselves”
If you are reading this from inside a large MedTech organization, you may be thinking: we have ten times the engineering staff. Why can’t we just build this ourselves?
We-Should-Just-Build-This-Ourse…
It is a fair question.
But software has a well-known paradox. Adding more people to a complex project does not make it go faster. It usually makes it go slower. More coordination. More handoffs. More meetings about meetings. More surface area for misalignment
A large IT organization is optimized for breadth — supporting dozens of systems, managing infrastructure, keeping the lights on across the enterprise
That is valuable work.
But it is fundamentally different from building and sustaining a deep vertical product over a decade.
The people on your team have day jobs. They run devices through regulatory pathways, manage quality systems, support manufacturing, and commercialize products globally
Building a regulated platform is not a side quest.
It is a second company
What the Numbers Actually Look Like
When people compare license fees to internal builds, they stop at the wrong baseline
The real comparison is:
Licensing a specialized platform
versus
Standing up and operating a regulated software company inside your enterprise
Product management.
UX research.
Engineering.
Regulatory SMEs.
Validation and QA.
Security operations.
Compliance programs.
24/7 support.
Infrastructure.
Multi-year modernization
AI makes some of that faster.
It does not make any of it optional
With a specialized vendor, that investment is amortized across an entire customer base.
With an internal build, the full long tail of ownership falls on you
And most of that spend ends up recreating the 80 percent that has already been solved — all because someone decided the remaining 20 percent justified building from scratch
The return on that 20 percent rarely survives honest scrutiny.
The Questions That Should Keep You Honest
It is easy to get excited about how fast something can be built.
The harder exercise is asking what happens in year three, year five, year eight
When your VP of Regulatory Affairs leaves, who maintains validation documentation?
When regulations change across jurisdictions simultaneously, who redesigns workflows and pushes a validated release before the deadline?
When an auditor asks for change control history and disaster recovery test results, who is accountable?
Internal initiatives often stumble not because engineers cannot prototype, but because sustaining them for a decade is brutally hard
Sponsors move on. Budgets change. Teams reorganize.
Regulatory systems do not get to pause.
They must remain inspection-ready through acquisitions, divestitures, and leadership turnover
Systems of record are commitments, not experiments
AI Changed the Tools, Not the Gravity
I am genuinely excited about what AI enables. It will reshape regulatory operations, reduce headcount growth, compress timelines, and raise expectations for every vendor in this space
What it has not done is repeal gravity.
Most of what AI replaces today is busy work. That is enormously valuable. But busy work was never the strategic bottleneck
The hard parts remain.
- Deciding submission strategy.
- Interpreting regulator feedback.
- Designing defensible workflows.
- Staying inspection-ready.
- Running global rollouts
Agents help teams move faster.
They do not decide what is safe, defensible, or durable
In MedTech, software is not just built.
It is designed, governed, operated, and defended
And gravity still applies.